Jump to
S1C2
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -477.254642 |
| Energy at 298.15K | |
| HF Energy | -476.762525 |
| Nuclear repulsion energy | 106.453238 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3135 |
2988 |
|
|
|
|
| 2 |
A' |
3067 |
2923 |
|
|
|
|
| 3 |
A' |
3047 |
2904 |
|
|
|
|
| 4 |
A' |
2714 |
2586 |
|
|
|
|
| 5 |
A' |
1493 |
1423 |
|
|
|
|
| 6 |
A' |
1482 |
1413 |
|
|
|
|
| 7 |
A' |
1404 |
1339 |
|
|
|
|
| 8 |
A' |
1298 |
1237 |
|
|
|
|
| 9 |
A' |
1114 |
1062 |
|
|
|
|
| 10 |
A' |
1004 |
957 |
|
|
|
|
| 11 |
A' |
864 |
823 |
|
|
|
|
| 12 |
A' |
680 |
649 |
|
|
|
|
| 13 |
A' |
304 |
289 |
|
|
|
|
| 14 |
A" |
3142 |
2995 |
|
|
|
|
| 15 |
A" |
3117 |
2971 |
|
|
|
|
| 16 |
A" |
1482 |
1413 |
|
|
|
|
| 17 |
A" |
1263 |
1204 |
|
|
|
|
| 18 |
A" |
1041 |
993 |
|
|
|
|
| 19 |
A" |
785 |
748 |
|
|
|
|
| 20 |
A" |
261 |
249 |
|
|
|
|
| 21 |
A" |
178 |
170 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16437.2 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 15666.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.527 |
0.693 |
0.000 |
| C2 |
0.000 |
0.837 |
0.000 |
| S3 |
-0.762 |
-0.845 |
0.000 |
| H4 |
2.005 |
1.691 |
0.000 |
| H5 |
1.871 |
0.144 |
0.896 |
| H6 |
1.871 |
0.144 |
-0.896 |
| H7 |
-0.336 |
1.389 |
0.897 |
| H8 |
-0.336 |
1.389 |
-0.897 |
| H9 |
-2.049 |
-0.425 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5339 | 2.7578 | 1.1061 | 1.1061 | 1.1061 | 2.1816 | 2.1816 | 3.7469 |
C2 | 1.5339 | | 1.8464 | 2.1794 | 2.1875 | 2.1875 | 1.1055 | 1.1055 | 2.4065 | S3 | 2.7578 | 1.8464 | | 3.7532 | 2.9517 | 2.9517 | 2.4447 | 2.4447 | 1.3539 | H4 | 1.1061 | 2.1794 | 3.7532 | | 1.7930 | 1.7930 | 2.5250 | 2.5250 | 4.5731 | H5 | 1.1061 | 2.1875 | 2.9517 | 1.7930 | | 1.7923 | 2.5341 | 3.1044 | 4.0613 | H6 | 1.1061 | 2.1875 | 2.9517 | 1.7930 | 1.7923 | | 3.1044 | 2.5341 | 4.0613 | H7 | 2.1816 | 1.1055 | 2.4447 | 2.5250 | 2.5341 | 3.1044 | | 1.7941 | 2.6515 | H8 | 2.1816 | 1.1055 | 2.4447 | 2.5250 | 3.1044 | 2.5341 | 1.7941 | | 2.6515 | H9 | 3.7469 | 2.4065 | 1.3539 | 4.5731 | 4.0613 | 4.0613 | 2.6515 | 2.6515 | |
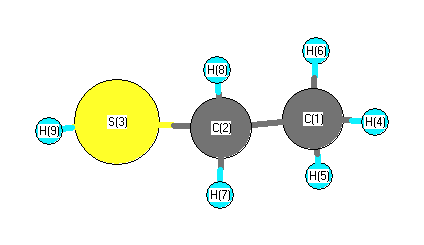 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
108.990 |
|
C1 |
C2 |
H7 |
110.437 |
| C1 |
C2 |
H8 |
110.437 |
|
C2 |
C1 |
H4 |
110.227 |
| C2 |
C1 |
H5 |
110.866 |
|
C2 |
C1 |
H6 |
110.866 |
| C2 |
S3 |
H9 |
96.302 |
|
S3 |
C2 |
H7 |
109.237 |
| S3 |
C2 |
H8 |
109.237 |
|
H4 |
C1 |
H5 |
108.286 |
| H4 |
C1 |
H6 |
108.286 |
|
H5 |
C1 |
H6 |
108.220 |
| H7 |
C2 |
H8 |
108.478 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -477.255766 |
| Energy at 298.15K | -477.262095 |
| HF Energy | -476.763240 |
| Nuclear repulsion energy | 106.233842 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3145 |
2997 |
|
|
|
|
| 2 |
A |
3125 |
2978 |
|
|
|
|
| 3 |
A |
3115 |
2969 |
|
|
|
|
| 4 |
A |
3068 |
2924 |
|
|
|
|
| 5 |
A |
3040 |
2897 |
|
|
|
|
| 6 |
A |
2705 |
2578 |
|
|
|
|
| 7 |
A |
1487 |
1417 |
|
|
|
|
| 8 |
A |
1481 |
1411 |
|
|
|
|
| 9 |
A |
1468 |
1400 |
|
|
|
|
| 10 |
A |
1403 |
1337 |
|
|
|
|
| 11 |
A |
1303 |
1242 |
|
|
|
|
| 12 |
A |
1275 |
1215 |
|
|
|
|
| 13 |
A |
1120 |
1068 |
|
|
|
|
| 14 |
A |
1072 |
1022 |
|
|
|
|
| 15 |
A |
995 |
948 |
|
|
|
|
| 16 |
A |
869 |
828 |
|
|
|
|
| 17 |
A |
734 |
700 |
|
|
|
|
| 18 |
A |
666 |
635 |
|
|
|
|
| 19 |
A |
330 |
315 |
|
|
|
|
| 20 |
A |
270 |
258 |
|
|
|
|
| 21 |
A |
218 |
208 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16444.4 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 15673.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.647 |
-0.355 |
-0.054 |
| C2 |
0.501 |
0.651 |
0.090 |
| S3 |
-1.172 |
-0.102 |
-0.081 |
| H4 |
2.626 |
0.161 |
0.008 |
| H5 |
1.609 |
-1.114 |
0.752 |
| H6 |
1.586 |
-0.882 |
-1.023 |
| H7 |
0.560 |
1.191 |
1.053 |
| H8 |
0.545 |
1.411 |
-0.714 |
| H9 |
-1.067 |
-0.912 |
1.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5325 | 2.8306 | 1.1077 | 1.1069 | 1.1052 | 2.1904 | 2.1843 | 2.9649 |
C2 | 1.5325 | | 1.8424 | 2.1823 | 2.1866 | 2.1838 | 1.1057 | 1.1065 | 2.3943 | S3 | 2.8306 | 1.8424 | | 3.8075 | 3.0739 | 3.0169 | 2.4411 | 2.3738 | 1.3549 | H4 | 1.1077 | 2.1823 | 3.8075 | | 1.7917 | 1.7980 | 2.5335 | 2.5328 | 3.9715 | H5 | 1.1069 | 2.1866 | 3.0739 | 1.7917 | | 1.7899 | 2.5500 | 3.1069 | 2.6953 | H6 | 1.1052 | 2.1838 | 3.0169 | 1.7980 | 1.7899 | | 3.1084 | 2.5377 | 3.3367 | H7 | 2.1904 | 1.1057 | 2.4411 | 2.5335 | 2.5500 | 3.1084 | | 1.7803 | 2.6603 | H8 | 2.1843 | 1.1065 | 2.3738 | 2.5328 | 3.1069 | 2.5377 | 1.7803 | | 3.3063 | H9 | 2.9649 | 2.3943 | 1.3549 | 3.9715 | 2.6953 | 3.3367 | 2.6603 | 3.3063 | |
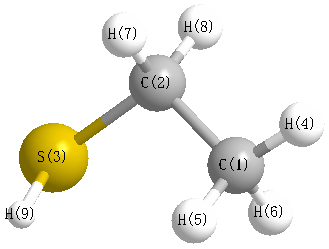 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
113.698 |
|
C1 |
C2 |
H7 |
111.217 |
| C1 |
C2 |
H8 |
110.689 |
|
C2 |
C1 |
H4 |
110.460 |
| C2 |
C1 |
H5 |
110.850 |
|
C2 |
C1 |
H6 |
110.726 |
| C2 |
S3 |
H9 |
95.776 |
|
S3 |
C2 |
H7 |
109.230 |
| S3 |
C2 |
H8 |
104.435 |
|
H4 |
C1 |
H5 |
108.005 |
| H4 |
C1 |
H6 |
108.684 |
|
H5 |
C1 |
H6 |
108.023 |
| H7 |
C2 |
H8 |
107.175 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
