Jump to
S1C2
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -477.241516 |
| Energy at 298.15K | -477.247801 |
| HF Energy | -476.762745 |
| Nuclear repulsion energy | 106.607952 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3147 |
3019 |
23.16 |
|
|
|
| 2 |
A' |
3083 |
2958 |
24.55 |
|
|
|
| 3 |
A' |
3061 |
2937 |
22.25 |
|
|
|
| 4 |
A' |
2730 |
2619 |
12.71 |
|
|
|
| 5 |
A' |
1503 |
1442 |
1.26 |
|
|
|
| 6 |
A' |
1494 |
1434 |
2.00 |
|
|
|
| 7 |
A' |
1418 |
1360 |
1.85 |
|
|
|
| 8 |
A' |
1313 |
1259 |
35.40 |
|
|
|
| 9 |
A' |
1125 |
1079 |
1.57 |
|
|
|
| 10 |
A' |
1014 |
973 |
1.90 |
|
|
|
| 11 |
A' |
875 |
840 |
1.62 |
|
|
|
| 12 |
A' |
691 |
663 |
0.86 |
|
|
|
| 13 |
A' |
308 |
295 |
1.89 |
|
|
|
| 14 |
A" |
3155 |
3027 |
34.27 |
|
|
|
| 15 |
A" |
3132 |
3005 |
1.49 |
|
|
|
| 16 |
A" |
1492 |
1431 |
6.29 |
|
|
|
| 17 |
A" |
1274 |
1222 |
0.58 |
|
|
|
| 18 |
A" |
1052 |
1009 |
0.40 |
|
|
|
| 19 |
A" |
793 |
761 |
2.13 |
|
|
|
| 20 |
A" |
265 |
254 |
1.01 |
|
|
|
| 21 |
A" |
184 |
177 |
17.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16553.2 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 15881.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
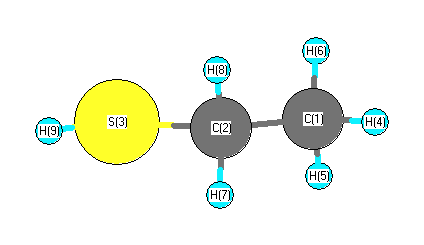
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.526 |
0.693 |
0.000 |
| C2 |
0.000 |
0.834 |
0.000 |
| S3 |
-0.761 |
-0.843 |
0.000 |
| H4 |
2.001 |
1.691 |
0.000 |
| H5 |
1.871 |
0.145 |
0.895 |
| H6 |
1.871 |
0.145 |
-0.895 |
| H7 |
-0.334 |
1.387 |
0.895 |
| H8 |
-0.334 |
1.387 |
-0.895 |
| H9 |
-2.048 |
-0.427 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5321 | 2.7549 | 1.1049 | 1.1050 | 1.1050 | 2.1778 | 2.1778 | 3.7452 |
C2 | 1.5321 | | 1.8419 | 2.1762 | 2.1859 | 2.1859 | 1.1041 | 1.1041 | 2.4053 | S3 | 2.7549 | 1.8419 | | 3.7480 | 2.9511 | 2.9511 | 2.4408 | 2.4408 | 1.3523 | H4 | 1.1049 | 2.1762 | 3.7480 | | 1.7907 | 1.7907 | 2.5190 | 2.5190 | 4.5691 | H5 | 1.1050 | 2.1859 | 2.9511 | 1.7907 | | 1.7901 | 2.5315 | 3.1005 | 4.0610 | H6 | 1.1050 | 2.1859 | 2.9511 | 1.7907 | 1.7901 | | 3.1005 | 2.5315 | 4.0610 | H7 | 2.1778 | 1.1041 | 2.4408 | 2.5190 | 2.5315 | 3.1005 | | 1.7903 | 2.6515 | H8 | 2.1778 | 1.1041 | 2.4408 | 2.5190 | 3.1005 | 2.5315 | 1.7903 | | 2.6515 | H9 | 3.7452 | 2.4053 | 1.3523 | 4.5691 | 4.0610 | 4.0610 | 2.6515 | 2.6515 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
109.127 |
|
C1 |
C2 |
H7 |
110.345 |
| C1 |
C2 |
H8 |
110.345 |
|
C2 |
C1 |
H4 |
110.174 |
| C2 |
C1 |
H5 |
110.937 |
|
C2 |
C1 |
H6 |
110.937 |
| C2 |
S3 |
H9 |
96.503 |
|
S3 |
C2 |
H7 |
109.328 |
| S3 |
C2 |
H8 |
109.328 |
|
H4 |
C1 |
H5 |
108.251 |
| H4 |
C1 |
H6 |
108.251 |
|
H5 |
C1 |
H6 |
108.195 |
| H7 |
C2 |
H8 |
108.347 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -477.242555 |
| Energy at 298.15K | -477.248910 |
| HF Energy | -476.763489 |
| Nuclear repulsion energy | 106.355356 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3158 |
3030 |
30.04 |
|
|
|
| 2 |
A |
3138 |
3011 |
18.04 |
|
|
|
| 3 |
A |
3130 |
3003 |
13.89 |
|
|
|
| 4 |
A |
3086 |
2960 |
18.99 |
|
|
|
| 5 |
A |
3055 |
2931 |
26.14 |
|
|
|
| 6 |
A |
2723 |
2613 |
11.93 |
|
|
|
| 7 |
A |
1497 |
1436 |
1.72 |
|
|
|
| 8 |
A |
1490 |
1430 |
7.34 |
|
|
|
| 9 |
A |
1480 |
1420 |
0.51 |
|
|
|
| 10 |
A |
1417 |
1359 |
1.40 |
|
|
|
| 11 |
A |
1317 |
1264 |
22.33 |
|
|
|
| 12 |
A |
1285 |
1233 |
2.23 |
|
|
|
| 13 |
A |
1131 |
1085 |
6.67 |
|
|
|
| 14 |
A |
1082 |
1038 |
0.69 |
|
|
|
| 15 |
A |
1005 |
964 |
4.19 |
|
|
|
| 16 |
A |
878 |
842 |
5.46 |
|
|
|
| 17 |
A |
741 |
711 |
1.21 |
|
|
|
| 18 |
A |
675 |
648 |
2.83 |
|
|
|
| 19 |
A |
333 |
319 |
1.62 |
|
|
|
| 20 |
A |
273 |
262 |
1.94 |
|
|
|
| 21 |
A |
220 |
211 |
15.51 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16556.4 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 15884.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
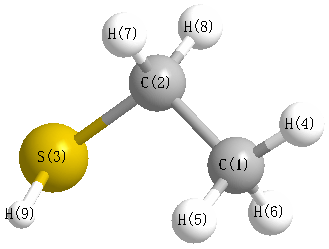
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.647 |
-0.354 |
-0.053 |
| C2 |
0.500 |
0.649 |
0.090 |
| S3 |
-1.171 |
-0.102 |
-0.081 |
| H4 |
2.623 |
0.162 |
0.018 |
| H5 |
1.606 |
-1.117 |
0.746 |
| H6 |
1.593 |
-0.875 |
-1.025 |
| H7 |
0.558 |
1.187 |
1.053 |
| H8 |
0.544 |
1.409 |
-0.711 |
| H9 |
-1.076 |
-0.911 |
0.999 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5303 | 2.8286 | 1.1065 | 1.1055 | 1.1040 | 2.1871 | 2.1813 | 2.9713 |
C2 | 1.5303 | | 1.8396 | 2.1790 | 2.1846 | 2.1820 | 1.1041 | 1.1050 | 2.3965 | S3 | 2.8286 | 1.8396 | | 3.8037 | 3.0695 | 3.0210 | 2.4358 | 2.3704 | 1.3531 | H4 | 1.1065 | 2.1790 | 3.8037 | | 1.7891 | 1.7956 | 2.5267 | 2.5314 | 3.9739 | H5 | 1.1055 | 2.1846 | 3.0695 | 1.7891 | | 1.7874 | 2.5497 | 3.1035 | 2.7011 | H6 | 1.1040 | 2.1820 | 3.0210 | 1.7956 | 1.7874 | | 3.1048 | 2.5331 | 3.3499 | H7 | 2.1871 | 1.1041 | 2.4358 | 2.5267 | 2.5497 | 3.1048 | | 1.7775 | 2.6592 | H8 | 2.1813 | 1.1050 | 2.3704 | 2.5314 | 3.1035 | 2.5331 | 1.7775 | | 3.3061 | H9 | 2.9713 | 2.3965 | 1.3531 | 3.9739 | 2.7011 | 3.3499 | 2.6592 | 3.3061 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
113.834 |
|
C1 |
C2 |
H7 |
111.206 |
| C1 |
C2 |
H8 |
110.691 |
|
C2 |
C1 |
H4 |
110.423 |
| C2 |
C1 |
H5 |
110.921 |
|
C2 |
C1 |
H6 |
110.812 |
| C2 |
S3 |
H9 |
96.090 |
|
S3 |
C2 |
H7 |
109.112 |
| S3 |
C2 |
H8 |
104.440 |
|
H4 |
C1 |
H5 |
107.955 |
| H4 |
C1 |
H6 |
108.641 |
|
H5 |
C1 |
H6 |
107.991 |
| H7 |
C2 |
H8 |
107.151 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
