Jump to
S1C2
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -477.722090 |
| Energy at 298.15K | -477.728183 |
| HF Energy | -477.722090 |
| Nuclear repulsion energy | 106.280251 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3057 |
3039 |
16.28 |
|
|
|
| 2 |
A' |
2987 |
2970 |
23.28 |
|
|
|
| 3 |
A' |
2974 |
2957 |
19.77 |
|
|
|
| 4 |
A' |
2608 |
2593 |
8.47 |
|
|
|
| 5 |
A' |
1427 |
1419 |
2.26 |
|
|
|
| 6 |
A' |
1414 |
1406 |
1.06 |
|
|
|
| 7 |
A' |
1343 |
1335 |
3.06 |
|
|
|
| 8 |
A' |
1232 |
1224 |
38.31 |
|
|
|
| 9 |
A' |
1079 |
1073 |
0.93 |
|
|
|
| 10 |
A' |
976 |
970 |
4.97 |
|
|
|
| 11 |
A' |
827 |
822 |
0.84 |
|
|
|
| 12 |
A' |
654 |
650 |
1.51 |
|
|
|
| 13 |
A' |
291 |
289 |
2.36 |
|
|
|
| 14 |
A" |
3066 |
3048 |
25.09 |
|
|
|
| 15 |
A" |
3037 |
3019 |
1.96 |
|
|
|
| 16 |
A" |
1413 |
1405 |
8.01 |
|
|
|
| 17 |
A" |
1214 |
1207 |
0.92 |
|
|
|
| 18 |
A" |
994 |
988 |
0.17 |
|
|
|
| 19 |
A" |
765 |
760 |
4.54 |
|
|
|
| 20 |
A" |
251 |
249 |
0.47 |
|
|
|
| 21 |
A" |
143 |
142 |
17.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15874.5 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 15782.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.521 |
0.706 |
0.000 |
| C2 |
0.000 |
0.838 |
0.000 |
| S3 |
-0.757 |
-0.852 |
0.000 |
| H4 |
1.992 |
1.710 |
0.000 |
| H5 |
1.878 |
0.161 |
0.897 |
| H6 |
1.878 |
0.161 |
-0.897 |
| H7 |
-0.347 |
1.385 |
0.900 |
| H8 |
-0.347 |
1.385 |
-0.900 |
| H9 |
-2.060 |
-0.438 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5264 | 2.7600 | 1.1091 | 1.1090 | 1.1090 | 2.1814 | 2.1814 | 3.7593 |
C2 | 1.5264 | | 1.8516 | 2.1748 | 2.1886 | 2.1886 | 1.1087 | 1.1087 | 2.4232 | S3 | 2.7600 | 1.8516 | | 3.7582 | 2.9625 | 2.9625 | 2.4456 | 2.4456 | 1.3672 | H4 | 1.1091 | 2.1748 | 3.7582 | | 1.7938 | 1.7938 | 2.5272 | 2.5272 | 4.5865 | H5 | 1.1090 | 2.1886 | 2.9625 | 1.7938 | | 1.7942 | 2.5396 | 3.1109 | 4.0834 | H6 | 1.1090 | 2.1886 | 2.9625 | 1.7938 | 1.7942 | | 3.1109 | 2.5396 | 4.0834 | H7 | 2.1814 | 1.1087 | 2.4456 | 2.5272 | 2.5396 | 3.1109 | | 1.7992 | 2.6583 | H8 | 2.1814 | 1.1087 | 2.4456 | 2.5272 | 3.1109 | 2.5396 | 1.7992 | | 2.6583 | H9 | 3.7593 | 2.4232 | 1.3672 | 4.5865 | 4.0834 | 4.0834 | 2.6583 | 2.6583 | |
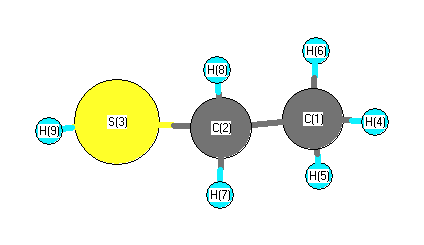 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
109.201 |
|
C1 |
C2 |
H7 |
110.760 |
| C1 |
C2 |
H8 |
110.760 |
|
C2 |
C1 |
H4 |
110.210 |
| C2 |
C1 |
H5 |
111.311 |
|
C2 |
C1 |
H6 |
111.311 |
| C2 |
S3 |
H9 |
96.510 |
|
S3 |
C2 |
H7 |
108.803 |
| S3 |
C2 |
H8 |
108.803 |
|
H4 |
C1 |
H5 |
107.944 |
| H4 |
C1 |
H6 |
107.944 |
|
H5 |
C1 |
H6 |
107.985 |
| H7 |
C2 |
H8 |
108.468 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.065 |
|
|
|
| 2 |
C |
-0.171 |
|
|
|
| 3 |
S |
-0.121 |
|
|
|
| 4 |
H |
0.043 |
|
|
|
| 5 |
H |
0.050 |
|
|
|
| 6 |
H |
0.050 |
|
|
|
| 7 |
H |
0.065 |
|
|
|
| 8 |
H |
0.065 |
|
|
|
| 9 |
H |
0.083 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.037 |
1.578 |
0.000 |
1.579 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.608 |
-0.152 |
0.000 |
| y |
-0.152 |
-27.971 |
0.000 |
| z |
0.000 |
0.000 |
-28.686 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.720 |
-0.152 |
0.000 |
| y |
-0.152 |
-1.324 |
0.000 |
| z |
0.000 |
0.000 |
-2.396 |
|
| Polar |
|---|
| 3z2-r2 | -4.793 |
|---|
| x2-y2 | 3.363 |
|---|
| xy | -0.152 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.961 |
0.861 |
0.000 |
| y |
0.861 |
6.318 |
0.000 |
| z |
0.000 |
0.000 |
4.498 |
<r2> (average value of r
2) Å
2
| <r2> |
84.528 |
| (<r2>)1/2 |
9.194 |
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -477.723445 |
| Energy at 298.15K | -477.729684 |
| HF Energy | -477.723445 |
| Nuclear repulsion energy | 106.023398 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3070 |
3052 |
21.13 |
|
|
|
| 2 |
A |
3047 |
3029 |
18.27 |
|
|
|
| 3 |
A |
3036 |
3019 |
7.97 |
|
|
|
| 4 |
A |
2991 |
2974 |
16.65 |
|
|
|
| 5 |
A |
2966 |
2948 |
26.06 |
|
|
|
| 6 |
A |
2601 |
2586 |
8.38 |
|
|
|
| 7 |
A |
1421 |
1413 |
2.24 |
|
|
|
| 8 |
A |
1410 |
1402 |
9.19 |
|
|
|
| 9 |
A |
1401 |
1393 |
1.03 |
|
|
|
| 10 |
A |
1339 |
1331 |
2.16 |
|
|
|
| 11 |
A |
1239 |
1232 |
19.96 |
|
|
|
| 12 |
A |
1221 |
1214 |
6.40 |
|
|
|
| 13 |
A |
1076 |
1069 |
4.58 |
|
|
|
| 14 |
A |
1032 |
1026 |
1.97 |
|
|
|
| 15 |
A |
960 |
955 |
7.64 |
|
|
|
| 16 |
A |
839 |
834 |
5.97 |
|
|
|
| 17 |
A |
713 |
709 |
2.40 |
|
|
|
| 18 |
A |
639 |
635 |
3.68 |
|
|
|
| 19 |
A |
319 |
318 |
2.26 |
|
|
|
| 20 |
A |
262 |
260 |
2.92 |
|
|
|
| 21 |
A |
220 |
219 |
14.66 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15902.0 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 15809.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.652 |
-0.351 |
-0.055 |
| C2 |
0.505 |
0.644 |
0.093 |
| S3 |
-1.176 |
-0.100 |
-0.082 |
| H4 |
2.630 |
0.173 |
0.005 |
| H5 |
1.629 |
-1.113 |
0.751 |
| H6 |
1.598 |
-0.882 |
-1.026 |
| H7 |
0.557 |
1.187 |
1.059 |
| H8 |
0.535 |
1.409 |
-0.710 |
| H9 |
-1.081 |
-0.938 |
0.996 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5262 | 2.8393 | 1.1110 | 1.1094 | 1.1079 | 2.1927 | 2.1851 | 2.9866 |
C2 | 1.5262 | | 1.8458 | 2.1790 | 2.1870 | 2.1856 | 1.1089 | 1.1096 | 2.4148 | S3 | 2.8393 | 1.8458 | | 3.8167 | 3.0958 | 3.0325 | 2.4406 | 2.3654 | 1.3684 | H4 | 1.1110 | 2.1790 | 3.8167 | | 1.7925 | 1.8006 | 2.5377 | 2.5359 | 3.9985 | H5 | 1.1094 | 2.1870 | 3.0958 | 1.7925 | | 1.7917 | 2.5568 | 3.1129 | 2.7263 | H6 | 1.1079 | 2.1856 | 3.0325 | 1.8006 | 1.7917 | | 3.1165 | 2.5453 | 3.3564 | H7 | 2.1927 | 1.1089 | 2.4406 | 2.5377 | 2.5568 | 3.1165 | | 1.7824 | 2.6837 | H8 | 2.1851 | 1.1096 | 2.3654 | 2.5359 | 3.1129 | 2.5453 | 1.7824 | | 3.3207 | H9 | 2.9866 | 2.4148 | 1.3684 | 3.9985 | 2.7263 | 3.3564 | 2.6837 | 3.3207 | |
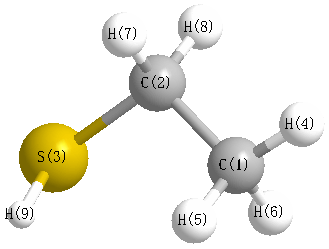 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
114.378 |
|
C1 |
C2 |
H7 |
111.657 |
| C1 |
C2 |
H8 |
111.012 |
|
C2 |
C1 |
H4 |
110.445 |
| C2 |
C1 |
H5 |
111.171 |
|
C2 |
C1 |
H6 |
111.144 |
| C2 |
S3 |
H9 |
96.269 |
|
S3 |
C2 |
H7 |
108.811 |
| S3 |
C2 |
H8 |
103.514 |
|
H4 |
C1 |
H5 |
107.661 |
| H4 |
C1 |
H6 |
108.483 |
|
H5 |
C1 |
H6 |
107.808 |
| H7 |
C2 |
H8 |
106.925 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.047 |
|
|
|
| 2 |
C |
-0.178 |
|
|
|
| 3 |
S |
-0.119 |
|
|
|
| 4 |
H |
0.037 |
|
|
|
| 5 |
H |
0.038 |
|
|
|
| 6 |
H |
0.048 |
|
|
|
| 7 |
H |
0.066 |
|
|
|
| 8 |
H |
0.072 |
|
|
|
| 9 |
H |
0.083 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.418 |
0.069 |
0.721 |
1.592 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.750 |
1.268 |
-0.724 |
| y |
1.268 |
-26.937 |
-1.615 |
| z |
-0.724 |
-1.615 |
-26.772 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.896 |
1.268 |
-0.724 |
| y |
1.268 |
0.824 |
-1.615 |
| z |
-0.724 |
-1.615 |
1.071 |
|
| Polar |
|---|
| 3z2-r2 | 2.143 |
|---|
| x2-y2 | -1.813 |
|---|
| xy | 1.268 |
|---|
| xz | -0.724 |
|---|
| yz | -1.615 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.345 |
0.484 |
0.034 |
| y |
0.484 |
5.437 |
-0.453 |
| z |
0.034 |
-0.453 |
5.197 |
<r2> (average value of r
2) Å
2
| <r2> |
85.197 |
| (<r2>)1/2 |
9.230 |
