Jump to
S1C2
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -169.465756 |
| Energy at 298.15K | |
| HF Energy | -168.946131 |
| Nuclear repulsion energy | 70.665776 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3757 |
3580 |
|
|
|
|
| 2 |
A' |
3603 |
3434 |
|
|
|
|
| 3 |
A' |
2960 |
2821 |
|
|
|
|
| 4 |
A' |
1829 |
1743 |
|
|
|
|
| 5 |
A' |
1601 |
1526 |
|
|
|
|
| 6 |
A' |
1419 |
1352 |
|
|
|
|
| 7 |
A' |
1276 |
1216 |
|
|
|
|
| 8 |
A' |
1047 |
998 |
|
|
|
|
| 9 |
A' |
560 |
533 |
|
|
|
|
| 10 |
A" |
1044 |
995 |
|
|
|
|
| 11 |
A" |
639 |
609 |
|
|
|
|
| 12 |
A" |
288i |
274i |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9722.6 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 9266.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.421 |
0.000 |
| O2 |
1.207 |
0.241 |
0.000 |
| N3 |
-0.944 |
-0.571 |
0.000 |
| H4 |
-0.472 |
1.438 |
0.000 |
| H5 |
-0.641 |
-1.540 |
0.000 |
| H6 |
-1.934 |
-0.357 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2201 | 1.3698 | 1.1204 | 2.0636 | 2.0844 |
O2 | 1.2201 | | 2.2992 | 2.0610 | 2.5667 | 3.1969 | N3 | 1.3698 | 2.2992 | | 2.0636 | 1.0154 | 1.0125 | H4 | 1.1204 | 2.0610 | 2.0636 | | 2.9826 | 2.3147 | H5 | 2.0636 | 2.5667 | 1.0154 | 2.9826 | | 1.7527 | H6 | 2.0844 | 3.1969 | 1.0125 | 2.3147 | 1.7527 | |
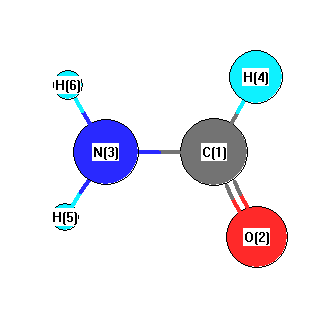 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.056 |
|
C1 |
N3 |
H6 |
121.351 |
| O2 |
C1 |
N3 |
125.090 |
|
O2 |
C1 |
H4 |
123.371 |
| N3 |
C1 |
H4 |
111.539 |
|
H5 |
N3 |
H6 |
119.594 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -169.466139 |
| Energy at 298.15K | -169.470011 |
| HF Energy | -168.945948 |
| Nuclear repulsion energy | 70.580648 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3710 |
3536 |
|
|
|
|
| 2 |
A |
3571 |
3403 |
|
|
|
|
| 3 |
A |
2965 |
2826 |
|
|
|
|
| 4 |
A |
1829 |
1743 |
|
|
|
|
| 5 |
A |
1609 |
1534 |
|
|
|
|
| 6 |
A |
1420 |
1353 |
|
|
|
|
| 7 |
A |
1275 |
1215 |
|
|
|
|
| 8 |
A |
1075 |
1024 |
|
|
|
|
| 9 |
A |
1028 |
980 |
|
|
|
|
| 10 |
A |
620 |
591 |
|
|
|
|
| 11 |
A |
559 |
533 |
|
|
|
|
| 12 |
A |
382 |
364 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10020.6 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 9550.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.164 |
0.390 |
0.001 |
| O2 |
1.203 |
-0.248 |
0.010 |
| N3 |
-1.100 |
-0.160 |
-0.068 |
| H4 |
0.129 |
1.509 |
-0.001 |
| H5 |
-1.159 |
-1.156 |
0.137 |
| H6 |
-1.881 |
0.408 |
0.247 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2188 | 1.3800 | 1.1200 | 2.0392 | 2.0596 |
O2 | 1.2188 | | 2.3057 | 2.0589 | 2.5337 | 3.1615 | N3 | 1.3800 | 2.3057 | | 2.0740 | 1.0185 | 1.0161 | H4 | 1.1200 | 2.0589 | 2.0740 | | 2.9635 | 2.3051 | H5 | 2.0392 | 2.5337 | 1.0185 | 2.9635 | | 1.7265 | H6 | 2.0596 | 3.1615 | 1.0161 | 2.3051 | 1.7265 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
115.638 |
|
C1 |
N3 |
H6 |
117.733 |
| O2 |
C1 |
N3 |
124.933 |
|
O2 |
C1 |
H4 |
123.309 |
| N3 |
C1 |
H4 |
111.697 |
|
H5 |
N3 |
H6 |
116.109 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
