Jump to
S1C2
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -169.856485 |
| Energy at 298.15K | |
| HF Energy | -169.856485 |
| Nuclear repulsion energy | 70.316661 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3582 |
3588 |
19.45 |
|
|
|
| 2 |
A' |
3434 |
3440 |
11.24 |
|
|
|
| 3 |
A' |
2795 |
2799 |
141.02 |
|
|
|
| 4 |
A' |
1755 |
1758 |
307.88 |
|
|
|
| 5 |
A' |
1529 |
1531 |
50.52 |
|
|
|
| 6 |
A' |
1363 |
1366 |
4.80 |
|
|
|
| 7 |
A' |
1222 |
1224 |
76.28 |
|
|
|
| 8 |
A' |
1009 |
1011 |
2.32 |
|
|
|
| 9 |
A' |
545 |
546 |
9.45 |
|
|
|
| 10 |
A" |
994 |
996 |
1.25 |
|
|
|
| 11 |
A" |
640 |
641 |
10.94 |
|
|
|
| 12 |
A" |
40i |
40i |
177.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9414.6 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 9429.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.423 |
0.000 |
| O2 |
1.213 |
0.242 |
0.000 |
| N3 |
-0.947 |
-0.573 |
0.000 |
| H4 |
-0.482 |
1.444 |
0.000 |
| H5 |
-0.646 |
-1.550 |
0.000 |
| H6 |
-1.945 |
-0.360 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2263 | 1.3745 | 1.1287 | 2.0765 | 2.0969 |
O2 | 1.2263 | | 2.3085 | 2.0776 | 2.5818 | 3.2148 | N3 | 1.3745 | 2.3085 | | 2.0699 | 1.0228 | 1.0204 | H4 | 1.1287 | 2.0776 | 2.0699 | | 2.9988 | 2.3231 | H5 | 2.0765 | 2.5818 | 1.0228 | 2.9988 | | 1.7620 | H6 | 2.0969 | 3.2148 | 1.0204 | 2.3231 | 1.7620 | |
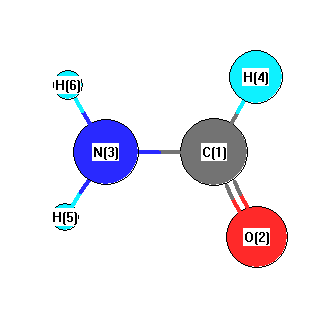 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.306 |
|
C1 |
N3 |
H6 |
121.522 |
| O2 |
C1 |
N3 |
125.052 |
|
O2 |
C1 |
H4 |
123.764 |
| N3 |
C1 |
H4 |
111.184 |
|
H5 |
N3 |
H6 |
119.172 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.128 |
|
|
|
| 2 |
O |
-0.222 |
|
|
|
| 3 |
N |
-0.040 |
|
|
|
| 4 |
H |
-0.035 |
|
|
|
| 5 |
H |
0.087 |
|
|
|
| 6 |
H |
0.083 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.525 |
-0.882 |
0.000 |
3.634 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.607 |
0.139 |
0.000 |
| y |
0.139 |
-15.044 |
0.000 |
| z |
0.000 |
0.000 |
-18.330 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.921 |
0.139 |
0.000 |
| y |
0.139 |
2.925 |
0.000 |
| z |
0.000 |
0.000 |
-2.004 |
|
| Polar |
|---|
| 3z2-r2 | -4.008 |
|---|
| x2-y2 | -2.564 |
|---|
| xy | 0.139 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.448 |
0.287 |
0.000 |
| y |
0.287 |
3.730 |
0.000 |
| z |
0.000 |
0.000 |
1.599 |
<r2> (average value of r
2) Å
2
| <r2> |
41.551 |
| (<r2>)1/2 |
6.446 |
Jump to
S1C1
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -169.856489 |
| Energy at 298.15K | -169.859904 |
| HF Energy | -169.856489 |
| Nuclear repulsion energy | 70.313644 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3581 |
3586 |
19.22 |
|
|
|
| 2 |
A |
3434 |
3439 |
11.07 |
|
|
|
| 3 |
A |
2798 |
2802 |
140.44 |
|
|
|
| 4 |
A |
1754 |
1757 |
306.63 |
|
|
|
| 5 |
A |
1529 |
1531 |
50.13 |
|
|
|
| 6 |
A |
1363 |
1365 |
4.71 |
|
|
|
| 7 |
A |
1223 |
1225 |
76.02 |
|
|
|
| 8 |
A |
1011 |
1013 |
2.50 |
|
|
|
| 9 |
A |
993 |
995 |
1.17 |
|
|
|
| 10 |
A |
639 |
640 |
11.95 |
|
|
|
| 11 |
A |
546 |
546 |
9.39 |
|
|
|
| 12 |
A |
75 |
75 |
178.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9472.3 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 9487.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.164 |
0.391 |
0.001 |
| O2 |
1.212 |
-0.247 |
0.002 |
| N3 |
-1.095 |
-0.161 |
-0.014 |
| H4 |
0.116 |
1.518 |
0.000 |
| H5 |
-1.194 |
-1.179 |
0.028 |
| H6 |
-1.931 |
0.420 |
0.048 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2265 | 1.3747 | 1.1284 | 2.0749 | 2.0958 |
O2 | 1.2265 | | 2.3086 | 2.0772 | 2.5797 | 3.2134 | N3 | 1.3747 | 2.3086 | | 2.0706 | 1.0229 | 1.0205 | H4 | 1.1284 | 2.0772 | 2.0706 | | 2.9979 | 2.3234 | H5 | 2.0749 | 2.5797 | 1.0229 | 2.9979 | | 1.7612 | H6 | 2.0958 | 3.2134 | 1.0205 | 2.3234 | 1.7612 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.135 |
|
C1 |
N3 |
H6 |
121.387 |
| O2 |
C1 |
N3 |
125.027 |
|
O2 |
C1 |
H4 |
123.724 |
| N3 |
C1 |
H4 |
111.247 |
|
H5 |
N3 |
H6 |
119.066 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.128 |
|
|
|
| 2 |
O |
-0.222 |
|
|
|
| 3 |
N |
-0.040 |
|
|
|
| 4 |
H |
-0.035 |
|
|
|
| 5 |
H |
0.087 |
|
|
|
| 6 |
H |
0.083 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.583 |
0.554 |
0.139 |
3.628 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.144 |
1.011 |
-0.258 |
| y |
1.011 |
-15.529 |
-0.026 |
| z |
-0.258 |
-0.026 |
-18.325 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.217 |
1.011 |
-0.258 |
| y |
1.011 |
2.206 |
-0.026 |
| z |
-0.258 |
-0.026 |
-1.988 |
|
| Polar |
|---|
| 3z2-r2 | -3.977 |
|---|
| x2-y2 | -1.615 |
|---|
| xy | 1.011 |
|---|
| xz | -0.258 |
|---|
| yz | -0.026 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.544 |
-0.056 |
-0.014 |
| y |
-0.056 |
3.632 |
-0.003 |
| z |
-0.014 |
-0.003 |
1.602 |
<r2> (average value of r
2) Å
2
| <r2> |
41.548 |
| (<r2>)1/2 |
6.446 |
