Jump to
S1C2
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -169.711279 |
| Energy at 298.15K | -169.714698 |
| HF Energy | -169.711279 |
| Nuclear repulsion energy | 70.526078 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3620 |
3599 |
26.91 |
|
|
|
| 2 |
A' |
3468 |
3448 |
15.04 |
|
|
|
| 3 |
A' |
2819 |
2803 |
134.92 |
|
|
|
| 4 |
A' |
1786 |
1776 |
317.40 |
|
|
|
| 5 |
A' |
1525 |
1517 |
58.52 |
|
|
|
| 6 |
A' |
1359 |
1351 |
3.82 |
|
|
|
| 7 |
A' |
1238 |
1231 |
68.34 |
|
|
|
| 8 |
A' |
1013 |
1007 |
0.87 |
|
|
|
| 9 |
A' |
543 |
540 |
9.65 |
|
|
|
| 10 |
A" |
995 |
989 |
1.60 |
|
|
|
| 11 |
A" |
646 |
643 |
11.59 |
|
|
|
| 12 |
A" |
75 |
74 |
183.38 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9544.0 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 9488.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.421 |
0.000 |
| O2 |
1.210 |
0.237 |
0.000 |
| N3 |
-0.945 |
-0.568 |
0.000 |
| H4 |
-0.479 |
1.444 |
0.000 |
| H5 |
-0.643 |
-1.544 |
0.000 |
| H6 |
-1.941 |
-0.354 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2236 | 1.3677 | 1.1290 | 2.0679 | 2.0904 |
O2 | 1.2236 | | 2.2999 | 2.0753 | 2.5703 | 3.2060 | N3 | 1.3677 | 2.2999 | | 2.0646 | 1.0219 | 1.0193 | H4 | 1.1290 | 2.0753 | 2.0646 | | 2.9922 | 2.3172 | H5 | 2.0679 | 2.5703 | 1.0219 | 2.9922 | | 1.7613 | H6 | 2.0904 | 3.2060 | 1.0193 | 2.3172 | 1.7613 | |
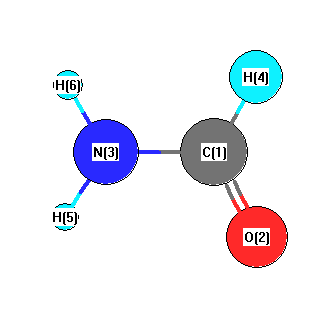 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.142 |
|
C1 |
N3 |
H6 |
121.578 |
| O2 |
C1 |
N3 |
125.041 |
|
O2 |
C1 |
H4 |
123.754 |
| N3 |
C1 |
H4 |
111.205 |
|
H5 |
N3 |
H6 |
119.280 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.125 |
|
|
|
| 2 |
O |
-0.232 |
|
|
|
| 3 |
N |
-0.080 |
|
|
|
| 4 |
H |
-0.012 |
|
|
|
| 5 |
H |
0.101 |
|
|
|
| 6 |
H |
0.098 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.563 |
-0.892 |
0.000 |
3.673 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.460 |
0.130 |
0.000 |
| y |
0.130 |
-14.914 |
0.000 |
| z |
0.000 |
0.000 |
-18.350 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.828 |
0.130 |
0.000 |
| y |
0.130 |
2.991 |
0.000 |
| z |
0.000 |
0.000 |
-2.163 |
|
| Polar |
|---|
| 3z2-r2 | -4.327 |
|---|
| x2-y2 | -2.546 |
|---|
| xy | 0.130 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.406 |
0.278 |
0.000 |
| y |
0.278 |
3.683 |
0.000 |
| z |
0.000 |
0.000 |
1.612 |
<r2> (average value of r
2) Å
2
| <r2> |
41.292 |
| (<r2>)1/2 |
6.426 |
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -169.711278 |
| Energy at 298.15K | -169.714684 |
| HF Energy | -169.711278 |
| Nuclear repulsion energy | 70.526406 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3621 |
3600 |
26.97 |
|
|
|
| 2 |
A |
3468 |
3448 |
15.04 |
|
|
|
| 3 |
A |
2821 |
2804 |
134.65 |
|
|
|
| 4 |
A |
1785 |
1775 |
317.25 |
|
|
|
| 5 |
A |
1525 |
1516 |
58.50 |
|
|
|
| 6 |
A |
1358 |
1350 |
3.80 |
|
|
|
| 7 |
A |
1239 |
1232 |
68.13 |
|
|
|
| 8 |
A |
1013 |
1007 |
0.85 |
|
|
|
| 9 |
A |
995 |
990 |
1.61 |
|
|
|
| 10 |
A |
647 |
643 |
11.79 |
|
|
|
| 11 |
A |
543 |
540 |
9.68 |
|
|
|
| 12 |
A |
68 |
67 |
183.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9540.9 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 9485.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.162 |
0.389 |
0.000 |
| O2 |
1.208 |
-0.245 |
0.000 |
| N3 |
-1.090 |
-0.161 |
-0.000 |
| H4 |
0.113 |
1.517 |
-0.000 |
| H5 |
-1.186 |
-1.179 |
0.000 |
| H6 |
-1.928 |
0.418 |
0.001 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2239 | 1.3675 | 1.1288 | 2.0673 | 2.0902 |
O2 | 1.2239 | | 2.2998 | 2.0750 | 2.5693 | 3.2060 | N3 | 1.3675 | 2.2998 | | 2.0649 | 1.0219 | 1.0192 | H4 | 1.1288 | 2.0750 | 2.0649 | | 2.9922 | 2.3179 | H5 | 2.0673 | 2.5693 | 1.0219 | 2.9922 | | 1.7616 | H6 | 2.0902 | 3.2060 | 1.0192 | 2.3179 | 1.7616 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.101 |
|
C1 |
N3 |
H6 |
121.583 |
| O2 |
C1 |
N3 |
125.025 |
|
O2 |
C1 |
H4 |
123.717 |
| N3 |
C1 |
H4 |
111.259 |
|
H5 |
N3 |
H6 |
119.316 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.125 |
|
|
|
| 2 |
O |
-0.233 |
|
|
|
| 3 |
N |
-0.079 |
|
|
|
| 4 |
H |
-0.012 |
|
|
|
| 5 |
H |
0.101 |
|
|
|
| 6 |
H |
0.098 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.634 |
0.545 |
0.002 |
3.674 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.994 |
0.994 |
-0.004 |
| y |
0.994 |
-15.379 |
-0.000 |
| z |
-0.004 |
-0.000 |
-18.350 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.129 |
0.994 |
-0.004 |
| y |
0.994 |
2.293 |
-0.000 |
| z |
-0.004 |
-0.000 |
-2.163 |
|
| Polar |
|---|
| 3z2-r2 | -4.327 |
|---|
| x2-y2 | -1.615 |
|---|
| xy | 0.994 |
|---|
| xz | -0.004 |
|---|
| yz | -0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.496 |
-0.060 |
-0.000 |
| y |
-0.060 |
3.593 |
0.000 |
| z |
-0.000 |
0.000 |
1.612 |
<r2> (average value of r
2) Å
2
| <r2> |
41.289 |
| (<r2>)1/2 |
6.426 |
