return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHBrCH3 (i-propyl bromide)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-2690.700932
Energy at 298.15K-2690.711452
HF Energy-2690.079958
Nuclear repulsion energy249.636125
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3169 3040 22.91      
2 A' 3145 3017 36.66      
3 A' 3122 2996 1.29      
4 A' 3059 2935 27.53      
5 A' 1503 1442 5.23      
6 A' 1491 1431 7.27      
7 A' 1424 1366 2.61      
8 A' 1271 1219 46.43      
9 A' 1191 1143 20.41      
10 A' 1067 1024 6.89      
11 A' 916 879 5.48      
12 A' 563 540 15.05      
13 A' 405 389 1.23      
14 A' 307 294 1.10      
15 A' 273 262 0.53      
16 A" 3165 3036 11.55      
17 A" 3139 3011 2.10      
18 A" 3055 2931 13.97      
19 A" 1484 1424 0.51      
20 A" 1480 1420 1.55      
21 A" 1416 1359 6.47      
22 A" 1364 1308 0.85      
23 A" 1166 1119 0.73      
24 A" 964 925 0.05      
25 A" 939 901 0.85      
26 A" 291 279 0.59      
27 A" 254 244 0.04      

Unscaled Zero Point Vibrational Energy (zpe) 20810.8 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 19965.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.26553 0.09578 0.07548

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.910 -0.432 0.000
Br2 -0.676 0.758 0.000
H3 1.763 0.268 0.000
C4 0.910 -1.272 1.275
C5 0.910 -1.272 -1.275
H6 0.023 -1.929 1.308
H7 1.817 -1.906 1.294
H8 0.909 -0.635 2.176
H9 0.023 -1.929 -1.308
H10 1.817 -1.906 -1.294
H11 0.909 -0.635 -2.176

Atom - Atom Distances (Å)
  C1 Br2 H3 C4 C5 H6 H7 H8 H9 H10 H11
C11.98221.10271.52731.52732.17722.16142.18542.17722.16142.1854
Br21.98222.48682.87412.87413.06873.87113.03083.06873.87113.0308
H31.10272.48682.17352.17353.09252.53082.50593.09252.53082.5059
C41.52732.87412.17352.55071.10451.10701.10272.80952.79743.5094
C51.52732.87412.17352.55072.80952.79743.50941.10451.10701.1027
H62.17723.06873.09251.10452.80951.79431.79182.61633.16063.8204
H72.16143.87112.53081.10702.79741.79431.79403.16062.58753.8051
H82.18543.03082.50591.10273.50941.79181.79403.82043.80514.3517
H92.17723.06873.09252.80951.10452.61633.16063.82041.79431.7918
H102.16143.87112.53082.79741.10703.16062.58753.80511.79431.7940
H112.18543.03082.50593.50941.10273.82043.80514.35171.79181.7940

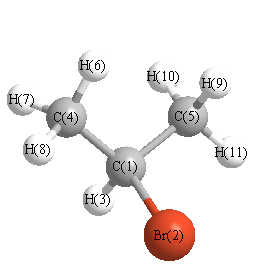
picture of i-propyl bromide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H6 110.614 C1 C4 H7 109.227
C1 C4 H8 111.370 C1 C5 H9 110.614
C1 C5 H10 109.227 C1 C5 H11 111.370
Br2 C1 H3 103.753 Br2 C1 C4 109.278
Br2 C1 C5 109.278 H3 C1 C4 110.428
H3 C1 C5 110.428 C4 C1 C5 113.238
H6 C4 H7 108.458 H6 C4 H8 108.543
H7 C4 H8 108.558 H9 C5 H10 108.458
H9 C5 H11 108.543 H10 C5 H11 108.558
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability