return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHClCH3 (Propane, 2-chloro-)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-577.828665
Energy at 298.15K-577.836430
HF Energy-577.198853
Nuclear repulsion energy164.981356
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3167 3039 25.30      
2 A' 3149 3021 39.84      
3 A' 3114 2987 5.87      
4 A' 3063 2938 20.48      
5 A' 1504 1443 5.73      
6 A' 1491 1431 6.29      
7 A' 1425 1367 3.68      
8 A' 1301 1248 33.81      
9 A' 1196 1147 12.90      
10 A' 1088 1044 10.53      
11 A' 922 885 8.32      
12 A' 649 623 23.57      
13 A' 425 408 1.69      
14 A' 341 327 0.99      
15 A' 282 271 0.10      
16 A" 3163 3035 12.89      
17 A" 3143 3015 1.75      
18 A" 3059 2935 14.79      
19 A" 1485 1424 0.39      
20 A" 1481 1421 1.09      
21 A" 1418 1360 7.81      
22 A" 1367 1311 1.28      
23 A" 1169 1122 1.90      
24 A" 963 924 0.01      
25 A" 940 902 0.70      
26 A" 328 315 1.74      
27 A" 251 241 0.03      

Unscaled Zero Point Vibrational Energy (zpe) 20939.9 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 20089.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.26636 0.15018 0.10562

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Cl1 -0.892 1.022 0.000
C2 0.571 -0.064 0.000
C3 0.571 -0.906 1.274
C4 0.571 -0.906 -1.274
H5 1.432 0.625 0.000
H6 1.478 -1.538 1.299
H7 1.478 -1.538 -1.299
H8 0.563 -0.266 2.173
H9 0.563 -0.266 -2.173
H10 -0.317 -1.563 1.303
H11 -0.317 -1.563 -1.303

Atom - Atom Distances (Å)
  Cl1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11
Cl11.82152.73462.73462.35753.72243.72242.91472.91472.95092.9509
C21.82151.52721.52721.10342.16412.16412.18232.18232.17492.1749
C32.73461.52722.54892.17021.10632.80121.10293.50631.10452.8037
C42.73461.52722.54892.17022.80121.10633.50631.10292.80371.1045
H52.35751.10342.17022.17022.52372.52372.50442.50443.08913.0891
H63.72242.16411.10632.80122.52372.59841.79423.80941.79503.1610
H73.72242.16412.80121.10632.52372.59843.80941.79423.16101.7950
H82.91472.18231.10293.50632.50441.79423.80944.34591.79223.8124
H92.91472.18233.50631.10292.50443.80941.79424.34593.81241.7922
H102.95092.17491.10452.80373.08911.79503.16101.79223.81242.6054
H112.95092.17492.80371.10453.08913.16101.79503.81241.79222.6054

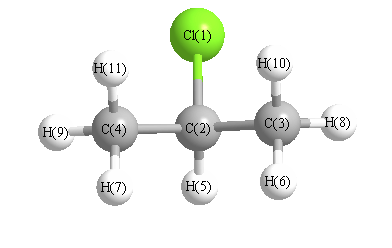
picture of Propane, 2-chloro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
Cl1 C2 C3 109.180 Cl1 C2 C4 109.180
Cl1 C3 H5 56.042 C2 C3 H6 109.487
C2 C3 H8 111.117 C2 C3 H10 110.434
C2 C4 H7 109.487 C2 C4 H9 111.117
C2 C4 H11 110.434 C3 C2 C4 113.127
C3 C2 H5 110.134 C4 C2 H5 110.134
H6 C3 H8 108.614 H6 C3 H10 108.567
H7 C4 H9 108.614 H7 C4 H11 108.567
H8 C3 H10 108.563 H9 C4 H11 108.563
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability