return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C(CH3)3SH (2-Propanethiol, 2-methyl-)

using model chemistry: QCISD(T)/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD(T)/cc-pVDZ
 hartrees
Energy at 0K-555.660036
Energy at 298.15K-555.670820
HF Energy-554.838291
Nuclear repulsion energy242.834646
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3137 2989        
2 A' 3122 2975        
3 A' 3111 2966        
4 A' 3037 2894        
5 A' 3030 2888        
6 A' 2700 2574        
7 A' 1500 1430        
8 A' 1487 1417        
9 A' 1473 1404        
10 A' 1417 1351        
11 A' 1391 1326        
12 A' 1267 1208        
13 A' 1200 1144        
14 A' 1052 1003        
15 A' 949 905        
16 A' 876 835        
17 A' 841 802        
18 A' 593 565        
19 A' 391 373        
20 A' 366 349        
21 A' 304 289        
22 A' 286 272        
23 A" 3135 2988        
24 A" 3130 2983        
25 A" 3107 2961        
26 A" 3026 2884        
27 A" 1488 1418        
28 A" 1470 1401        
29 A" 1464 1395        
30 A" 1392 1326        
31 A" 1254 1195        
32 A" 1043 994        
33 A" 955 911        
34 A" 941 897        
35 A" 397 379        
36 A" 302 288        
37 A" 286 273        
38 A" 254 242        
39 A" 201 191        

Unscaled Zero Point Vibrational Energy (zpe) 28686.6 cm-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 27341.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/cc-pVDZ
ABC
0.14783 0.09816 0.09726

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.350 -0.006 0.000
S2 -1.514 0.088 0.000
C3 0.840 1.454 0.000
C4 0.840 -0.733 1.265
C5 0.840 -0.733 -1.265
H6 -1.716 -1.252 0.000
H7 1.947 1.472 0.000
H8 0.483 1.993 -0.898
H9 0.483 1.993 0.898
H10 1.949 -0.757 1.281
H11 1.949 -0.757 -1.281
H12 0.484 -0.220 2.177
H13 0.474 -1.778 1.289
H14 0.484 -0.220 -2.177
H15 0.474 -1.778 -1.289

Atom - Atom Distances (Å)
  C1 S2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.86581.54061.53871.53872.41232.17672.19622.19622.18182.18182.19172.19452.19172.1945
S21.86582.72112.79522.79521.35573.72742.90252.90253.78723.78722.97103.01632.97103.0163
C31.54062.72112.52652.52653.72241.10781.10641.10642.78562.78562.76953.49872.76953.4987
C41.53872.79522.52652.52932.89852.77273.49822.77381.10932.77641.10531.10703.49782.7834
C51.53872.79522.52652.52932.89852.77272.77383.49822.77641.10933.49782.78341.10531.1070
H62.41231.35573.72242.89852.89854.56524.02224.02223.91353.91353.26282.59573.26282.5957
H72.17673.72741.10782.77272.77274.56521.79481.79482.57072.57073.12153.79363.12153.7936
H82.19622.90251.10643.49822.77384.02221.79481.79623.80253.13983.78914.35962.55663.7914
H92.19622.90251.10642.77383.49824.02221.79481.79623.13983.80252.55663.79143.78914.3596
H102.18183.78722.78561.10932.77643.91352.57073.80253.13982.56111.79901.79293.79323.1335
H112.18183.78722.78562.77641.10933.91352.57073.13983.80252.56113.79323.13351.79901.7929
H122.19172.97102.76951.10533.49783.26283.12153.78912.55661.79903.79321.79284.35423.8002
H132.19453.01633.49871.10702.78342.59573.79364.35963.79141.79293.13351.79283.80022.5785
H142.19172.97102.76953.49781.10533.26283.12152.55663.78913.79321.79904.35423.80021.7928
H152.19453.01633.49872.78341.10702.59573.79363.79144.35963.13351.79293.80022.57851.7928

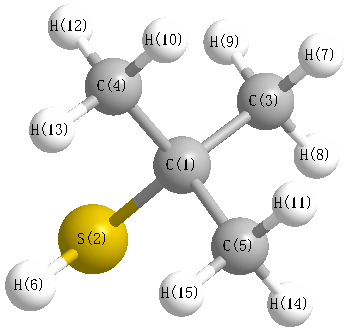
picture of 2-Propanethiol, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S2 H6 95.674 C1 C3 H7 109.465
C1 C3 H8 111.075 C1 C3 H9 111.075
C1 C4 H10 109.906 C1 C4 H12 110.916
C1 C4 H13 111.033 C1 C5 H11 109.906
C1 C5 H14 110.916 C1 C5 H15 111.033
S2 C1 C3 105.639 S2 C1 C4 110.003
S2 C1 C5 110.003 C3 C1 C4 110.266
C3 C1 C5 110.266 C4 C1 C5 110.553
H7 C3 H8 108.307 H7 C3 H9 108.307
H8 C3 H9 108.526 H10 C4 H12 108.649
H10 C4 H13 107.988 H11 C5 H14 108.649
H11 C5 H15 107.988 H12 C4 H13 108.260
H14 C5 H15 108.260
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability