return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C(CH3)3SH (2-Propanethiol, 2-methyl-)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-555.635872
Energy at 298.15K-555.646711
HF Energy-554.838690
Nuclear repulsion energy243.185508
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3152 3024 26.78      
2 A' 3138 3010 30.85      
3 A' 3128 3001 39.51      
4 A' 3055 2931 21.65      
5 A' 3048 2924 32.23      
6 A' 2722 2611 12.82      
7 A' 1511 1450 6.06      
8 A' 1498 1437 7.93      
9 A' 1484 1424 0.07      
10 A' 1432 1373 0.46      
11 A' 1406 1349 5.79      
12 A' 1281 1229 0.24      
13 A' 1214 1165 32.51      
14 A' 1063 1020 1.55      
15 A' 958 919 0.20      
16 A' 886 850 3.68      
17 A' 852 817 1.37      
18 A' 603 578 3.69      
19 A' 396 380 1.13      
20 A' 370 355 0.09      
21 A' 305 293 0.12      
22 A' 288 277 0.75      
23 A" 3150 3022 27.73      
24 A" 3145 3017 4.64      
25 A" 3123 2996 0.89      
26 A" 3044 2921 19.33      
27 A" 1499 1438 6.12      
28 A" 1480 1420 0.14      
29 A" 1474 1415 0.00      
30 A" 1407 1349 5.65      
31 A" 1267 1215 3.01      
32 A" 1052 1010 0.08      
33 A" 965 926 0.01      
34 A" 951 912 0.11      
35 A" 402 386 0.63      
36 A" 306 293 1.34      
37 A" 288 276 0.03      
38 A" 255 245 6.00      
39 A" 201 193 10.35      

Unscaled Zero Point Vibrational Energy (zpe) 28897.7 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 27724.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.14822 0.09846 0.09756

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.348 -0.006 0.000
S2 -1.511 0.088 0.000
C3 0.838 1.452 0.000
C4 0.838 -0.732 1.263
C5 0.838 -0.732 -1.263
H6 -1.720 -1.249 0.000
H7 1.944 1.470 0.000
H8 0.483 1.992 -0.897
H9 0.483 1.992 0.897
H10 1.946 -0.757 1.278
H11 1.946 -0.757 -1.278
H12 0.485 -0.221 2.175
H13 0.474 -1.776 1.289
H14 0.485 -0.221 -2.175
H15 0.474 -1.776 -1.289

Atom - Atom Distances (Å)
  C1 S2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.86101.53881.53711.53712.41212.17482.19402.19402.17992.17992.18992.19252.18992.1925
S21.86102.71672.79072.79071.35353.72162.89912.89913.78133.78132.96833.01262.96833.0126
C31.53882.71672.52292.52293.72031.10611.10491.10492.78202.78202.76663.49432.76663.4943
C41.53712.79072.52292.52582.89942.76913.49392.77101.10762.77221.10381.10553.49372.7806
C51.53712.79072.52292.52582.89942.76912.77103.49392.77221.10763.49372.78061.10381.1055
H62.41211.35353.72032.89942.89944.56294.01994.01993.91313.91313.26342.59843.26342.5984
H72.17483.72161.10612.76912.76914.56291.79171.79172.56782.56783.11773.78903.11773.7890
H82.19402.89911.10493.49392.77104.01991.79171.79313.79773.13653.78534.35502.55523.7875
H92.19402.89911.10492.77103.49394.01991.79171.79313.13653.79772.55523.78753.78534.3550
H102.17993.78132.78201.10762.77223.91312.56783.79773.13652.55691.79581.78973.78793.1294
H112.17993.78132.78202.77221.10763.91312.56783.13653.79772.55693.78793.12941.79581.7897
H122.18992.96832.76661.10383.49373.26343.11773.78532.55521.79583.78791.78984.35013.7967
H132.19253.01263.49431.10552.78062.59843.78904.35503.78751.78973.12941.78983.79672.5773
H142.18992.96832.76663.49371.10383.26343.11772.55523.78533.78791.79584.35013.79671.7898
H152.19253.01263.49432.78061.10552.59843.78903.78754.35503.12941.78973.79672.57731.7898

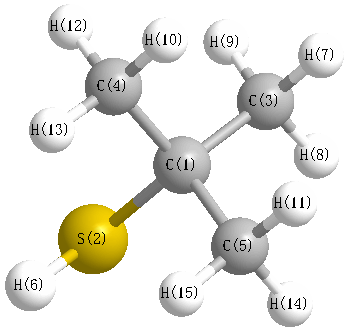
picture of 2-Propanethiol, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S2 H6 95.957 C1 C3 H7 109.542
C1 C3 H8 111.118 C1 C3 H9 111.118
C1 C4 H10 109.961 C1 C4 H12 110.976
C1 C4 H13 111.078 C1 C5 H11 109.961
C1 C5 H14 110.976 C1 C5 H15 111.078
S2 C1 C3 105.691 S2 C1 C4 110.057
S2 C1 C5 110.057 C3 C1 C4 110.219
C3 C1 C5 110.219 C4 C1 C5 110.494
H7 C3 H8 108.254 H7 C3 H9 108.254
H8 C3 H9 108.464 H10 C4 H12 108.589
H10 C4 H13 107.932 H11 C5 H14 108.589
H11 C5 H15 107.932 H12 C4 H13 108.210
H14 C5 H15 108.210
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability