return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C(CH3)3SH (2-Propanethiol, 2-methyl-)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-555.631143
Energy at 298.15K-555.641995
HF Energy-554.838799
Nuclear repulsion energy243.316183
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3164 3027 25.46      
2 A' 3150 3013 28.70      
3 A' 3140 3004 38.17      
4 A' 3064 2932 20.19      
5 A' 3058 2925 30.27      
6 A' 2736 2617 11.61      
7 A' 1514 1448 6.41      
8 A' 1500 1435 8.12      
9 A' 1486 1422 0.08      
10 A' 1435 1373 0.54      
11 A' 1409 1348 6.19      
12 A' 1284 1228 0.23      
13 A' 1220 1167 31.42      
14 A' 1065 1019 1.59      
15 A' 961 919 0.22      
16 A' 890 852 3.76      
17 A' 857 819 1.15      
18 A' 609 582 3.07      
19 A' 397 380 1.10      
20 A' 372 356 0.08      
21 A' 305 292 0.14      
22 A' 289 277 0.72      
23 A" 3162 3025 25.64      
24 A" 3157 3020 4.60      
25 A" 3135 3000 0.78      
26 A" 3054 2922 18.34      
27 A" 1501 1436 6.23      
28 A" 1483 1418 0.16      
29 A" 1477 1413 0.00      
30 A" 1410 1349 6.14      
31 A" 1270 1215 3.02      
32 A" 1055 1009 0.09      
33 A" 967 925 0.01      
34 A" 953 912 0.13      
35 A" 403 386 0.60      
36 A" 307 293 1.29      
37 A" 288 275 0.02      
38 A" 255 244 6.21      
39 A" 200 191 10.49      

Unscaled Zero Point Vibrational Energy (zpe) 28988.4 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 27733.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.14832 0.09858 0.09768

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.346 -0.006 0.000
S2 -1.510 0.088 0.000
C3 0.838 1.452 0.000
C4 0.838 -0.732 1.262
C5 0.838 -0.732 -1.262
H6 -1.719 -1.248 0.000
H7 1.943 1.470 0.000
H8 0.483 1.991 -0.896
H9 0.483 1.991 0.896
H10 1.945 -0.755 1.278
H11 1.945 -0.755 -1.278
H12 0.484 -0.221 2.174
H13 0.475 -1.775 1.288
H14 0.484 -0.221 -2.174
H15 0.475 -1.775 -1.288

Atom - Atom Distances (Å)
  C1 S2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.85841.53871.53681.53682.41002.17472.19362.19362.17952.17952.18932.19192.18932.1919
S21.85842.71512.78882.78881.35233.71932.89802.89803.77873.77872.96643.01162.96643.0116
C31.53872.71512.52212.52213.71861.10561.10441.10442.78032.78032.76613.49332.76613.4933
C41.53682.78882.52212.52502.89832.76793.49272.77021.10712.77151.10341.10503.49262.7797
C51.53682.78882.52212.52502.89832.76792.77023.49272.77151.10713.49262.77971.10341.1050
H62.41001.35233.71862.89832.89834.56084.01844.01843.91173.91173.26162.59853.26162.5985
H72.17473.71931.10562.76792.76794.56081.79071.79072.56612.56613.11693.78733.11693.7873
H82.19362.89801.10443.49272.77024.01841.79071.79213.79573.13453.78444.35392.55503.7866
H92.19362.89801.10442.77023.49274.01841.79071.79213.13453.79572.55503.78663.78444.3539
H102.17953.77872.78031.10712.77153.91172.56613.79573.13452.55671.79481.78883.78683.1285
H112.17953.77872.78032.77151.10713.91172.56613.13453.79572.55673.78683.12851.79481.7888
H122.18932.96642.76611.10343.49263.26163.11693.78442.55501.79483.78681.78894.34883.7954
H132.19193.01163.49331.10502.77972.59853.78734.35393.78661.78883.12851.78893.79542.5767
H142.18932.96642.76613.49261.10343.26163.11692.55503.78443.78681.79484.34883.79541.7889
H152.19193.01163.49332.77971.10502.59853.78733.78664.35393.12851.78883.79542.57671.7889

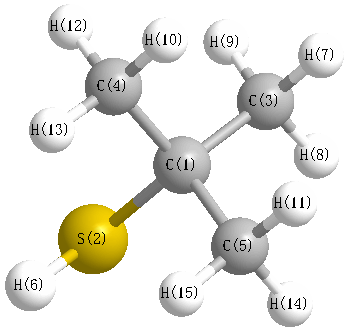
picture of 2-Propanethiol, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S2 H6 96.006 C1 C3 H7 109.565
C1 C3 H8 111.120 C1 C3 H9 111.120
C1 C4 H10 109.983 C1 C4 H12 110.974
C1 C4 H13 111.085 C1 C5 H11 109.983
C1 C5 H14 110.974 C1 C5 H15 111.085
S2 C1 C3 105.730 S2 C1 C4 110.090
S2 C1 C5 110.090 C3 C1 C4 110.180
C3 C1 C5 110.180 C4 C1 C5 110.469
H7 C3 H8 108.244 H7 C3 H9 108.244
H8 C3 H9 108.455 H10 C4 H12 108.575
H10 C4 H13 107.928 H11 C5 H14 108.575
H11 C5 H15 107.928 H12 C4 H13 108.200
H14 C5 H15 108.200
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability