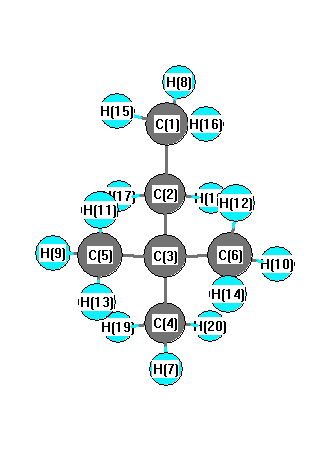
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3051 |
3033 |
41.77 |
|
|
|
| 2 |
A' |
3047 |
3029 |
49.73 |
|
|
|
| 3 |
A' |
3041 |
3023 |
59.25 |
|
|
|
| 4 |
A' |
3035 |
3018 |
0.96 |
|
|
|
| 5 |
A' |
2974 |
2957 |
22.21 |
|
|
|
| 6 |
A' |
2964 |
2947 |
10.78 |
|
|
|
| 7 |
A' |
2954 |
2937 |
34.50 |
|
|
|
| 8 |
A' |
2945 |
2928 |
32.67 |
|
|
|
| 9 |
A' |
1439 |
1431 |
10.03 |
|
|
|
| 10 |
A' |
1431 |
1423 |
4.76 |
|
|
|
| 11 |
A' |
1430 |
1421 |
6.40 |
|
|
|
| 12 |
A' |
1407 |
1399 |
0.02 |
|
|
|
| 13 |
A' |
1397 |
1389 |
0.26 |
|
|
|
| 14 |
A' |
1357 |
1349 |
4.21 |
|
|
|
| 15 |
A' |
1350 |
1342 |
4.48 |
|
|
|
| 16 |
A' |
1326 |
1319 |
5.41 |
|
|
|
| 17 |
A' |
1308 |
1300 |
0.21 |
|
|
|
| 18 |
A' |
1226 |
1219 |
3.49 |
|
|
|
| 19 |
A' |
1194 |
1187 |
8.59 |
|
|
|
| 20 |
A' |
1058 |
1052 |
0.85 |
|
|
|
| 21 |
A' |
1010 |
1004 |
4.88 |
|
|
|
| 22 |
A' |
967 |
962 |
2.53 |
|
|
|
| 23 |
A' |
906 |
901 |
1.14 |
|
|
|
| 24 |
A' |
857 |
852 |
0.29 |
|
|
|
| 25 |
A' |
703 |
699 |
0.37 |
|
|
|
| 26 |
A' |
469 |
467 |
0.42 |
|
|
|
| 27 |
A' |
392 |
389 |
0.05 |
|
|
|
| 28 |
A' |
340 |
338 |
0.01 |
|
|
|
| 29 |
A' |
251 |
250 |
0.06 |
|
|
|
| 30 |
A' |
248 |
246 |
0.07 |
|
|
|
| 31 |
A" |
3054 |
3036 |
28.51 |
|
|
|
| 32 |
A" |
3045 |
3027 |
20.63 |
|
|
|
| 33 |
A" |
3038 |
3020 |
30.14 |
|
|
|
| 34 |
A" |
3035 |
3018 |
0.04 |
|
|
|
| 35 |
A" |
2984 |
2966 |
20.58 |
|
|
|
| 36 |
A" |
2958 |
2941 |
37.45 |
|
|
|
| 37 |
A" |
1432 |
1424 |
0.00 |
|
|
|
| 38 |
A" |
1424 |
1416 |
9.42 |
|
|
|
| 39 |
A" |
1402 |
1394 |
0.05 |
|
|
|
| 40 |
A" |
1397 |
1389 |
0.07 |
|
|
|
| 41 |
A" |
1328 |
1320 |
7.35 |
|
|
|
| 42 |
A" |
1281 |
1274 |
0.55 |
|
|
|
| 43 |
A" |
1199 |
1192 |
1.54 |
|
|
|
| 44 |
A" |
1054 |
1048 |
0.60 |
|
|
|
| 45 |
A" |
956 |
951 |
0.52 |
|
|
|
| 46 |
A" |
911 |
905 |
0.44 |
|
|
|
| 47 |
A" |
908 |
903 |
0.10 |
|
|
|
| 48 |
A" |
764 |
760 |
2.70 |
|
|
|
| 49 |
A" |
406 |
404 |
0.06 |
|
|
|
| 50 |
A" |
325 |
323 |
0.01 |
|
|
|
| 51 |
A" |
274 |
273 |
0.00 |
|
|
|
| 52 |
A" |
245 |
244 |
0.00 |
|
|
|
| 53 |
A" |
202 |
201 |
0.01 |
|
|
|
| 54 |
A" |
92 |
92 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 39894.7 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 39663.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.048 |
|
|
|
| 2 |
C |
0.004 |
|
|
|
| 3 |
C |
-0.330 |
|
|
|
| 4 |
C |
0.032 |
|
|
|
| 5 |
C |
0.029 |
|
|
|
| 6 |
C |
0.029 |
|
|
|
| 7 |
H |
0.017 |
|
|
|
| 8 |
H |
0.028 |
|
|
|
| 9 |
H |
0.019 |
|
|
|
| 10 |
H |
0.019 |
|
|
|
| 11 |
H |
0.023 |
|
|
|
| 12 |
H |
0.023 |
|
|
|
| 13 |
H |
0.020 |
|
|
|
| 14 |
H |
0.020 |
|
|
|
| 15 |
H |
0.032 |
|
|
|
| 16 |
H |
0.032 |
|
|
|
| 17 |
H |
0.008 |
|
|
|
| 18 |
H |
0.008 |
|
|
|
| 19 |
H |
0.018 |
|
|
|
| 20 |
H |
0.018 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.049 |
-0.015 |
0.000 |
0.051 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.882 |
0.160 |
0.000 |
| y |
0.160 |
-42.371 |
0.000 |
| z |
0.000 |
0.000 |
-42.026 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.316 |
0.160 |
0.000 |
| y |
0.160 |
-0.417 |
0.000 |
| z |
0.000 |
0.000 |
0.100 |
|
| Polar |
|---|
| 3z2-r2 | 0.201 |
|---|
| x2-y2 | 0.489 |
|---|
| xy | 0.160 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.088 |
-0.488 |
0.000 |
| y |
-0.488 |
10.568 |
0.000 |
| z |
0.000 |
0.000 |
10.238 |
<r2> (average value of r
2) Å
2
| <r2> |
198.508 |
| (<r2>)1/2 |
14.089 |