Jump to
S1C2
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -637.668881 |
| Energy at 298.15K | -637.673775 |
| HF Energy | -637.009156 |
| Nuclear repulsion energy | 156.113740 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3104 |
2958 |
|
|
|
|
| 2 |
A' |
3072 |
2928 |
|
|
|
|
| 3 |
A' |
1511 |
1440 |
|
|
|
|
| 4 |
A' |
1489 |
1419 |
|
|
|
|
| 5 |
A' |
1421 |
1354 |
|
|
|
|
| 6 |
A' |
1273 |
1213 |
|
|
|
|
| 7 |
A' |
1097 |
1046 |
|
|
|
|
| 8 |
A' |
1071 |
1021 |
|
|
|
|
| 9 |
A' |
790 |
753 |
|
|
|
|
| 10 |
A' |
384 |
366 |
|
|
|
|
| 11 |
A' |
241 |
230 |
|
|
|
|
| 12 |
A" |
3175 |
3026 |
|
|
|
|
| 13 |
A" |
3132 |
2985 |
|
|
|
|
| 14 |
A" |
1294 |
1234 |
|
|
|
|
| 15 |
A" |
1217 |
1160 |
|
|
|
|
| 16 |
A" |
1058 |
1008 |
|
|
|
|
| 17 |
A" |
792 |
755 |
|
|
|
|
| 18 |
A" |
136 |
130 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13127.8 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 12512.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.618 |
0.000 |
| C2 |
0.997 |
-0.536 |
0.000 |
| Cl3 |
-1.675 |
-0.041 |
0.000 |
| F4 |
2.279 |
0.006 |
0.000 |
| H5 |
0.125 |
1.243 |
0.900 |
| H6 |
0.125 |
1.243 |
-0.900 |
| H7 |
0.870 |
-1.163 |
0.902 |
| H8 |
0.870 |
-1.163 |
-0.902 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5247 | 1.8002 | 2.3596 | 1.1027 | 1.1027 | 2.1770 | 2.1770 |
C2 | 1.5247 | | 2.7176 | 1.3914 | 2.1756 | 2.1756 | 1.1057 | 1.1057 | Cl3 | 1.8002 | 2.7176 | | 3.9543 | 2.3871 | 2.3871 | 2.9236 | 2.9236 | F4 | 2.3596 | 1.3914 | 3.9543 | | 2.6420 | 2.6420 | 2.0405 | 2.0405 | H5 | 1.1027 | 2.1756 | 2.3871 | 2.6420 | | 1.8000 | 2.5178 | 3.0962 | H6 | 1.1027 | 2.1756 | 2.3871 | 2.6420 | 1.8000 | | 3.0962 | 2.5178 | H7 | 2.1770 | 1.1057 | 2.9236 | 2.0405 | 2.5178 | 3.0962 | | 1.8039 | H8 | 2.1770 | 1.1057 | 2.9236 | 2.0405 | 3.0962 | 2.5178 | 1.8039 | |
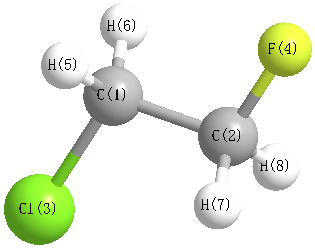 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.941 |
|
C1 |
C2 |
H7 |
110.705 |
| C1 |
C2 |
H8 |
110.705 |
|
C2 |
C1 |
Cl3 |
109.360 |
| C2 |
C1 |
H5 |
110.775 |
|
C2 |
C1 |
H6 |
110.775 |
| Cl3 |
C1 |
H5 |
108.222 |
|
Cl3 |
C1 |
H6 |
108.222 |
| F4 |
C2 |
H7 |
109.065 |
|
F4 |
C2 |
H8 |
109.065 |
| H5 |
C1 |
H6 |
109.414 |
|
H7 |
C2 |
H8 |
109.318 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -637.667835 |
| Energy at 298.15K | -637.672826 |
| HF Energy | -637.007751 |
| Nuclear repulsion energy | 160.214079 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3163 |
3015 |
|
|
|
|
| 2 |
A |
3117 |
2970 |
|
|
|
|
| 3 |
A |
3093 |
2948 |
|
|
|
|
| 4 |
A |
3049 |
2906 |
|
|
|
|
| 5 |
A |
1494 |
1424 |
|
|
|
|
| 6 |
A |
1458 |
1390 |
|
|
|
|
| 7 |
A |
1427 |
1360 |
|
|
|
|
| 8 |
A |
1331 |
1268 |
|
|
|
|
| 9 |
A |
1271 |
1212 |
|
|
|
|
| 10 |
A |
1215 |
1158 |
|
|
|
|
| 11 |
A |
1130 |
1077 |
|
|
|
|
| 12 |
A |
1064 |
1014 |
|
|
|
|
| 13 |
A |
981 |
935 |
|
|
|
|
| 14 |
A |
857 |
817 |
|
|
|
|
| 15 |
A |
693 |
661 |
|
|
|
|
| 16 |
A |
463 |
441 |
|
|
|
|
| 17 |
A |
285 |
271 |
|
|
|
|
| 18 |
A |
134 |
127 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13111.9 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 12497.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.087 |
0.854 |
-0.295 |
| C2 |
1.211 |
0.413 |
0.361 |
| Cl3 |
-1.435 |
-0.286 |
0.065 |
| F4 |
1.660 |
-0.770 |
-0.204 |
| H5 |
-0.379 |
1.850 |
0.080 |
| H6 |
0.032 |
0.892 |
-1.391 |
| H7 |
1.978 |
1.196 |
0.198 |
| H8 |
1.070 |
0.259 |
1.447 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5195 | 1.8014 | 2.3868 | 1.1040 | 1.1032 | 2.1506 | 2.1738 |
C2 | 1.5195 | | 2.7523 | 1.3855 | 2.1619 | 2.1657 | 1.1088 | 1.1060 | Cl3 | 1.8014 | 2.7523 | | 3.1441 | 2.3827 | 2.3788 | 3.7233 | 2.9117 | F4 | 2.3868 | 1.3855 | 3.1441 | | 3.3323 | 2.6123 | 2.0319 | 2.0332 | H5 | 1.1040 | 2.1619 | 2.3827 | 3.3323 | | 1.8026 | 2.4492 | 2.5495 | H6 | 1.1032 | 2.1657 | 2.3788 | 2.6123 | 1.8026 | | 2.5310 | 3.0876 | H7 | 2.1506 | 1.1088 | 3.7233 | 2.0319 | 2.4492 | 2.5310 | | 1.8069 | H8 | 2.1738 | 1.1060 | 2.9117 | 2.0332 | 2.5495 | 3.0876 | 1.8069 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.406 |
|
C1 |
C2 |
H7 |
108.811 |
| C1 |
C2 |
H8 |
110.790 |
|
C2 |
C1 |
Cl3 |
111.666 |
| C2 |
C1 |
H5 |
109.974 |
|
C2 |
C1 |
H6 |
110.319 |
| Cl3 |
C1 |
H5 |
107.767 |
|
Cl3 |
C1 |
H6 |
107.524 |
| F4 |
C2 |
H7 |
108.592 |
|
F4 |
C2 |
H8 |
108.868 |
| H5 |
C1 |
H6 |
109.513 |
|
H7 |
C2 |
H8 |
109.337 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
