Jump to
S1C2
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -637.654336 |
| Energy at 298.15K | -637.659252 |
| HF Energy | -637.009446 |
| Nuclear repulsion energy | 156.338139 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3122 |
2995 |
16.98 |
|
|
|
| 2 |
A' |
3091 |
2965 |
23.13 |
|
|
|
| 3 |
A' |
1522 |
1460 |
0.42 |
|
|
|
| 4 |
A' |
1500 |
1439 |
5.18 |
|
|
|
| 5 |
A' |
1435 |
1377 |
6.84 |
|
|
|
| 6 |
A' |
1286 |
1234 |
13.86 |
|
|
|
| 7 |
A' |
1109 |
1064 |
56.41 |
|
|
|
| 8 |
A' |
1084 |
1040 |
55.32 |
|
|
|
| 9 |
A' |
801 |
768 |
40.12 |
|
|
|
| 10 |
A' |
387 |
372 |
2.33 |
|
|
|
| 11 |
A' |
245 |
235 |
10.93 |
|
|
|
| 12 |
A" |
3192 |
3062 |
13.90 |
|
|
|
| 13 |
A" |
3149 |
3021 |
20.66 |
|
|
|
| 14 |
A" |
1305 |
1252 |
0.06 |
|
|
|
| 15 |
A" |
1228 |
1178 |
1.71 |
|
|
|
| 16 |
A" |
1067 |
1024 |
2.49 |
|
|
|
| 17 |
A" |
799 |
766 |
0.20 |
|
|
|
| 18 |
A" |
136 |
130 |
9.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13228.1 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 12691.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.616 |
0.000 |
| C2 |
0.996 |
-0.536 |
0.000 |
| Cl3 |
-1.673 |
-0.039 |
0.000 |
| F4 |
2.276 |
0.002 |
0.000 |
| H5 |
0.125 |
1.241 |
0.898 |
| H6 |
0.125 |
1.241 |
-0.898 |
| H7 |
0.869 |
-1.162 |
0.900 |
| H8 |
0.869 |
-1.162 |
-0.900 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5232 | 1.7971 | 2.3574 | 1.1011 | 1.1011 | 2.1745 | 2.1745 |
C2 | 1.5232 | | 2.7155 | 1.3879 | 2.1732 | 2.1732 | 1.1042 | 1.1042 | Cl3 | 1.7971 | 2.7155 | | 3.9495 | 2.3826 | 2.3826 | 2.9213 | 2.9213 | F4 | 2.3574 | 1.3879 | 3.9495 | | 2.6400 | 2.6400 | 2.0362 | 2.0362 | H5 | 1.1011 | 2.1732 | 2.3826 | 2.6400 | | 1.7969 | 2.5153 | 3.0923 | H6 | 1.1011 | 2.1732 | 2.3826 | 2.6400 | 1.7969 | | 3.0923 | 2.5153 | H7 | 2.1745 | 1.1042 | 2.9213 | 2.0362 | 2.5153 | 3.0923 | | 1.8005 | H8 | 2.1745 | 1.1042 | 2.9213 | 2.0362 | 3.0923 | 2.5153 | 1.8005 | |
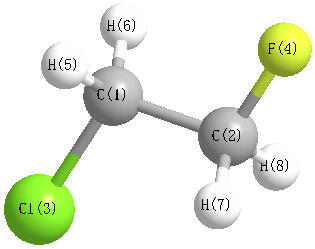 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.061 |
|
C1 |
C2 |
H7 |
110.699 |
| C1 |
C2 |
H8 |
110.699 |
|
C2 |
C1 |
Cl3 |
109.469 |
| C2 |
C1 |
H5 |
110.781 |
|
C2 |
C1 |
H6 |
110.781 |
| Cl3 |
C1 |
H5 |
108.183 |
|
Cl3 |
C1 |
H6 |
108.183 |
| F4 |
C2 |
H7 |
109.053 |
|
F4 |
C2 |
H8 |
109.053 |
| H5 |
C1 |
H6 |
109.369 |
|
H7 |
C2 |
H8 |
109.238 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -637.653299 |
| Energy at 298.15K | -637.658315 |
| HF Energy | -637.008043 |
| Nuclear repulsion energy | 160.450765 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3180 |
3051 |
9.29 |
|
|
|
| 2 |
A |
3134 |
3007 |
30.51 |
|
|
|
| 3 |
A |
3111 |
2985 |
15.19 |
|
|
|
| 4 |
A |
3068 |
2943 |
31.54 |
|
|
|
| 5 |
A |
1506 |
1445 |
1.87 |
|
|
|
| 6 |
A |
1470 |
1411 |
10.44 |
|
|
|
| 7 |
A |
1441 |
1383 |
16.54 |
|
|
|
| 8 |
A |
1344 |
1289 |
30.41 |
|
|
|
| 9 |
A |
1284 |
1231 |
0.30 |
|
|
|
| 10 |
A |
1227 |
1177 |
3.79 |
|
|
|
| 11 |
A |
1143 |
1097 |
58.97 |
|
|
|
| 12 |
A |
1075 |
1031 |
19.61 |
|
|
|
| 13 |
A |
991 |
951 |
5.14 |
|
|
|
| 14 |
A |
866 |
831 |
9.75 |
|
|
|
| 15 |
A |
703 |
674 |
26.06 |
|
|
|
| 16 |
A |
468 |
449 |
11.66 |
|
|
|
| 17 |
A |
287 |
275 |
0.75 |
|
|
|
| 18 |
A |
135 |
129 |
2.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13215.5 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 12679.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.088 |
0.851 |
-0.294 |
| C2 |
1.210 |
0.412 |
0.360 |
| Cl3 |
-1.433 |
-0.286 |
0.065 |
| F4 |
1.659 |
-0.768 |
-0.204 |
| H5 |
-0.380 |
1.846 |
0.081 |
| H6 |
0.030 |
0.892 |
-1.389 |
| H7 |
1.975 |
1.195 |
0.197 |
| H8 |
1.070 |
0.258 |
1.445 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5178 | 1.7979 | 2.3835 | 1.1024 | 1.1016 | 2.1483 | 2.1714 |
C2 | 1.5178 | | 2.7492 | 1.3824 | 2.1589 | 2.1634 | 1.1072 | 1.1045 | Cl3 | 1.7979 | 2.7492 | | 3.1412 | 2.3778 | 2.3746 | 3.7185 | 2.9094 | F4 | 2.3835 | 1.3824 | 3.1412 | | 3.3272 | 2.6103 | 2.0278 | 2.0292 | H5 | 1.1024 | 2.1589 | 2.3778 | 3.3272 | | 1.7996 | 2.4464 | 2.5459 | H6 | 1.1016 | 2.1634 | 2.3746 | 2.6103 | 1.7996 | | 2.5284 | 3.0841 | H7 | 2.1483 | 1.1072 | 3.7185 | 2.0278 | 2.4464 | 2.5284 | | 1.8037 | H8 | 2.1714 | 1.1045 | 2.9094 | 2.0292 | 2.5459 | 3.0841 | 1.8037 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.450 |
|
C1 |
C2 |
H7 |
108.843 |
| C1 |
C2 |
H8 |
110.811 |
|
C2 |
C1 |
Cl3 |
111.742 |
| C2 |
C1 |
H5 |
109.949 |
|
C2 |
C1 |
H6 |
110.353 |
| Cl3 |
C1 |
H5 |
107.713 |
|
Cl3 |
C1 |
H6 |
107.528 |
| F4 |
C2 |
H7 |
108.574 |
|
F4 |
C2 |
H8 |
108.845 |
| H5 |
C1 |
H6 |
109.475 |
|
H7 |
C2 |
H8 |
109.279 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
