Jump to
S1C2
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -637.649562 |
| Energy at 298.15K | -637.654496 |
| HF Energy | -637.009581 |
| Nuclear repulsion energy | 156.549026 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3129 |
2993 |
16.84 |
|
|
|
| 2 |
A' |
3100 |
2965 |
21.90 |
|
|
|
| 3 |
A' |
1526 |
1460 |
0.68 |
|
|
|
| 4 |
A' |
1504 |
1439 |
5.51 |
|
|
|
| 5 |
A' |
1444 |
1381 |
7.36 |
|
|
|
| 6 |
A' |
1294 |
1238 |
12.77 |
|
|
|
| 7 |
A' |
1123 |
1074 |
92.12 |
|
|
|
| 8 |
A' |
1096 |
1048 |
19.92 |
|
|
|
| 9 |
A' |
815 |
780 |
37.44 |
|
|
|
| 10 |
A' |
391 |
374 |
2.45 |
|
|
|
| 11 |
A' |
247 |
236 |
10.84 |
|
|
|
| 12 |
A" |
3200 |
3061 |
13.82 |
|
|
|
| 13 |
A" |
3158 |
3021 |
19.56 |
|
|
|
| 14 |
A" |
1311 |
1254 |
0.10 |
|
|
|
| 15 |
A" |
1234 |
1180 |
1.87 |
|
|
|
| 16 |
A" |
1072 |
1026 |
2.52 |
|
|
|
| 17 |
A" |
800 |
766 |
0.21 |
|
|
|
| 18 |
A" |
136 |
131 |
9.43 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13288.6 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 12713.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.616 |
0.000 |
| C2 |
0.995 |
-0.538 |
0.000 |
| Cl3 |
-1.671 |
-0.034 |
0.000 |
| F4 |
2.273 |
-0.006 |
0.000 |
| H5 |
0.128 |
1.240 |
0.898 |
| H6 |
0.128 |
1.240 |
-0.898 |
| H7 |
0.865 |
-1.164 |
0.900 |
| H8 |
0.865 |
-1.164 |
-0.900 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5232 | 1.7929 | 2.3561 | 1.1008 | 1.1008 | 2.1737 | 2.1737 |
C2 | 1.5232 | | 2.7135 | 1.3839 | 2.1721 | 2.1721 | 1.1038 | 1.1038 | Cl3 | 1.7929 | 2.7135 | | 3.9440 | 2.3797 | 2.3797 | 2.9190 | 2.9190 | F4 | 2.3561 | 1.3839 | 3.9440 | | 2.6380 | 2.6380 | 2.0326 | 2.0326 | H5 | 1.1008 | 2.1721 | 2.3797 | 2.6380 | | 1.7954 | 2.5145 | 3.0908 | H6 | 1.1008 | 2.1721 | 2.3797 | 2.6380 | 1.7954 | | 3.0908 | 2.5145 | H7 | 2.1737 | 1.1038 | 2.9190 | 2.0326 | 2.5145 | 3.0908 | | 1.7991 | H8 | 2.1737 | 1.1038 | 2.9190 | 2.0326 | 3.0908 | 2.5145 | 1.7991 | |
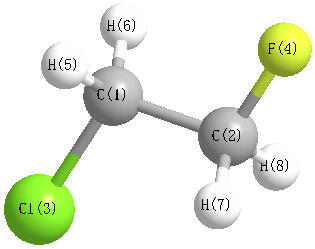 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.186 |
|
C1 |
C2 |
H7 |
110.659 |
| C1 |
C2 |
H8 |
110.659 |
|
C2 |
C1 |
Cl3 |
109.560 |
| C2 |
C1 |
H5 |
110.713 |
|
C2 |
C1 |
H6 |
110.713 |
| Cl3 |
C1 |
H5 |
108.261 |
|
Cl3 |
C1 |
H6 |
108.261 |
| F4 |
C2 |
H7 |
109.065 |
|
F4 |
C2 |
H8 |
109.065 |
| H5 |
C1 |
H6 |
109.264 |
|
H7 |
C2 |
H8 |
109.171 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -637.648555 |
| Energy at 298.15K | -637.653588 |
| HF Energy | -637.008186 |
| Nuclear repulsion energy | 160.648288 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3188 |
3050 |
8.94 |
|
|
|
| 2 |
A |
3144 |
3008 |
29.25 |
|
|
|
| 3 |
A |
3119 |
2984 |
14.60 |
|
|
|
| 4 |
A |
3078 |
2945 |
29.98 |
|
|
|
| 5 |
A |
1511 |
1445 |
2.42 |
|
|
|
| 6 |
A |
1474 |
1410 |
11.19 |
|
|
|
| 7 |
A |
1448 |
1386 |
16.17 |
|
|
|
| 8 |
A |
1352 |
1293 |
29.33 |
|
|
|
| 9 |
A |
1290 |
1235 |
0.17 |
|
|
|
| 10 |
A |
1234 |
1181 |
5.19 |
|
|
|
| 11 |
A |
1157 |
1107 |
62.75 |
|
|
|
| 12 |
A |
1081 |
1034 |
15.70 |
|
|
|
| 13 |
A |
997 |
954 |
4.39 |
|
|
|
| 14 |
A |
871 |
833 |
8.82 |
|
|
|
| 15 |
A |
714 |
683 |
24.26 |
|
|
|
| 16 |
A |
471 |
451 |
11.59 |
|
|
|
| 17 |
A |
288 |
276 |
0.75 |
|
|
|
| 18 |
A |
135 |
130 |
2.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13276.7 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 12701.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.089 |
0.849 |
-0.293 |
| C2 |
1.209 |
0.410 |
0.359 |
| Cl3 |
-1.432 |
-0.285 |
0.065 |
| F4 |
1.659 |
-0.765 |
-0.204 |
| H5 |
-0.380 |
1.844 |
0.082 |
| H6 |
0.029 |
0.891 |
-1.387 |
| H7 |
1.974 |
1.194 |
0.197 |
| H8 |
1.071 |
0.256 |
1.444 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5178 | 1.7940 | 2.3810 | 1.1020 | 1.1014 | 2.1481 | 2.1708 |
C2 | 1.5178 | | 2.7468 | 1.3789 | 2.1580 | 2.1623 | 1.1066 | 1.1041 | Cl3 | 1.7940 | 2.7468 | | 3.1389 | 2.3751 | 2.3720 | 3.7152 | 2.9079 | F4 | 2.3810 | 1.3789 | 3.1389 | | 3.3234 | 2.6081 | 2.0245 | 2.0260 | H5 | 1.1020 | 2.1580 | 2.3751 | 3.3234 | | 1.7980 | 2.4444 | 2.5452 | H6 | 1.1014 | 2.1623 | 2.3720 | 2.6081 | 1.7980 | | 2.5270 | 3.0827 | H7 | 2.1481 | 1.1066 | 3.7152 | 2.0245 | 2.4444 | 2.5270 | | 1.8023 | H8 | 2.1708 | 1.1041 | 2.9079 | 2.0260 | 2.5452 | 3.0827 | 1.8023 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.481 |
|
C1 |
C2 |
H7 |
108.863 |
| C1 |
C2 |
H8 |
110.794 |
|
C2 |
C1 |
Cl3 |
111.807 |
| C2 |
C1 |
H5 |
109.904 |
|
C2 |
C1 |
H6 |
110.280 |
| Cl3 |
C1 |
H5 |
107.792 |
|
Cl3 |
C1 |
H6 |
107.604 |
| F4 |
C2 |
H7 |
108.584 |
|
F4 |
C2 |
H8 |
108.859 |
| H5 |
C1 |
H6 |
109.375 |
|
H7 |
C2 |
H8 |
109.220 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
