return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 ((Z)-hex-3-ene)

using model chemistry: QCISD(T)/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at QCISD(T)/cc-pVDZ
 hartrees
Energy at 0K-235.155075
Energy at 298.15K-235.167323
HF Energy-234.191869
Nuclear repulsion energy228.807478
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3147 2999        
2 A 3127 2980        
3 A 3123 2977        
4 A 3103 2958        
5 A 3038 2896        
6 A 3034 2892        
7 A 1717 1636        
8 A 1496 1426        
9 A 1485 1415        
10 A 1477 1408        
11 A 1402 1336        
12 A 1341 1278        
13 A 1292 1231        
14 A 1262 1202        
15 A 1088 1037        
16 A 1063 1013        
17 A 1029 981        
18 A 976 930        
19 A 849 809        
20 A 771 735        
21 A 501 477        
22 A 309 295        
23 A 209 199        
24 A 193 184        
25 A 43 41        
26 B 3127 2980        
27 B 3123 2977        
28 B 3119 2972        
29 B 3090 2945        
30 B 3037 2895        
31 B 3032 2890        
32 B 1494 1424        
33 B 1485 1416        
34 B 1468 1400        
35 B 1432 1365        
36 B 1402 1336        
37 B 1319 1258        
38 B 1279 1219        
39 B 1172 1117        
40 B 1092 1041        
41 B 1054 1005        
42 B 913 870        
43 B 803 765        
44 B 747 712        
45 B 557 531        
46 B 356 339        
47 B 244 233        
48 B 37 35        

Unscaled Zero Point Vibrational Energy (zpe) 36226.9 cm-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 34527.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/cc-pVDZ
ABC
0.28218 0.04971 0.04734

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/cc-pVDZ

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.004 0.679 0.862
C2 -0.004 -0.679 0.862
C3 -0.004 1.604 -0.338
C4 0.004 -1.604 -0.338
C5 -1.271 2.482 -0.377
C6 1.271 -2.482 -0.377
H7 0.001 1.186 1.841
H8 -0.001 -1.186 1.841
H9 0.081 1.030 -1.280
H10 -0.081 -1.030 -1.280
H11 0.890 2.261 -0.287
H12 -0.890 -2.261 -0.287
H13 -1.246 3.184 -1.233
H14 -2.178 1.854 -0.469
H15 -1.365 3.078 0.551
H16 1.246 -3.184 -1.233
H17 2.178 -1.854 -0.469
H18 1.365 -3.078 0.551

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.35731.51512.57862.53183.62321.10322.10622.17122.74072.14683.28033.49592.81272.78014.56553.59324.0080
C21.35732.57861.51513.62322.53182.10621.10322.74072.17123.28032.14684.56553.59324.00803.49592.81272.7801
C31.51512.57863.20801.54174.28002.21913.54011.10592.79841.11023.96572.19942.19222.19495.02814.09074.9586
C42.57861.51513.20804.28001.54173.54012.21912.79841.10593.96571.11025.02814.09074.95862.19942.19222.1949
C52.53183.62321.54174.28005.57612.86684.47052.17913.81592.17324.75901.10671.10691.10776.25765.54056.2228
C63.62322.53184.28001.54175.57614.47052.86683.81592.17914.75902.17326.25765.54056.22281.10671.10691.1077
H71.10322.10622.21913.54012.86684.47052.37163.12583.82842.54504.14793.87223.24522.66705.48544.39514.6592
H82.10621.10323.54012.21914.47052.86682.37163.82843.12584.14792.54505.48544.39514.65923.87223.24522.6670
H92.17122.74071.10592.79842.17913.81593.12583.82842.06671.77573.57222.52982.53743.10464.37183.65654.6775
H102.74072.17122.79841.10593.81592.17913.82843.12582.06673.57221.77574.37183.65654.67752.52982.53743.1046
H112.14683.28031.11023.96572.17324.75902.54504.14791.77573.57224.85972.51063.09942.54065.53774.31605.4258
H123.28032.14683.96571.11024.75902.17324.14792.54503.57221.77574.85975.53774.31605.42582.51063.09942.5406
H133.49594.56552.19945.02811.10676.25763.87225.48542.52984.37182.51065.53771.79431.79096.83706.13837.0149
H142.81273.59322.19224.09071.10695.54053.24524.39512.53743.65653.09944.31601.79431.78876.13835.72006.1580
H152.78014.00802.19494.95861.10776.22282.66704.65923.10464.67752.54065.42581.79091.78877.01496.15806.7349
H164.56553.49595.02812.19946.25761.10675.48543.87224.37182.52985.53772.51066.83706.13837.01491.79431.7909
H173.59322.81274.09072.19225.54051.10694.39513.24523.65652.53744.31603.09946.13835.72006.15801.79431.7887
H184.00802.78014.95862.19496.22281.10774.65922.66704.67753.10465.42582.54067.01496.15806.73491.79091.7887

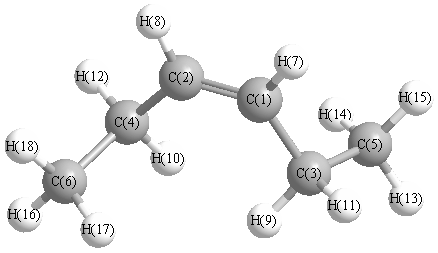
picture of (Z)-hex-3-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 127.644 C1 C2 H8 117.367
C1 C3 C5 111.840 C1 C3 H9 110.903
C1 C3 H11 108.747 C2 C1 C3 127.644
C2 C1 H7 117.367 C2 C4 C6 111.840
C2 C4 H10 110.903 C2 C4 H12 108.747
C3 C1 H7 114.983 C3 C5 H13 111.234
C3 C5 H14 110.653 C3 C5 H15 110.824
C4 C2 H8 114.983 C4 C6 H16 111.234
C4 C6 H17 110.653 C4 C6 H18 110.824
C5 C3 H9 109.687 C5 C3 H11 108.986
C6 C4 H10 109.687 C6 C4 H12 108.986
H9 C3 H11 106.512 H10 C4 H12 106.512
H13 C5 H14 108.305 H13 C5 H15 107.949
H14 C5 H15 107.748 H16 C6 H17 108.305
H16 C6 H18 107.949 H17 C6 H18 107.748
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability