return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 ((Z)-hex-3-ene)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-235.125295
Energy at 298.15K-235.137626
HF Energy-234.192696
Nuclear repulsion energy229.414187
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3168 3039 50.83      
2 A 3141 3014 13.92      
3 A 3138 3010 60.31      
4 A 3120 2993 0.06      
5 A 3056 2932 5.48      
6 A 3051 2927 0.33      
7 A 1751 1680 0.67      
8 A 1508 1446 6.19      
9 A 1495 1434 2.71      
10 A 1490 1430 0.04      
11 A 1415 1358 0.08      
12 A 1357 1302 0.74      
13 A 1305 1252 0.02      
14 A 1274 1222 0.15      
15 A 1099 1054 0.18      
16 A 1075 1031 0.30      
17 A 1038 996 0.30      
18 A 999 959 0.56      
19 A 855 821 0.69      
20 A 780 748 0.47      
21 A 515 494 0.17      
22 A 315 302 0.01      
23 A 212 203 0.01      
24 A 197 189 0.01      
25 A 42 40 0.01      
26 B 3144 3016 44.14      
27 B 3139 3012 42.91      
28 B 3134 3007 0.11      
29 B 3108 2981 6.45      
30 B 3054 2930 40.16      
31 B 3050 2926 76.09      
32 B 1504 1443 2.41      
33 B 1495 1434 6.30      
34 B 1481 1421 0.56      
35 B 1447 1388 1.33      
36 B 1415 1358 1.53      
37 B 1336 1282 15.81      
38 B 1292 1239 0.06      
39 B 1184 1136 0.10      
40 B 1104 1059 6.34      
41 B 1064 1021 1.61      
42 B 920 883 4.31      
43 B 812 779 2.56      
44 B 761 730 27.86      
45 B 562 539 4.18      
46 B 361 346 0.72      
47 B 246 236 0.04      
48 B 42 40 0.06      

Unscaled Zero Point Vibrational Energy (zpe) 36523.8 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 35040.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.27456 0.05053 0.04811

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.005 0.676 0.873
C2 -0.005 -0.676 0.873
C3 -0.005 1.602 -0.324
C4 0.005 -1.602 -0.324
C5 -1.300 2.431 -0.399
C6 1.300 -2.431 -0.399
H7 -0.000 1.182 1.851
H8 0.000 -1.182 1.851
H9 0.128 1.035 -1.263
H10 -0.128 -1.035 -1.263
H11 0.860 2.290 -0.248
H12 -0.860 -2.290 -0.248
H13 -1.274 3.137 -1.249
H14 -2.180 1.773 -0.521
H15 -1.445 3.019 0.527
H16 1.274 -3.137 -1.249
H17 2.180 -1.773 -0.521
H18 1.445 -3.019 0.527

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.35151.51382.57302.53053.59831.10142.09942.16932.73962.14343.28663.49292.81452.77764.54463.55923.9805
C21.35152.57301.51383.59832.53052.09941.10142.73962.16933.28662.14344.54463.55923.98053.49292.81452.7776
C31.51382.57303.20321.53954.23962.21583.53291.10422.80121.10843.98502.19632.18992.19284.99514.02514.9174
C42.57301.51383.20324.23961.53953.53292.21582.80121.10423.98501.10844.99514.02514.91742.19632.18992.1928
C52.53053.59831.53954.23965.51422.88324.45062.17633.75972.16984.74421.10541.10551.10636.19385.45906.1727
C63.59832.53054.23961.53955.51424.45062.88323.75972.17634.74422.16986.19385.45906.17271.10541.10551.1063
H71.10142.09942.21583.53292.88324.45062.36373.12013.82452.52514.14743.88043.27512.68645.46734.37154.6358
H82.09941.10143.53292.21584.45062.88322.36373.82453.12014.14742.52515.46734.37154.63583.88043.27512.6864
H92.16932.73961.10422.80122.17633.75973.12013.82452.08561.77193.61392.52752.53403.10114.32703.55584.6232
H102.73962.16932.80121.10423.75972.17633.82453.12012.08563.61391.77194.32703.55584.62322.52752.53403.1011
H112.14343.28661.10843.98502.16984.74422.52514.14741.77193.61394.89232.50513.09532.53935.53444.28075.3972
H123.28662.14343.98501.10844.74422.16984.14742.52513.61391.77194.89235.53444.28075.39722.50513.09532.5393
H133.49294.54462.19634.99511.10546.19383.88045.46732.52754.32702.50515.53441.79181.78816.77296.04776.9609
H142.81453.55922.18994.02511.10555.45903.27514.37152.53403.55583.09534.28071.79181.78626.04775.61956.0995
H152.77763.98052.19284.91741.10636.17272.68644.63583.10114.62322.53935.39721.78811.78626.96096.09956.6944
H164.54463.49294.99512.19636.19381.10545.46733.88044.32702.52755.53442.50516.77296.04776.96091.79181.7881
H173.55922.81454.02512.18995.45901.10554.37153.27513.55582.53404.28073.09536.04775.61956.09951.79181.7862
H183.98052.77764.91742.19286.17271.10634.63582.68644.62323.10115.39722.53936.96096.09956.69441.78811.7862

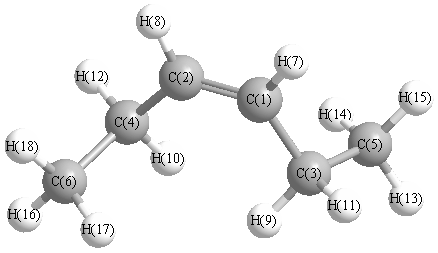
picture of (Z)-hex-3-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 127.703 C1 C2 H8 117.356
C1 C3 C5 111.943 C1 C3 H9 110.944
C1 C3 H11 108.666 C2 C1 C3 127.703
C2 C1 H7 117.356 C2 C4 C6 111.943
C2 C4 H10 110.944 C2 C4 H12 108.666
C3 C1 H7 114.927 C3 C5 H13 111.222
C3 C5 H14 110.702 C3 C5 H15 110.886
C4 C2 H8 114.927 C4 C6 H16 111.222
C4 C6 H17 110.702 C4 C6 H18 110.886
C5 C3 H9 109.716 C5 C3 H11 108.973
C6 C4 H10 109.716 C6 C4 H12 108.973
H9 C3 H11 106.420 H10 C4 H12 106.420
H13 C5 H14 108.277 H13 C5 H15 107.903
H14 C5 H15 107.719 H16 C6 H17 108.277
H16 C6 H18 107.903 H17 C6 H18 107.719
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability