return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 ((Z)-hex-3-ene)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-235.119439
Energy at 298.15K-235.131786
HF Energy-234.192939
Nuclear repulsion energy229.567479
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3177 3040 50.54      
2 A 3154 3017 9.07      
3 A 3150 3014 59.33      
4 A 3132 2997 0.00      
5 A 3066 2934 5.76      
6 A 3062 2929 0.05      
7 A 1765 1689 0.71      
8 A 1510 1445 6.50      
9 A 1497 1433 2.80      
10 A 1493 1428 0.07      
11 A 1418 1357 0.06      
12 A 1360 1302 0.72      
13 A 1308 1251 0.02      
14 A 1277 1222 0.13      
15 A 1102 1055 0.20      
16 A 1077 1031 0.29      
17 A 1042 997 0.25      
18 A 1006 962 0.61      
19 A 858 821 0.75      
20 A 782 748 0.50      
21 A 518 496 0.17      
22 A 316 302 0.01      
23 A 212 203 0.01      
24 A 197 189 0.01      
25 A 42 40 0.01      
26 B 3154 3018 66.66      
27 B 3151 3015 15.27      
28 B 3147 3010 0.28      
29 B 3120 2985 6.70      
30 B 3064 2931 35.97      
31 B 3061 2928 73.95      
32 B 1507 1442 2.72      
33 B 1497 1433 6.49      
34 B 1483 1419 0.51      
35 B 1451 1388 1.36      
36 B 1419 1357 1.78      
37 B 1340 1282 14.00      
38 B 1294 1238 0.06      
39 B 1187 1135 0.10      
40 B 1106 1059 6.43      
41 B 1068 1021 1.46      
42 B 922 882 4.11      
43 B 814 778 2.62      
44 B 764 731 28.18      
45 B 563 539 4.16      
46 B 362 346 0.70      
47 B 246 235 0.04      
48 B 42 40 0.06      

Unscaled Zero Point Vibrational Energy (zpe) 36640.7 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 35054.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.27367 0.05068 0.04825

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.005 0.675 0.875
C2 -0.005 -0.675 0.875
C3 -0.005 1.600 -0.322
C4 0.005 -1.600 -0.322
C5 -1.303 2.424 -0.401
C6 1.303 -2.424 -0.401
H7 -0.000 1.181 1.852
H8 0.000 -1.181 1.852
H9 0.134 1.035 -1.260
H10 -0.134 -1.035 -1.260
H11 0.856 2.292 -0.243
H12 -0.856 -2.292 -0.243
H13 -1.278 3.130 -1.251
H14 -2.179 1.762 -0.527
H15 -1.455 3.010 0.524
H16 1.278 -3.130 -1.251
H17 2.179 -1.762 -0.527
H18 1.455 -3.010 0.524

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.34911.51332.57062.53013.59351.10102.09702.16862.73832.14263.28553.49202.81412.77714.54003.55323.9753
C21.34912.57061.51333.59352.53012.09701.10102.73832.16863.28552.14264.54003.55323.97533.49202.81412.7771
C31.51332.57063.20071.53934.23252.21493.53031.10372.80011.10793.98552.19582.18932.19224.98854.01464.9099
C42.57061.51333.20074.23251.53933.53032.21492.80011.10373.98551.10794.98854.01464.90992.19582.18932.1922
C52.53013.59351.53934.23255.50432.88514.44652.17573.75072.16944.74021.10481.10501.10576.18325.44646.1641
C63.59352.53014.23251.53935.50434.44652.88513.75072.17574.74022.16946.18325.44646.16411.10481.10501.1057
H71.10102.09702.21493.53032.88514.44652.36123.11863.82272.52214.14563.88103.27832.68895.46304.36684.6311
H82.09701.10103.53032.21494.44652.88512.36123.82273.11864.14562.52215.46304.36684.63113.88103.27832.6889
H92.16862.73831.10372.80012.17573.75073.11863.82272.08731.77093.61752.52672.53323.10004.31923.54114.6141
H102.73832.16862.80011.10373.75072.17573.82273.11862.08733.61751.77094.31923.54114.61412.52672.53323.1000
H112.14263.28551.10793.98552.16944.74022.52214.14561.77093.61754.89422.50433.09432.53895.53144.27405.3913
H123.28552.14263.98551.10794.74022.16944.14562.52213.61751.77094.89425.53144.27405.39132.50433.09432.5389
H133.49204.54002.19584.98851.10486.18323.88105.46302.52674.31922.50435.53141.79101.78736.76186.03366.9515
H142.81413.55322.18934.01461.10505.44643.27834.36682.53323.54113.09434.27401.79101.78536.03365.60436.0897
H152.77713.97532.19224.90991.10576.16412.68894.63113.10004.61412.53895.39131.78731.78536.95156.08976.6872
H164.54003.49204.98852.19586.18321.10485.46303.88104.31922.52675.53142.50436.76186.03366.95151.79101.7873
H173.55322.81414.01462.18935.44641.10504.36683.27833.54112.53324.27403.09436.03365.60436.08971.79101.7853
H183.97532.77714.90992.19226.16411.10574.63112.68894.61413.10005.39132.53896.95156.08976.68721.78731.7853

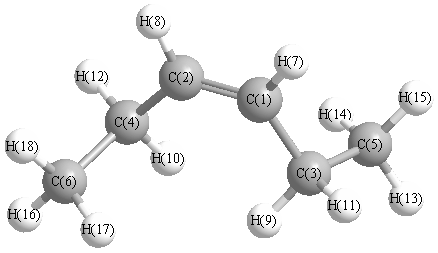
picture of (Z)-hex-3-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 127.714 C1 C2 H8 117.362
C1 C3 C5 111.957 C1 C3 H9 110.953
C1 C3 H11 108.669 C2 C1 C3 127.714
C2 C1 H7 117.362 C2 C4 C6 111.957
C2 C4 H10 110.953 C2 C4 H12 108.669
C3 C1 H7 114.911 C3 C5 H13 111.222
C3 C5 H14 110.698 C3 C5 H15 110.887
C4 C2 H8 114.911 C4 C6 H16 111.222
C4 C6 H17 110.698 C4 C6 H18 110.887
C5 C3 H9 109.708 C5 C3 H11 108.977
C6 C4 H10 109.708 C6 C4 H12 108.977
H9 C3 H11 106.397 H10 C4 H12 106.397
H13 C5 H14 108.281 H13 C5 H15 107.904
H14 C5 H15 107.717 H16 C6 H17 108.281
H16 C6 H18 107.904 H17 C6 H18 107.717
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability