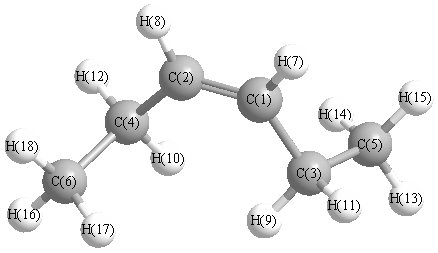
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3069 |
3051 |
50.30 |
|
|
|
| 2 |
A |
3054 |
3036 |
4.53 |
|
|
|
| 3 |
A |
3048 |
3030 |
53.55 |
|
|
|
| 4 |
A |
3011 |
2994 |
7.18 |
|
|
|
| 5 |
A |
2967 |
2950 |
0.33 |
|
|
|
| 6 |
A |
2947 |
2929 |
6.74 |
|
|
|
| 7 |
A |
1686 |
1677 |
1.44 |
|
|
|
| 8 |
A |
1430 |
1422 |
7.88 |
|
|
|
| 9 |
A |
1418 |
1409 |
3.21 |
|
|
|
| 10 |
A |
1407 |
1399 |
0.34 |
|
|
|
| 11 |
A |
1340 |
1332 |
0.16 |
|
|
|
| 12 |
A |
1284 |
1277 |
0.68 |
|
|
|
| 13 |
A |
1252 |
1245 |
0.07 |
|
|
|
| 14 |
A |
1221 |
1214 |
0.12 |
|
|
|
| 15 |
A |
1049 |
1043 |
0.27 |
|
|
|
| 16 |
A |
1030 |
1024 |
1.33 |
|
|
|
| 17 |
A |
1003 |
998 |
0.10 |
|
|
|
| 18 |
A |
966 |
960 |
0.86 |
|
|
|
| 19 |
A |
832 |
827 |
1.94 |
|
|
|
| 20 |
A |
757 |
753 |
0.65 |
|
|
|
| 21 |
A |
499 |
496 |
0.37 |
|
|
|
| 22 |
A |
298 |
297 |
0.00 |
|
|
|
| 23 |
A |
199 |
198 |
0.00 |
|
|
|
| 24 |
A |
190 |
189 |
0.05 |
|
|
|
| 25 |
A |
58 |
57 |
0.01 |
|
|
|
| 26 |
B |
3054 |
3037 |
59.63 |
|
|
|
| 27 |
B |
3048 |
3031 |
5.41 |
|
|
|
| 28 |
B |
3043 |
3026 |
3.90 |
|
|
|
| 29 |
B |
3006 |
2989 |
6.96 |
|
|
|
| 30 |
B |
2966 |
2949 |
73.87 |
|
|
|
| 31 |
B |
2946 |
2929 |
44.95 |
|
|
|
| 32 |
B |
1430 |
1422 |
3.20 |
|
|
|
| 33 |
B |
1418 |
1410 |
8.12 |
|
|
|
| 34 |
B |
1399 |
1391 |
0.35 |
|
|
|
| 35 |
B |
1380 |
1372 |
3.16 |
|
|
|
| 36 |
B |
1339 |
1331 |
1.23 |
|
|
|
| 37 |
B |
1266 |
1258 |
14.61 |
|
|
|
| 38 |
B |
1233 |
1225 |
0.53 |
|
|
|
| 39 |
B |
1136 |
1129 |
0.03 |
|
|
|
| 40 |
B |
1052 |
1045 |
5.77 |
|
|
|
| 41 |
B |
1026 |
1020 |
3.67 |
|
|
|
| 42 |
B |
890 |
885 |
8.83 |
|
|
|
| 43 |
B |
786 |
781 |
5.26 |
|
|
|
| 44 |
B |
732 |
727 |
26.04 |
|
|
|
| 45 |
B |
555 |
551 |
6.52 |
|
|
|
| 46 |
B |
346 |
344 |
0.76 |
|
|
|
| 47 |
B |
235 |
234 |
0.03 |
|
|
|
| 48 |
B |
48 |
48 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35173.3 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 34969.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.045 |
|
|
|
| 2 |
C |
-0.045 |
|
|
|
| 3 |
C |
-0.026 |
|
|
|
| 4 |
C |
-0.026 |
|
|
|
| 5 |
C |
-0.062 |
|
|
|
| 6 |
C |
-0.062 |
|
|
|
| 7 |
H |
-0.015 |
|
|
|
| 8 |
H |
-0.015 |
|
|
|
| 9 |
H |
0.017 |
|
|
|
| 10 |
H |
0.017 |
|
|
|
| 11 |
H |
0.035 |
|
|
|
| 12 |
H |
0.035 |
|
|
|
| 13 |
H |
0.026 |
|
|
|
| 14 |
H |
0.038 |
|
|
|
| 15 |
H |
0.032 |
|
|
|
| 16 |
H |
0.026 |
|
|
|
| 17 |
H |
0.038 |
|
|
|
| 18 |
H |
0.032 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.277 |
0.277 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.502 |
0.458 |
0.000 |
| y |
0.458 |
-38.484 |
0.000 |
| z |
0.000 |
0.000 |
-39.383 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.569 |
0.458 |
0.000 |
| y |
0.458 |
1.959 |
0.000 |
| z |
0.000 |
0.000 |
0.610 |
|
| Polar |
|---|
| 3z2-r2 | 1.220 |
|---|
| x2-y2 | -3.019 |
|---|
| xy | 0.458 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.481 |
-1.175 |
0.000 |
| y |
-1.175 |
14.091 |
0.000 |
| z |
0.000 |
0.000 |
9.597 |
<r2> (average value of r
2) Å
2
| <r2> |
261.314 |
| (<r2>)1/2 |
16.165 |