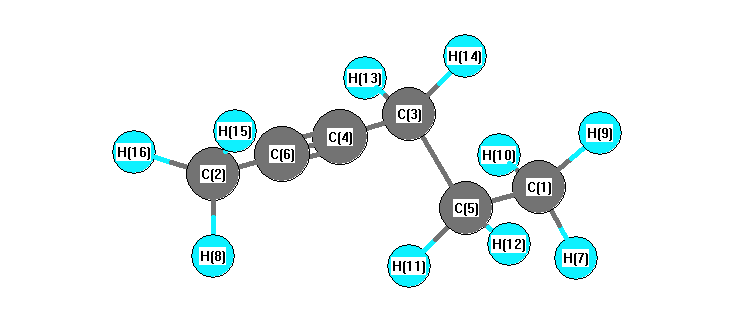
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3053 |
3035 |
27.27 |
|
|
|
| 2 |
A |
3029 |
3012 |
8.53 |
|
|
|
| 3 |
A |
2976 |
2959 |
26.00 |
|
|
|
| 4 |
A |
2965 |
2948 |
29.37 |
|
|
|
| 5 |
A |
2956 |
2939 |
44.49 |
|
|
|
| 6 |
A |
2935 |
2918 |
27.05 |
|
|
|
| 7 |
A |
2301 |
2288 |
0.03 |
|
|
|
| 8 |
A |
1431 |
1423 |
4.22 |
|
|
|
| 9 |
A |
1416 |
1408 |
0.45 |
|
|
|
| 10 |
A |
1397 |
1389 |
6.71 |
|
|
|
| 11 |
A |
1393 |
1385 |
1.47 |
|
|
|
| 12 |
A |
1351 |
1343 |
0.50 |
|
|
|
| 13 |
A |
1344 |
1336 |
6.28 |
|
|
|
| 14 |
A |
1326 |
1318 |
1.51 |
|
|
|
| 15 |
A |
1249 |
1242 |
12.77 |
|
|
|
| 16 |
A |
1161 |
1154 |
5.54 |
|
|
|
| 17 |
A |
1076 |
1069 |
1.86 |
|
|
|
| 18 |
A |
1039 |
1033 |
0.74 |
|
|
|
| 19 |
A |
996 |
990 |
1.50 |
|
|
|
| 20 |
A |
879 |
874 |
3.39 |
|
|
|
| 21 |
A |
761 |
756 |
0.24 |
|
|
|
| 22 |
A |
469 |
466 |
1.32 |
|
|
|
| 23 |
A |
328 |
326 |
0.21 |
|
|
|
| 24 |
A |
267 |
266 |
5.85 |
|
|
|
| 25 |
A |
98 |
98 |
1.63 |
|
|
|
| 26 |
A |
3044 |
3026 |
48.25 |
|
|
|
| 27 |
A |
3028 |
3011 |
8.66 |
|
|
|
| 28 |
A |
3014 |
2997 |
6.85 |
|
|
|
| 29 |
A |
2967 |
2950 |
8.63 |
|
|
|
| 30 |
A |
1421 |
1413 |
6.67 |
|
|
|
| 31 |
A |
1397 |
1389 |
6.60 |
|
|
|
| 32 |
A |
1268 |
1261 |
0.00 |
|
|
|
| 33 |
A |
1201 |
1194 |
0.05 |
|
|
|
| 34 |
A |
1070 |
1064 |
0.48 |
|
|
|
| 35 |
A |
996 |
990 |
1.66 |
|
|
|
| 36 |
A |
839 |
834 |
0.67 |
|
|
|
| 37 |
A |
726 |
721 |
3.48 |
|
|
|
| 38 |
A |
355 |
353 |
0.00 |
|
|
|
| 39 |
A |
246 |
245 |
0.01 |
|
|
|
| 40 |
A |
214 |
213 |
8.35 |
|
|
|
| 41 |
A |
85 |
84 |
0.42 |
|
|
|
| 42 |
A |
20 |
20 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30042.4 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 29868.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.045 |
|
|
|
| 2 |
C |
-0.010 |
|
|
|
| 3 |
C |
-0.054 |
|
|
|
| 4 |
C |
0.006 |
|
|
|
| 5 |
C |
-0.073 |
|
|
|
| 6 |
C |
-0.272 |
|
|
|
| 7 |
H |
0.030 |
|
|
|
| 8 |
H |
0.065 |
|
|
|
| 9 |
H |
0.031 |
|
|
|
| 10 |
H |
0.031 |
|
|
|
| 11 |
H |
0.032 |
|
|
|
| 12 |
H |
0.032 |
|
|
|
| 13 |
H |
0.048 |
|
|
|
| 14 |
H |
0.048 |
|
|
|
| 15 |
H |
0.065 |
|
|
|
| 16 |
H |
0.065 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.132 |
-0.068 |
0.000 |
0.149 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.208 |
-3.141 |
0.000 |
| y |
-3.141 |
-34.802 |
0.000 |
| z |
0.000 |
0.000 |
-38.538 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.462 |
-3.141 |
0.000 |
| y |
-3.141 |
2.571 |
0.000 |
| z |
0.000 |
0.000 |
-3.033 |
|
| Polar |
|---|
| 3z2-r2 | -6.066 |
|---|
| x2-y2 | -1.405 |
|---|
| xy | -3.141 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.291 |
-3.584 |
0.000 |
| y |
-3.584 |
13.273 |
0.000 |
| z |
0.000 |
0.000 |
7.425 |
<r2> (average value of r
2) Å
2
| <r2> |
297.411 |
| (<r2>)1/2 |
17.246 |