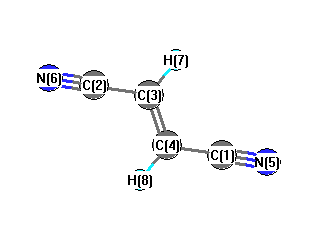
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3117 |
3099 |
0.00 |
|
|
|
| 2 |
Ag |
2237 |
2224 |
0.00 |
|
|
|
| 3 |
Ag |
1616 |
1607 |
0.00 |
|
|
|
| 4 |
Ag |
1256 |
1249 |
0.00 |
|
|
|
| 5 |
Ag |
1001 |
996 |
0.00 |
|
|
|
| 6 |
Ag |
525 |
522 |
0.00 |
|
|
|
| 7 |
Ag |
247 |
246 |
0.00 |
|
|
|
| 8 |
Au |
930 |
925 |
41.82 |
|
|
|
| 9 |
Au |
544 |
541 |
1.70 |
|
|
|
| 10 |
Au |
122 |
121 |
12.62 |
|
|
|
| 11 |
Bg |
820 |
815 |
0.00 |
|
|
|
| 12 |
Bg |
376 |
373 |
0.00 |
|
|
|
| 13 |
Bu |
3124 |
3106 |
5.36 |
|
|
|
| 14 |
Bu |
2258 |
2245 |
0.55 |
|
|
|
| 15 |
Bu |
1219 |
1212 |
1.73 |
|
|
|
| 16 |
Bu |
1024 |
1018 |
7.65 |
|
|
|
| 17 |
Bu |
520 |
517 |
0.98 |
|
|
|
| 18 |
Bu |
131 |
130 |
14.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10533.6 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 10472.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.047 |
|
|
|
| 2 |
C |
0.047 |
|
|
|
| 3 |
C |
-0.026 |
|
|
|
| 4 |
C |
-0.026 |
|
|
|
| 5 |
N |
-0.088 |
|
|
|
| 6 |
N |
-0.088 |
|
|
|
| 7 |
H |
0.067 |
|
|
|
| 8 |
H |
0.067 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.195 |
-7.821 |
0.000 |
| y |
-7.821 |
-51.091 |
0.000 |
| z |
0.000 |
0.000 |
-33.681 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
10.191 |
-7.821 |
0.000 |
| y |
-7.821 |
-18.153 |
0.000 |
| z |
0.000 |
0.000 |
7.961 |
|
| Polar |
|---|
| 3z2-r2 | 15.923 |
|---|
| x2-y2 | 18.896 |
|---|
| xy | -7.821 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.970 |
2.253 |
0.000 |
| y |
2.253 |
14.800 |
0.000 |
| z |
0.000 |
0.000 |
3.372 |
<r2> (average value of r
2) Å
2
| <r2> |
205.496 |
| (<r2>)1/2 |
14.335 |