return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8O (Methyl cyclopropyl ketone)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-269.768348
Energy at 298.15K-269.777059
HF Energy-268.853013
Nuclear repulsion energy229.272875
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3285 3142 5.45      
2 A' 3211 3072 7.75      
3 A' 3197 3058 9.06      
4 A' 3179 3042 7.87      
5 A' 3070 2937 4.34      
6 A' 1859 1778 108.62      
7 A' 1519 1453 19.23      
8 A' 1469 1405 11.94      
9 A' 1443 1380 91.48      
10 A' 1389 1329 6.29      
11 A' 1238 1185 2.23      
12 A' 1211 1159 77.91      
13 A' 1120 1071 10.01      
14 A' 1079 1032 7.59      
15 A' 990 947 34.69      
16 A' 945 905 9.56      
17 A' 811 776 1.21      
18 A' 761 728 0.25      
19 A' 599 573 14.10      
20 A' 375 359 3.82      
21 A' 243 233 3.71      
22 A" 3272 3130 0.44      
23 A" 3174 3036 10.81      
24 A" 3151 3014 9.07      
25 A" 1480 1416 8.27      
26 A" 1456 1393 2.58      
27 A" 1214 1161 0.27      
28 A" 1135 1086 1.69      
29 A" 1102 1054 1.46      
30 A" 1061 1015 2.24      
31 A" 911 871 1.38      
32 A" 844 807 4.66      
33 A" 607 580 1.42      
34 A" 266 255 0.36      
35 A" 156 150 0.05      
36 A" 67 64 1.51      

Unscaled Zero Point Vibrational Energy (zpe) 26442.1 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 25297.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.23822 0.08686 0.07603

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 -1.482 -0.555 0.000
C2 -0.284 -0.762 0.000
C3 0.299 -2.166 0.000
C4 0.704 0.372 0.000
C5 0.299 1.637 0.751
C6 0.299 1.637 -0.751
H7 -0.512 -2.911 0.000
H8 0.939 -2.306 -0.890
H9 0.939 -2.306 0.890
H10 1.765 0.101 0.000
H11 -0.674 1.597 1.252
H12 1.089 2.183 1.275
H13 -0.674 1.597 -1.252
H14 1.089 2.183 -1.275

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
O11.21562.40122.37482.92212.92212.54753.11723.11723.31312.61773.96652.61773.9665
C21.21561.51991.50452.58042.58042.16032.16102.16102.22412.69913.49072.69913.4907
C32.40121.51992.56993.87603.87601.10051.10541.10542.70044.08314.60014.08314.6001
C42.37481.50452.56991.52591.52593.50032.83132.83131.09482.22872.24792.22872.2479
C52.92212.58043.87601.52591.50184.67954.31803.99642.25211.09441.09442.22662.2421
C62.92212.58043.87601.52591.50184.67953.99644.31802.25212.22662.24211.09441.0944
H72.54752.16031.10053.50034.67954.67951.80611.80613.77584.68105.48904.68105.4890
H83.11722.16101.10542.83134.31803.99641.80611.78072.69634.73494.98564.23804.5073
H93.11722.16101.10542.83133.99644.31801.80611.78072.69634.23804.50734.73494.9856
H103.31312.22412.70041.09482.25212.25213.77582.69632.69633.12282.53253.12282.5325
H112.61772.69914.08312.22871.09442.22664.68104.73494.23803.12281.85772.50363.1361
H123.96653.49074.60012.24791.09442.24215.48904.98564.50732.53251.85773.13612.5498
H132.61772.69914.08312.22872.22661.09444.68104.23804.73493.12282.50363.13611.8577
H143.96653.49074.60012.24792.24211.09445.48904.50734.98562.53253.13612.54981.8577

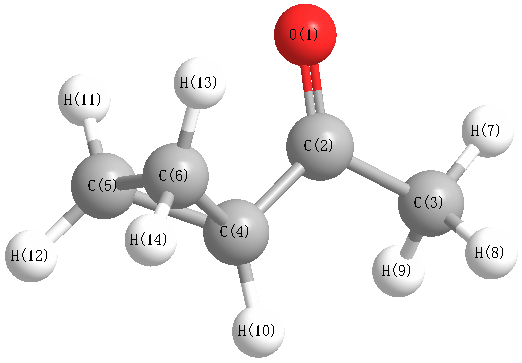
picture of Methyl cyclopropyl ketone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C3 122.364 O1 C2 C4 121.269
C2 C3 H7 110.032 C2 C3 H8 109.803
C2 C3 H9 109.803 C2 C4 C5 116.750
C2 C4 C6 116.750 C2 C4 H10 116.787
C3 C2 C4 116.367 C4 C5 C6 60.522
C4 C5 H11 115.558 C4 C5 H12 117.196
C4 C6 C5 60.522 C4 C6 H13 115.558
C4 C6 H14 117.196 C5 C4 C6 58.957
C5 C4 H10 117.538 C5 C6 H13 117.239
C5 C6 H14 118.605 C6 C4 H10 117.538
C6 C5 H11 117.239 C6 C5 H12 118.605
H7 C3 H8 109.924 H7 C3 H9 109.924
H8 C3 H9 107.313 H11 C5 H12 116.150
H13 C6 H14 116.150
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability