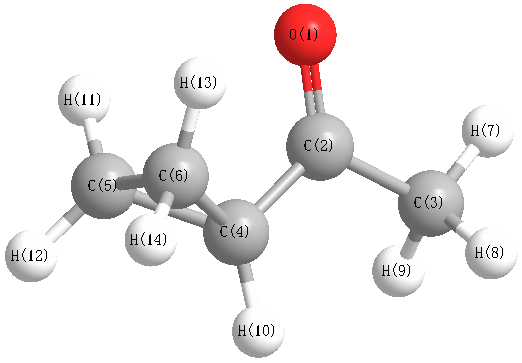
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3176 |
3158 |
3.39 |
|
|
|
| 2 |
A' |
3102 |
3084 |
11.16 |
|
|
|
| 3 |
A' |
3100 |
3082 |
6.48 |
|
|
|
| 4 |
A' |
3066 |
3048 |
9.55 |
|
|
|
| 5 |
A' |
2968 |
2951 |
2.68 |
|
|
|
| 6 |
A' |
1725 |
1715 |
112.86 |
|
|
|
| 7 |
A' |
1418 |
1409 |
20.38 |
|
|
|
| 8 |
A' |
1382 |
1374 |
15.66 |
|
|
|
| 9 |
A' |
1354 |
1346 |
109.18 |
|
|
|
| 10 |
A' |
1300 |
1292 |
14.51 |
|
|
|
| 11 |
A' |
1193 |
1186 |
1.36 |
|
|
|
| 12 |
A' |
1149 |
1142 |
81.98 |
|
|
|
| 13 |
A' |
1065 |
1059 |
15.46 |
|
|
|
| 14 |
A' |
1007 |
1001 |
9.40 |
|
|
|
| 15 |
A' |
933 |
928 |
54.10 |
|
|
|
| 16 |
A' |
911 |
905 |
0.69 |
|
|
|
| 17 |
A' |
777 |
773 |
4.42 |
|
|
|
| 18 |
A' |
733 |
729 |
0.18 |
|
|
|
| 19 |
A' |
575 |
571 |
8.11 |
|
|
|
| 20 |
A' |
365 |
363 |
4.38 |
|
|
|
| 21 |
A' |
236 |
234 |
3.90 |
|
|
|
| 22 |
A" |
3162 |
3144 |
0.51 |
|
|
|
| 23 |
A" |
3065 |
3047 |
12.47 |
|
|
|
| 24 |
A" |
3041 |
3024 |
5.21 |
|
|
|
| 25 |
A" |
1394 |
1385 |
8.99 |
|
|
|
| 26 |
A" |
1365 |
1357 |
4.65 |
|
|
|
| 27 |
A" |
1156 |
1150 |
0.16 |
|
|
|
| 28 |
A" |
1070 |
1064 |
1.59 |
|
|
|
| 29 |
A" |
1031 |
1025 |
0.92 |
|
|
|
| 30 |
A" |
994 |
988 |
4.88 |
|
|
|
| 31 |
A" |
843 |
838 |
0.31 |
|
|
|
| 32 |
A" |
808 |
803 |
3.55 |
|
|
|
| 33 |
A" |
580 |
577 |
0.36 |
|
|
|
| 34 |
A" |
245 |
244 |
0.16 |
|
|
|
| 35 |
A" |
138 |
137 |
0.34 |
|
|
|
| 36 |
A" |
67 |
66 |
1.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25244.9 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 25098.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.201 |
|
|
|
| 2 |
C |
0.052 |
|
|
|
| 3 |
C |
-0.032 |
|
|
|
| 4 |
C |
-0.135 |
|
|
|
| 5 |
C |
0.000 |
|
|
|
| 6 |
C |
0.000 |
|
|
|
| 7 |
H |
0.044 |
|
|
|
| 8 |
H |
0.059 |
|
|
|
| 9 |
H |
0.059 |
|
|
|
| 10 |
H |
-0.014 |
|
|
|
| 11 |
H |
0.048 |
|
|
|
| 12 |
H |
0.037 |
|
|
|
| 13 |
H |
0.048 |
|
|
|
| 14 |
H |
0.037 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.575 |
-0.006 |
0.000 |
2.575 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.077 |
-0.798 |
0.000 |
| y |
-0.798 |
-34.314 |
0.000 |
| z |
0.000 |
0.000 |
-36.076 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.881 |
-0.798 |
0.000 |
| y |
-0.798 |
3.262 |
0.000 |
| z |
0.000 |
0.000 |
0.619 |
|
| Polar |
|---|
| 3z2-r2 | 1.238 |
|---|
| x2-y2 | -4.762 |
|---|
| xy | -0.798 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.188 |
0.180 |
0.000 |
| y |
0.180 |
10.318 |
0.000 |
| z |
0.000 |
0.000 |
6.979 |
<r2> (average value of r
2) Å
2
| <r2> |
169.623 |
| (<r2>)1/2 |
13.024 |