Jump to
S1C2
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Geometric Data calculated at QCISD/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -235.125780 |
| Energy at 298.15K | -235.138001 |
| HF Energy | -234.193294 |
| Nuclear repulsion energy | 228.567129 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3177 |
3048 |
55.08 |
|
|
|
| 2 |
A |
3162 |
3033 |
8.47 |
|
|
|
| 3 |
A |
3147 |
3019 |
11.76 |
|
|
|
| 4 |
A |
3136 |
3008 |
36.84 |
|
|
|
| 5 |
A |
3131 |
3004 |
56.39 |
|
|
|
| 6 |
A |
3116 |
2990 |
21.87 |
|
|
|
| 7 |
A |
3111 |
2985 |
16.01 |
|
|
|
| 8 |
A |
3087 |
2962 |
6.29 |
|
|
|
| 9 |
A |
3054 |
2930 |
25.54 |
|
|
|
| 10 |
A |
3051 |
2928 |
14.91 |
|
|
|
| 11 |
A |
3048 |
2924 |
61.94 |
|
|
|
| 12 |
A |
3041 |
2918 |
13.76 |
|
|
|
| 13 |
A |
1754 |
1683 |
1.54 |
|
|
|
| 14 |
A |
1507 |
1446 |
3.94 |
|
|
|
| 15 |
A |
1496 |
1436 |
5.83 |
|
|
|
| 16 |
A |
1495 |
1435 |
3.28 |
|
|
|
| 17 |
A |
1490 |
1430 |
0.47 |
|
|
|
| 18 |
A |
1484 |
1424 |
3.24 |
|
|
|
| 19 |
A |
1482 |
1422 |
1.10 |
|
|
|
| 20 |
A |
1449 |
1390 |
6.22 |
|
|
|
| 21 |
A |
1423 |
1366 |
1.62 |
|
|
|
| 22 |
A |
1410 |
1353 |
2.13 |
|
|
|
| 23 |
A |
1389 |
1333 |
0.79 |
|
|
|
| 24 |
A |
1327 |
1273 |
0.06 |
|
|
|
| 25 |
A |
1298 |
1245 |
6.47 |
|
|
|
| 26 |
A |
1293 |
1241 |
1.82 |
|
|
|
| 27 |
A |
1257 |
1206 |
0.07 |
|
|
|
| 28 |
A |
1176 |
1128 |
0.14 |
|
|
|
| 29 |
A |
1127 |
1081 |
0.69 |
|
|
|
| 30 |
A |
1084 |
1040 |
1.28 |
|
|
|
| 31 |
A |
1065 |
1022 |
0.96 |
|
|
|
| 32 |
A |
1061 |
1018 |
1.45 |
|
|
|
| 33 |
A |
1017 |
976 |
3.78 |
|
|
|
| 34 |
A |
999 |
958 |
0.28 |
|
|
|
| 35 |
A |
929 |
891 |
2.66 |
|
|
|
| 36 |
A |
908 |
872 |
3.54 |
|
|
|
| 37 |
A |
874 |
838 |
2.78 |
|
|
|
| 38 |
A |
750 |
720 |
1.17 |
|
|
|
| 39 |
A |
722 |
693 |
26.17 |
|
|
|
| 40 |
A |
564 |
541 |
5.52 |
|
|
|
| 41 |
A |
470 |
451 |
0.68 |
|
|
|
| 42 |
A |
364 |
349 |
0.00 |
|
|
|
| 43 |
A |
289 |
277 |
0.04 |
|
|
|
| 44 |
A |
249 |
239 |
0.00 |
|
|
|
| 45 |
A |
199 |
191 |
0.01 |
|
|
|
| 46 |
A |
117 |
112 |
0.24 |
|
|
|
| 47 |
A |
92 |
88 |
0.00 |
|
|
|
| 48 |
A |
40 |
38 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36456.0 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 34975.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.472 |
0.967 |
-0.063 |
| C2 |
2.006 |
-0.469 |
-0.150 |
| C3 |
0.776 |
-0.954 |
0.126 |
| C4 |
-0.446 |
-0.180 |
0.571 |
| C5 |
-1.578 |
-0.222 |
-0.473 |
| C6 |
-2.843 |
0.508 |
-0.003 |
| H7 |
1.668 |
1.659 |
0.235 |
| H8 |
3.297 |
1.065 |
0.669 |
| H9 |
2.868 |
1.304 |
-1.040 |
| H10 |
2.771 |
-1.191 |
-0.473 |
| H11 |
0.620 |
-2.036 |
-0.001 |
| H12 |
-0.191 |
0.873 |
0.793 |
| H13 |
-0.822 |
-0.616 |
1.520 |
| H14 |
-1.212 |
0.227 |
-1.417 |
| H15 |
-1.820 |
-1.278 |
-0.704 |
| H16 |
-3.639 |
0.467 |
-0.768 |
| H17 |
-3.240 |
0.054 |
0.925 |
| H18 |
-2.631 |
1.573 |
0.209 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.5118 | 2.5696 | 3.1986 | 4.2405 | 5.3355 | 1.1022 | 1.1067 | 1.1065 | 2.2165 | 3.5291 | 2.7993 | 3.9824 | 3.9941 | 4.8862 | 6.1726 | 5.8682 | 5.1460 |
C2 | 1.5118 | | 1.3513 | 2.5722 | 3.6071 | 4.9490 | 2.1885 | 2.1651 | 2.1629 | 1.1004 | 2.0981 | 2.7421 | 3.2873 | 3.5282 | 3.9502 | 5.7562 | 5.3805 | 5.0792 | C3 | 2.5696 | 1.3513 | | 1.5130 | 2.5364 | 3.9049 | 2.7627 | 3.2748 | 3.2918 | 2.0972 | 1.1011 | 2.1717 | 2.1463 | 2.7799 | 2.7447 | 4.7236 | 4.2162 | 4.2418 | C4 | 3.1986 | 2.5722 | 1.5130 | | 1.5406 | 2.5593 | 2.8211 | 3.9449 | 3.9721 | 3.5302 | 2.2158 | 1.1052 | 1.1095 | 2.1696 | 2.1733 | 3.5231 | 2.8260 | 2.8242 | C5 | 4.2405 | 3.6071 | 2.5364 | 1.5406 | | 1.5342 | 3.8168 | 5.1687 | 4.7341 | 4.4556 | 2.8885 | 2.1728 | 2.1673 | 1.1077 | 1.1084 | 2.1937 | 2.1890 | 2.1890 | C6 | 5.3355 | 4.9490 | 3.9049 | 2.5593 | 1.5342 | | 4.6614 | 6.2013 | 5.8587 | 5.8844 | 4.2967 | 2.7925 | 2.7690 | 2.1771 | 2.1741 | 1.1050 | 1.1063 | 1.1064 | H7 | 1.1022 | 2.1885 | 2.7627 | 2.8211 | 3.8168 | 4.6614 | | 1.7871 | 1.7863 | 3.1367 | 3.8480 | 2.0940 | 3.6086 | 3.6152 | 4.6552 | 5.5310 | 5.2090 | 4.2993 | H8 | 1.1067 | 2.1651 | 3.2748 | 3.9449 | 5.1687 | 6.2013 | 1.7871 | | 1.7779 | 2.5826 | 4.1512 | 3.4953 | 4.5286 | 5.0377 | 5.7927 | 7.1085 | 6.6188 | 5.9666 | H9 | 1.1065 | 2.1629 | 3.2918 | 3.9721 | 4.7341 | 5.8587 | 1.7863 | 1.7779 | | 2.5603 | 4.1584 | 3.5921 | 4.8837 | 4.2363 | 5.3626 | 6.5667 | 6.5365 | 5.6449 | H10 | 2.2165 | 1.1004 | 2.0972 | 3.5302 | 4.4556 | 5.8844 | 3.1367 | 2.5826 | 2.5603 | | 2.3596 | 3.8261 | 4.1487 | 4.3319 | 4.5979 | 6.6281 | 6.2957 | 6.1059 | H11 | 3.5291 | 2.0981 | 1.1011 | 2.2158 | 2.8885 | 4.2967 | 3.8480 | 4.1512 | 4.1584 | 2.3596 | | 3.1227 | 2.5312 | 3.2377 | 2.6498 | 4.9993 | 4.4856 | 4.8613 | H12 | 2.7993 | 2.7421 | 2.1717 | 1.1052 | 2.1728 | 2.7925 | 2.0940 | 3.4953 | 3.5921 | 3.8261 | 3.1227 | | 1.7727 | 2.5183 | 3.0855 | 3.8066 | 3.1592 | 2.6040 | H13 | 3.9824 | 3.2873 | 2.1463 | 1.1095 | 2.1673 | 2.7690 | 3.6086 | 4.5286 | 4.8837 | 4.1487 | 2.5312 | 1.7727 | | 3.0803 | 2.5261 | 3.7876 | 2.5787 | 3.1275 | H14 | 3.9941 | 3.5282 | 2.7799 | 2.1696 | 1.1077 | 2.1771 | 3.6152 | 5.0377 | 4.2363 | 4.3319 | 3.2377 | 2.5183 | 3.0803 | | 1.7731 | 2.5244 | 3.1028 | 2.5429 | H15 | 4.8862 | 3.9502 | 2.7447 | 2.1733 | 1.1084 | 2.1741 | 4.6552 | 5.7927 | 5.3626 | 4.5979 | 2.6498 | 3.0855 | 2.5261 | 1.7731 | | 2.5217 | 2.5384 | 3.1010 | H16 | 6.1726 | 5.7562 | 4.7236 | 3.5231 | 2.1937 | 1.1050 | 5.5310 | 7.1085 | 6.5667 | 6.6281 | 4.9993 | 3.8066 | 3.7876 | 2.5244 | 2.5217 | | 1.7877 | 1.7872 | H17 | 5.8682 | 5.3805 | 4.2162 | 2.8260 | 2.1890 | 1.1063 | 5.2090 | 6.6188 | 6.5365 | 6.2957 | 4.4856 | 3.1592 | 2.5787 | 3.1028 | 2.5384 | 1.7877 | | 1.7861 | H18 | 5.1460 | 5.0792 | 4.2418 | 2.8242 | 2.1890 | 1.1064 | 4.2993 | 5.9666 | 5.6449 | 6.1059 | 4.8613 | 2.6040 | 3.1275 | 2.5429 | 3.1010 | 1.7872 | 1.7861 | |
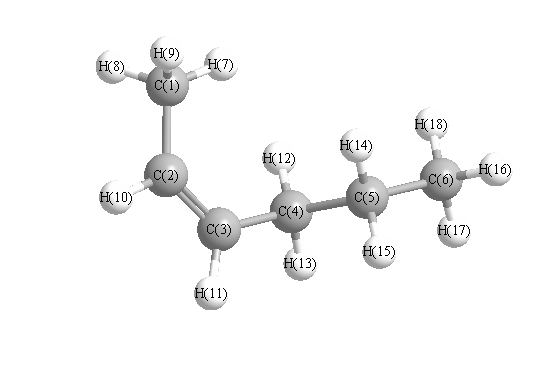 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.565 |
|
C1 |
C2 |
H10 |
115.192 |
| C2 |
C1 |
H7 |
112.750 |
|
C2 |
C1 |
H8 |
110.595 |
| C2 |
C1 |
H9 |
110.436 |
|
C2 |
C3 |
C4 |
127.714 |
| C2 |
C3 |
H11 |
117.273 |
|
C3 |
C2 |
H10 |
117.242 |
| C3 |
C4 |
C5 |
112.326 |
|
C3 |
C4 |
H12 |
111.130 |
| C3 |
C4 |
H13 |
108.886 |
|
C4 |
C3 |
H11 |
115.001 |
| C4 |
C5 |
C6 |
112.682 |
|
C4 |
C5 |
H14 |
108.916 |
| C4 |
C5 |
H15 |
109.158 |
|
C5 |
C4 |
H12 |
109.307 |
| C5 |
C4 |
H13 |
108.638 |
|
C5 |
C6 |
H16 |
111.408 |
| C5 |
C6 |
H17 |
110.956 |
|
C5 |
C6 |
H18 |
110.957 |
| C6 |
C5 |
H14 |
109.936 |
|
C6 |
C5 |
H15 |
109.665 |
| H7 |
C1 |
H8 |
108.003 |
|
H7 |
C1 |
H9 |
107.945 |
| H8 |
C1 |
H9 |
106.889 |
|
H12 |
C4 |
H13 |
106.345 |
| H14 |
C5 |
H16 |
93.934 |
|
H16 |
C6 |
H17 |
107.881 |
| H16 |
C6 |
H18 |
107.838 |
|
H17 |
C6 |
H18 |
107.641 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
