Jump to
S1C2
Vibrational Frequencies calculated at CCD/cc-pVDZ
Geometric Data calculated at CCD/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -235.119908 |
| Energy at 298.15K | -235.132144 |
| HF Energy | -234.193543 |
| Nuclear repulsion energy | 228.699399 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3187 |
3049 |
54.23 |
|
|
|
| 2 |
A |
3173 |
3035 |
6.76 |
|
|
|
| 3 |
A |
3156 |
3020 |
12.22 |
|
|
|
| 4 |
A |
3149 |
3012 |
34.44 |
|
|
|
| 5 |
A |
3143 |
3007 |
51.71 |
|
|
|
| 6 |
A |
3128 |
2993 |
20.58 |
|
|
|
| 7 |
A |
3123 |
2988 |
16.62 |
|
|
|
| 8 |
A |
3100 |
2966 |
5.48 |
|
|
|
| 9 |
A |
3063 |
2931 |
24.78 |
|
|
|
| 10 |
A |
3062 |
2929 |
14.33 |
|
|
|
| 11 |
A |
3059 |
2926 |
58.43 |
|
|
|
| 12 |
A |
3052 |
2920 |
12.74 |
|
|
|
| 13 |
A |
1768 |
1692 |
1.59 |
|
|
|
| 14 |
A |
1509 |
1444 |
4.28 |
|
|
|
| 15 |
A |
1499 |
1434 |
6.03 |
|
|
|
| 16 |
A |
1498 |
1433 |
3.42 |
|
|
|
| 17 |
A |
1493 |
1428 |
0.47 |
|
|
|
| 18 |
A |
1486 |
1422 |
2.67 |
|
|
|
| 19 |
A |
1485 |
1421 |
1.82 |
|
|
|
| 20 |
A |
1453 |
1390 |
6.56 |
|
|
|
| 21 |
A |
1427 |
1365 |
1.77 |
|
|
|
| 22 |
A |
1414 |
1352 |
1.73 |
|
|
|
| 23 |
A |
1393 |
1332 |
0.64 |
|
|
|
| 24 |
A |
1330 |
1272 |
0.06 |
|
|
|
| 25 |
A |
1301 |
1245 |
5.42 |
|
|
|
| 26 |
A |
1297 |
1241 |
1.95 |
|
|
|
| 27 |
A |
1260 |
1205 |
0.07 |
|
|
|
| 28 |
A |
1178 |
1127 |
0.16 |
|
|
|
| 29 |
A |
1130 |
1081 |
0.67 |
|
|
|
| 30 |
A |
1087 |
1040 |
1.18 |
|
|
|
| 31 |
A |
1068 |
1021 |
0.96 |
|
|
|
| 32 |
A |
1065 |
1019 |
1.39 |
|
|
|
| 33 |
A |
1020 |
976 |
3.58 |
|
|
|
| 34 |
A |
1005 |
961 |
0.35 |
|
|
|
| 35 |
A |
932 |
892 |
2.49 |
|
|
|
| 36 |
A |
911 |
871 |
3.73 |
|
|
|
| 37 |
A |
876 |
838 |
2.78 |
|
|
|
| 38 |
A |
752 |
719 |
1.27 |
|
|
|
| 39 |
A |
725 |
694 |
26.36 |
|
|
|
| 40 |
A |
566 |
541 |
5.46 |
|
|
|
| 41 |
A |
473 |
452 |
0.68 |
|
|
|
| 42 |
A |
365 |
349 |
0.00 |
|
|
|
| 43 |
A |
290 |
278 |
0.04 |
|
|
|
| 44 |
A |
249 |
239 |
0.00 |
|
|
|
| 45 |
A |
200 |
191 |
0.01 |
|
|
|
| 46 |
A |
117 |
112 |
0.24 |
|
|
|
| 47 |
A |
92 |
88 |
0.00 |
|
|
|
| 48 |
A |
40 |
38 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36572.6 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 34989.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.468 |
0.968 |
-0.061 |
| C2 |
2.004 |
-0.468 |
-0.151 |
| C3 |
0.777 |
-0.955 |
0.125 |
| C4 |
-0.445 |
-0.185 |
0.573 |
| C5 |
-1.574 |
-0.218 |
-0.475 |
| C6 |
-2.841 |
0.507 |
-0.002 |
| H7 |
1.663 |
1.658 |
0.239 |
| H8 |
3.293 |
1.066 |
0.670 |
| H9 |
2.862 |
1.308 |
-1.037 |
| H10 |
2.770 |
-1.187 |
-0.477 |
| H11 |
0.623 |
-2.037 |
-0.006 |
| H12 |
-0.192 |
0.866 |
0.803 |
| H13 |
-0.823 |
-0.628 |
1.516 |
| H14 |
-1.206 |
0.239 |
-1.414 |
| H15 |
-1.816 |
-1.272 |
-0.715 |
| H16 |
-3.635 |
0.472 |
-0.769 |
| H17 |
-3.240 |
0.046 |
0.920 |
| H18 |
-2.630 |
1.570 |
0.219 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.5114 | 2.5674 | 3.1965 | 4.2330 | 5.3295 | 1.1018 | 1.1063 | 1.1061 | 2.2153 | 3.5266 | 2.7988 | 3.9837 | 3.9827 | 4.8782 | 6.1640 | 5.8649 | 5.1408 |
C2 | 1.5114 | | 1.3489 | 2.5698 | 3.6020 | 4.9447 | 2.1879 | 2.1644 | 2.1622 | 1.1000 | 2.0957 | 2.7407 | 3.2865 | 3.5216 | 3.9445 | 5.7504 | 5.3772 | 5.0756 | C3 | 2.5674 | 1.3489 | | 1.5125 | 2.5360 | 3.9043 | 2.7615 | 3.2722 | 3.2892 | 2.0948 | 1.1007 | 2.1710 | 2.1457 | 2.7793 | 2.7440 | 4.7225 | 4.2155 | 4.2411 | C4 | 3.1965 | 2.5698 | 1.5125 | | 1.5404 | 2.5591 | 2.8201 | 3.9425 | 3.9697 | 3.5276 | 2.2149 | 1.1047 | 1.1090 | 2.1690 | 2.1727 | 3.5224 | 2.8258 | 2.8239 | C5 | 4.2330 | 3.6020 | 2.5360 | 1.5404 | | 1.5339 | 3.8094 | 5.1618 | 4.7253 | 4.4511 | 2.8907 | 2.1721 | 2.1668 | 1.1072 | 1.1079 | 2.1930 | 2.1884 | 2.1885 | C6 | 5.3295 | 4.9447 | 3.9043 | 2.5591 | 1.5339 | | 4.6555 | 6.1956 | 5.8515 | 5.8803 | 4.2975 | 2.7921 | 2.7686 | 2.1765 | 2.1735 | 1.1045 | 1.1058 | 1.1059 | H7 | 1.1018 | 2.1879 | 2.7615 | 2.8201 | 3.8094 | 4.6555 | | 1.7863 | 1.7854 | 3.1355 | 3.8464 | 2.0948 | 3.6115 | 3.6029 | 4.6477 | 5.5222 | 5.2065 | 4.2941 | H8 | 1.1063 | 2.1644 | 3.2722 | 3.9425 | 5.1618 | 6.1956 | 1.7863 | | 1.7771 | 2.5814 | 4.1484 | 3.4927 | 4.5307 | 5.0264 | 5.7862 | 7.1003 | 6.6165 | 5.9608 | H9 | 1.1061 | 2.1622 | 3.2892 | 3.9697 | 4.7253 | 5.8515 | 1.7854 | 1.7771 | | 2.5588 | 4.1555 | 3.5930 | 4.8839 | 4.2235 | 5.3520 | 6.5561 | 6.5316 | 5.6397 | H10 | 2.2153 | 1.1000 | 2.0948 | 3.5276 | 4.4511 | 5.8803 | 3.1355 | 2.5814 | 2.5588 | | 2.3569 | 3.8243 | 4.1471 | 4.3265 | 4.5927 | 6.6228 | 6.2922 | 6.1024 | H11 | 3.5266 | 2.0957 | 1.1007 | 2.2149 | 2.8907 | 4.2975 | 3.8464 | 4.1484 | 4.1555 | 2.3569 | | 3.1212 | 2.5281 | 3.2412 | 2.6525 | 5.0007 | 4.4849 | 4.8616 | H12 | 2.7988 | 2.7407 | 2.1710 | 1.1047 | 2.1721 | 2.7921 | 2.0948 | 3.4927 | 3.5930 | 3.8243 | 3.1212 | | 1.7717 | 2.5174 | 3.0844 | 3.8056 | 3.1587 | 2.6038 | H13 | 3.9837 | 3.2865 | 2.1457 | 1.1090 | 2.1668 | 2.7686 | 3.6115 | 4.5307 | 4.8839 | 4.1471 | 2.5281 | 1.7717 | | 3.0793 | 2.5258 | 3.7867 | 2.5786 | 3.1267 | H14 | 3.9827 | 3.5216 | 2.7793 | 2.1690 | 1.1072 | 2.1765 | 3.6029 | 5.0264 | 4.2235 | 4.3265 | 3.2412 | 2.5174 | 3.0793 | | 1.7722 | 2.5236 | 3.1018 | 2.5423 | H15 | 4.8782 | 3.9445 | 2.7440 | 2.1727 | 1.1079 | 2.1735 | 4.6477 | 5.7862 | 5.3520 | 4.5927 | 2.6525 | 3.0844 | 2.5258 | 1.7722 | | 2.5208 | 2.5377 | 3.0999 | H16 | 6.1640 | 5.7504 | 4.7225 | 3.5224 | 2.1930 | 1.1045 | 5.5222 | 7.1003 | 6.5561 | 6.6228 | 5.0007 | 3.8056 | 3.7867 | 2.5236 | 2.5208 | | 1.7868 | 1.7864 | H17 | 5.8649 | 5.3772 | 4.2155 | 2.8258 | 2.1884 | 1.1058 | 5.2065 | 6.6165 | 6.5316 | 6.2922 | 4.4849 | 3.1587 | 2.5786 | 3.1018 | 2.5377 | 1.7868 | | 1.7852 | H18 | 5.1408 | 5.0756 | 4.2411 | 2.8239 | 2.1885 | 1.1059 | 4.2941 | 5.9608 | 5.6397 | 6.1024 | 4.8616 | 2.6038 | 3.1267 | 2.5423 | 3.0999 | 1.7864 | 1.7852 | |
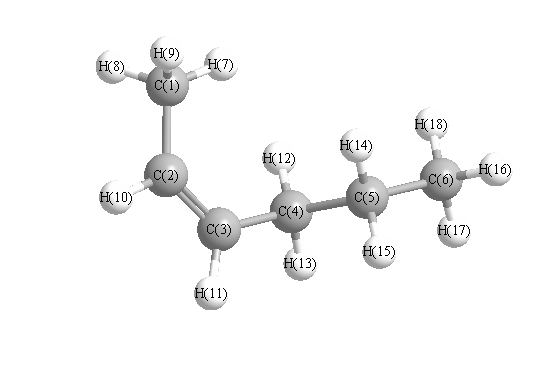 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.595 |
|
C1 |
C2 |
H10 |
115.158 |
| C2 |
C1 |
H7 |
112.770 |
|
C2 |
C1 |
H8 |
110.599 |
| C2 |
C1 |
H9 |
110.437 |
|
C2 |
C3 |
C4 |
127.725 |
| C2 |
C3 |
H11 |
117.276 |
|
C3 |
C2 |
H10 |
117.247 |
| C3 |
C4 |
C5 |
112.335 |
|
C3 |
C4 |
H12 |
111.138 |
| C3 |
C4 |
H13 |
108.894 |
|
C4 |
C3 |
H11 |
114.987 |
| C4 |
C5 |
C6 |
112.695 |
|
C4 |
C5 |
H14 |
108.909 |
| C4 |
C5 |
H15 |
109.159 |
|
C5 |
C4 |
H12 |
109.296 |
| C5 |
C4 |
H13 |
108.644 |
|
C5 |
C6 |
H16 |
111.404 |
| C5 |
C6 |
H17 |
110.959 |
|
C5 |
C6 |
H18 |
110.961 |
| C6 |
C5 |
H14 |
109.938 |
|
C6 |
C5 |
H15 |
109.665 |
| H7 |
C1 |
H8 |
107.996 |
|
H7 |
C1 |
H9 |
107.930 |
| H8 |
C1 |
H9 |
106.885 |
|
H12 |
C4 |
H13 |
106.324 |
| H14 |
C5 |
H16 |
93.936 |
|
H16 |
C6 |
H17 |
107.879 |
| H16 |
C6 |
H18 |
107.837 |
|
H17 |
C6 |
H18 |
107.641 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
