Jump to
S1C2
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
Geometric Data calculated at PBEPBE/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -235.511307 |
| Energy at 298.15K | -235.523416 |
| HF Energy | -235.511307 |
| Nuclear repulsion energy | 228.362968 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3080 |
3062 |
51.61 |
|
|
|
| 2 |
A |
3064 |
3046 |
8.75 |
|
|
|
| 3 |
A |
3050 |
3033 |
30.32 |
|
|
|
| 4 |
A |
3049 |
3032 |
9.66 |
|
|
|
| 5 |
A |
3040 |
3023 |
42.48 |
|
|
|
| 6 |
A |
3012 |
2994 |
16.25 |
|
|
|
| 7 |
A |
3007 |
2990 |
23.15 |
|
|
|
| 8 |
A |
2988 |
2970 |
6.58 |
|
|
|
| 9 |
A |
2963 |
2946 |
33.39 |
|
|
|
| 10 |
A |
2957 |
2940 |
26.83 |
|
|
|
| 11 |
A |
2956 |
2939 |
31.97 |
|
|
|
| 12 |
A |
2934 |
2916 |
22.50 |
|
|
|
| 13 |
A |
1692 |
1682 |
2.95 |
|
|
|
| 14 |
A |
1431 |
1423 |
4.32 |
|
|
|
| 15 |
A |
1421 |
1412 |
6.86 |
|
|
|
| 16 |
A |
1416 |
1408 |
2.16 |
|
|
|
| 17 |
A |
1413 |
1405 |
2.31 |
|
|
|
| 18 |
A |
1403 |
1395 |
1.39 |
|
|
|
| 19 |
A |
1401 |
1392 |
4.49 |
|
|
|
| 20 |
A |
1375 |
1367 |
12.07 |
|
|
|
| 21 |
A |
1348 |
1340 |
1.42 |
|
|
|
| 22 |
A |
1335 |
1327 |
2.77 |
|
|
|
| 23 |
A |
1316 |
1308 |
0.33 |
|
|
|
| 24 |
A |
1271 |
1263 |
0.04 |
|
|
|
| 25 |
A |
1247 |
1240 |
1.69 |
|
|
|
| 26 |
A |
1233 |
1226 |
5.81 |
|
|
|
| 27 |
A |
1200 |
1193 |
0.12 |
|
|
|
| 28 |
A |
1129 |
1122 |
0.37 |
|
|
|
| 29 |
A |
1076 |
1070 |
0.85 |
|
|
|
| 30 |
A |
1046 |
1040 |
2.03 |
|
|
|
| 31 |
A |
1024 |
1018 |
2.90 |
|
|
|
| 32 |
A |
1007 |
1001 |
0.42 |
|
|
|
| 33 |
A |
983 |
977 |
7.48 |
|
|
|
| 34 |
A |
966 |
960 |
0.30 |
|
|
|
| 35 |
A |
905 |
900 |
5.21 |
|
|
|
| 36 |
A |
877 |
871 |
6.12 |
|
|
|
| 37 |
A |
843 |
838 |
3.38 |
|
|
|
| 38 |
A |
732 |
728 |
2.12 |
|
|
|
| 39 |
A |
694 |
690 |
26.40 |
|
|
|
| 40 |
A |
556 |
553 |
8.41 |
|
|
|
| 41 |
A |
458 |
456 |
0.88 |
|
|
|
| 42 |
A |
355 |
353 |
0.03 |
|
|
|
| 43 |
A |
278 |
276 |
0.05 |
|
|
|
| 44 |
A |
246 |
245 |
0.00 |
|
|
|
| 45 |
A |
196 |
195 |
0.10 |
|
|
|
| 46 |
A |
140 |
140 |
0.36 |
|
|
|
| 47 |
A |
94 |
94 |
0.02 |
|
|
|
| 48 |
A |
55 |
55 |
0.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35129.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 34926.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.502 |
0.942 |
-0.082 |
| C2 |
2.012 |
-0.476 |
-0.135 |
| C3 |
0.771 |
-0.931 |
0.141 |
| C4 |
-0.437 |
-0.132 |
0.548 |
| C5 |
-1.607 |
-0.262 |
-0.447 |
| C6 |
-2.854 |
0.512 |
-0.011 |
| H7 |
1.713 |
1.664 |
0.203 |
| H8 |
3.336 |
1.048 |
0.646 |
| H9 |
2.913 |
1.258 |
-1.066 |
| H10 |
2.769 |
-1.225 |
-0.430 |
| H11 |
0.599 |
-2.019 |
0.045 |
| H12 |
-0.181 |
0.941 |
0.676 |
| H13 |
-0.788 |
-0.485 |
1.546 |
| H14 |
-1.270 |
0.091 |
-1.446 |
| H15 |
-1.859 |
-1.338 |
-0.575 |
| H16 |
-3.680 |
0.401 |
-0.742 |
| H17 |
-3.227 |
0.156 |
0.973 |
| H18 |
-2.641 |
1.597 |
0.091 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.5017 | 2.5608 | 3.1924 | 4.2977 | 5.3735 | 1.1069 | 1.1119 | 1.1117 | 2.2109 | 3.5225 | 2.7879 | 3.9385 | 4.1007 | 4.9457 | 6.2405 | 5.8781 | 5.1870 |
C2 | 1.5017 | | 1.3505 | 2.5657 | 3.6388 | 4.9663 | 2.1876 | 2.1646 | 2.1646 | 1.1047 | 2.0998 | 2.7336 | 3.2659 | 3.5792 | 3.9895 | 5.7906 | 5.3917 | 5.0982 | C3 | 2.5608 | 1.3505 | | 1.5043 | 2.5397 | 3.9042 | 2.7621 | 3.2787 | 3.2923 | 2.0988 | 1.1055 | 2.1669 | 2.1455 | 2.7804 | 2.7554 | 4.7291 | 4.2255 | 4.2462 | C4 | 3.1924 | 2.5657 | 1.5043 | | 1.5413 | 2.5625 | 2.8233 | 3.9544 | 3.9698 | 3.5255 | 2.2107 | 1.1106 | 1.1153 | 2.1726 | 2.1762 | 3.5302 | 2.8365 | 2.8376 | C5 | 4.2977 | 3.6388 | 2.5397 | 1.5413 | | 1.5303 | 3.8932 | 5.2291 | 4.8087 | 4.4810 | 2.8636 | 2.1775 | 2.1666 | 1.1120 | 1.1130 | 2.1958 | 2.1938 | 2.1934 | C6 | 5.3735 | 4.9663 | 3.9042 | 2.5625 | 1.5303 | | 4.7148 | 6.2473 | 5.9094 | 5.8995 | 4.2815 | 2.7928 | 2.7722 | 2.1784 | 2.1753 | 1.1087 | 1.1106 | 1.1104 | H7 | 1.1069 | 2.1876 | 2.7621 | 2.8233 | 3.8932 | 4.7148 | | 1.7917 | 1.7926 | 3.1404 | 3.8516 | 2.0818 | 3.5605 | 3.7544 | 4.7305 | 5.6187 | 5.2222 | 4.3555 | H8 | 1.1119 | 2.1646 | 3.2787 | 3.9544 | 5.2291 | 6.2473 | 1.7917 | | 1.7758 | 2.5773 | 4.1540 | 3.5183 | 4.4903 | 5.1486 | 5.8451 | 7.1807 | 6.6311 | 6.0270 | H9 | 1.1117 | 2.1646 | 3.2923 | 3.9698 | 4.8087 | 5.9094 | 1.7926 | 1.7758 | | 2.5671 | 4.1628 | 3.5641 | 4.8533 | 4.3593 | 5.4541 | 6.6557 | 6.5624 | 5.6825 | H10 | 2.2109 | 1.1047 | 2.0988 | 3.5255 | 4.4810 | 5.8995 | 3.1404 | 2.5773 | 2.5671 | | 2.3591 | 3.8226 | 4.1358 | 4.3677 | 4.6311 | 6.6578 | 6.3106 | 6.1232 | H11 | 3.5225 | 2.0998 | 1.1055 | 2.2107 | 2.8636 | 4.2815 | 3.8516 | 4.1540 | 4.1628 | 2.3591 | | 3.1252 | 2.5557 | 3.1893 | 2.6247 | 4.9788 | 4.4978 | 4.8552 | H12 | 2.7879 | 2.7336 | 2.1669 | 1.1106 | 2.1775 | 2.7928 | 2.0818 | 3.5183 | 3.5641 | 3.8226 | 3.1252 | | 1.7771 | 2.5322 | 3.0941 | 3.8136 | 3.1596 | 2.6119 | H13 | 3.9385 | 3.2659 | 2.1455 | 1.1153 | 2.1666 | 2.7722 | 3.5605 | 4.4903 | 4.8533 | 4.1358 | 2.5557 | 1.7771 | | 3.0853 | 2.5250 | 3.7927 | 2.5861 | 3.1433 | H14 | 4.1007 | 3.5792 | 2.7804 | 2.1726 | 1.1120 | 2.1784 | 3.7544 | 5.1486 | 4.3593 | 4.3677 | 3.1893 | 2.5322 | 3.0853 | | 1.7742 | 2.5296 | 3.1120 | 2.5516 | H15 | 4.9457 | 3.9895 | 2.7554 | 2.1762 | 1.1130 | 2.1753 | 4.7305 | 5.8451 | 5.4541 | 4.6311 | 2.6247 | 3.0941 | 2.5250 | 1.7742 | | 2.5242 | 2.5497 | 3.1098 | H16 | 6.2405 | 5.7906 | 4.7291 | 3.5302 | 2.1958 | 1.1087 | 5.6187 | 7.1807 | 6.6557 | 6.6578 | 4.9788 | 3.8136 | 3.7927 | 2.5296 | 2.5242 | | 1.7905 | 1.7899 | H17 | 5.8781 | 5.3917 | 4.2255 | 2.8365 | 2.1938 | 1.1106 | 5.2222 | 6.6311 | 6.5624 | 6.3106 | 4.4978 | 3.1596 | 2.5861 | 3.1120 | 2.5497 | 1.7905 | | 1.7879 | H18 | 5.1870 | 5.0982 | 4.2462 | 2.8376 | 2.1934 | 1.1104 | 4.3555 | 6.0270 | 5.6825 | 6.1232 | 4.8552 | 2.6119 | 3.1433 | 2.5516 | 3.1098 | 1.7899 | 1.7879 | |
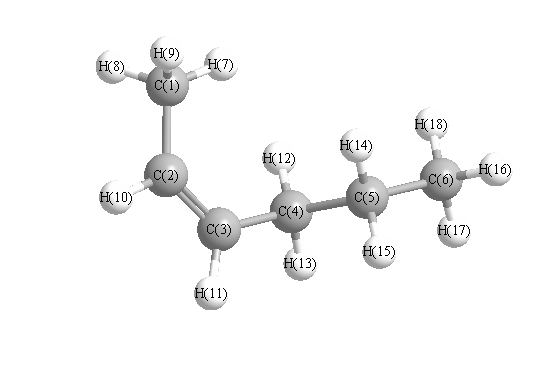 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.672 |
|
C1 |
C2 |
H10 |
115.192 |
| C2 |
C1 |
H7 |
113.112 |
|
C2 |
C1 |
H8 |
110.949 |
| C2 |
C1 |
H9 |
110.958 |
|
C2 |
C3 |
C4 |
127.898 |
| C2 |
C3 |
H11 |
117.165 |
|
C3 |
C2 |
H10 |
117.135 |
| C3 |
C4 |
C5 |
112.994 |
|
C3 |
C4 |
H12 |
111.028 |
| C3 |
C4 |
H13 |
109.064 |
|
C4 |
C3 |
H11 |
114.927 |
| C4 |
C5 |
C6 |
113.075 |
|
C4 |
C5 |
H14 |
108.860 |
| C4 |
C5 |
H15 |
109.078 |
|
C5 |
C4 |
H12 |
109.313 |
| C5 |
C4 |
H13 |
108.217 |
|
C5 |
C6 |
H16 |
111.627 |
| C5 |
C6 |
H17 |
111.352 |
|
C5 |
C6 |
H18 |
111.330 |
| C6 |
C5 |
H14 |
110.049 |
|
C6 |
C5 |
H15 |
109.751 |
| H7 |
C1 |
H8 |
107.702 |
|
H7 |
C1 |
H9 |
107.796 |
| H8 |
C1 |
H9 |
105.996 |
|
H12 |
C4 |
H13 |
105.943 |
| H14 |
C5 |
H16 |
93.996 |
|
H16 |
C6 |
H17 |
107.565 |
| H16 |
C6 |
H18 |
107.526 |
|
H17 |
C6 |
H18 |
107.218 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.020 |
|
|
|
| 2 |
C |
-0.089 |
|
|
|
| 3 |
C |
-0.046 |
|
|
|
| 4 |
C |
0.006 |
|
|
|
| 5 |
C |
-0.077 |
|
|
|
| 6 |
C |
-0.050 |
|
|
|
| 7 |
H |
0.031 |
|
|
|
| 8 |
H |
0.047 |
|
|
|
| 9 |
H |
0.046 |
|
|
|
| 10 |
H |
-0.011 |
|
|
|
| 11 |
H |
-0.015 |
|
|
|
| 12 |
H |
0.013 |
|
|
|
| 13 |
H |
0.030 |
|
|
|
| 14 |
H |
0.026 |
|
|
|
| 15 |
H |
0.022 |
|
|
|
| 16 |
H |
0.028 |
|
|
|
| 17 |
H |
0.030 |
|
|
|
| 18 |
H |
0.030 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.040 |
0.248 |
-0.013 |
0.252 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.791 |
0.878 |
-0.543 |
| y |
0.878 |
-39.390 |
0.001 |
| z |
-0.543 |
0.001 |
-41.041 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.424 |
0.878 |
-0.543 |
| y |
0.878 |
0.526 |
0.001 |
| z |
-0.543 |
0.001 |
-1.950 |
|
| Polar |
|---|
| 3z2-r2 | -3.901 |
|---|
| x2-y2 | 0.599 |
|---|
| xy | 0.878 |
|---|
| xz | -0.543 |
|---|
| yz | 0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.019 |
0.658 |
-0.664 |
| y |
0.658 |
9.902 |
0.075 |
| z |
-0.664 |
0.075 |
8.092 |
<r2> (average value of r
2) Å
2
| <r2> |
265.448 |
| (<r2>)1/2 |
16.293 |
