return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for H2OCH3OH (water methanol dimer)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-191.663370
Energy at 298.15K-191.669106
Counterpoise corrected energy-191.663370
CP Energy at 298.15K-191.669106
Counterpoise optimized geometry corrected energy 
CP opt. Energy at 298.15K 
Nuclear repulsion energy80.766513
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3917 3758 68.08      
2 A 3851 3694 34.52      
3 A 3797 3642 139.96      
4 A 3158 3029 29.28      
5 A 3122 2995 42.61      
6 A 3045 2921 43.39      
7 A 1730 1660 46.50      
8 A 1514 1452 3.44      
9 A 1501 1440 3.22      
10 A 1487 1427 4.78      
11 A 1405 1348 30.69      
12 A 1183 1135 0.82      
13 A 1095 1050 0.71      
14 A 1075 1031 106.12      
15 A 605 581 151.21      
16 A 476 457 38.13      
17 A 274 263 192.94      
18 A 190 183 1.68      
19 A 152 146 100.24      
20 A 121 117 16.46      
21 A 85 81 25.10      

Unscaled Zero Point Vibrational Energy (zpe) 16890.9 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 16205.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.78910 0.15897 0.13908

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -1.130 0.399 0.116
O2 -1.942 -0.078 -0.115
O3 0.814 0.754 0.150
H4 -2.159 -0.521 0.714
H5 0.886 1.187 -0.712
C6 1.233 -0.598 -0.036
H7 2.250 -0.659 -0.468
H8 1.254 -1.065 0.963
H9 0.523 -1.162 -0.671

Atom - Atom Distances (Å)
  H1 O2 O3 H4 H5 C6 H7 H8 H9
H10.96981.97641.50442.31712.56873.58962.92262.4054
O20.96982.89130.96483.15473.21804.24713.51462.7495
O31.97642.89133.28430.96661.42832.10792.04052.1048
H41.50440.96483.28433.77123.47484.56733.46563.0858
H52.31713.15470.96663.77121.94012.30872.83012.3770
C62.56873.21801.42833.47481.94011.10711.10271.1060
H73.58964.24712.10794.56732.30871.10711.78981.8102
H82.92263.51462.04053.46562.83011.10271.78981.7922
H92.40542.74952.10483.08582.37701.10601.81021.7922

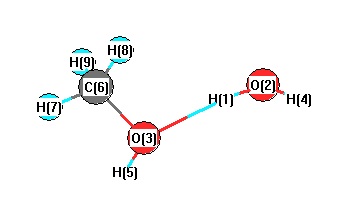
picture of water methanol dimer state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 O2 H4 102.082 H1 O3 H5 97.947
H1 O3 C6 96.630 O2 H1 O3 156.394
O3 C6 H7 111.856 O3 C6 H8 106.748
O3 C6 H9 111.667 H5 O3 C6 106.615
H7 C6 H8 108.180 H7 C6 H9 109.760
H8 C6 H9 108.469
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability