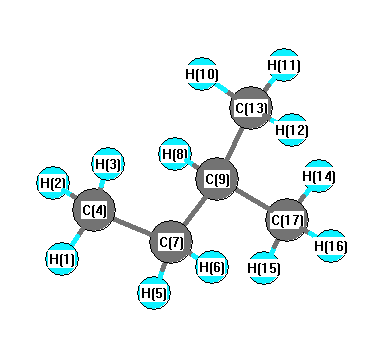
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3054 |
3036 |
25.38 |
|
|
|
| 2 |
A |
3048 |
3031 |
53.37 |
|
|
|
| 3 |
A |
3045 |
3027 |
17.96 |
|
|
|
| 4 |
A |
3042 |
3024 |
44.30 |
|
|
|
| 5 |
A |
3037 |
3020 |
41.79 |
|
|
|
| 6 |
A |
3033 |
3015 |
7.77 |
|
|
|
| 7 |
A |
2984 |
2967 |
23.13 |
|
|
|
| 8 |
A |
2965 |
2948 |
22.34 |
|
|
|
| 9 |
A |
2961 |
2944 |
31.15 |
|
|
|
| 10 |
A |
2956 |
2939 |
34.89 |
|
|
|
| 11 |
A |
2942 |
2925 |
34.59 |
|
|
|
| 12 |
A |
2925 |
2908 |
11.74 |
|
|
|
| 13 |
A |
1434 |
1426 |
3.95 |
|
|
|
| 14 |
A |
1429 |
1421 |
6.88 |
|
|
|
| 15 |
A |
1426 |
1418 |
4.16 |
|
|
|
| 16 |
A |
1420 |
1411 |
10.54 |
|
|
|
| 17 |
A |
1417 |
1409 |
1.83 |
|
|
|
| 18 |
A |
1406 |
1398 |
0.20 |
|
|
|
| 19 |
A |
1401 |
1393 |
0.18 |
|
|
|
| 20 |
A |
1351 |
1343 |
4.72 |
|
|
|
| 21 |
A |
1345 |
1337 |
1.26 |
|
|
|
| 22 |
A |
1336 |
1328 |
6.06 |
|
|
|
| 23 |
A |
1324 |
1316 |
1.10 |
|
|
|
| 24 |
A |
1306 |
1299 |
1.22 |
|
|
|
| 25 |
A |
1266 |
1258 |
3.60 |
|
|
|
| 26 |
A |
1241 |
1233 |
2.28 |
|
|
|
| 27 |
A |
1159 |
1152 |
1.28 |
|
|
|
| 28 |
A |
1150 |
1144 |
0.06 |
|
|
|
| 29 |
A |
1134 |
1127 |
2.84 |
|
|
|
| 30 |
A |
1040 |
1034 |
0.41 |
|
|
|
| 31 |
A |
997 |
991 |
3.10 |
|
|
|
| 32 |
A |
946 |
941 |
4.15 |
|
|
|
| 33 |
A |
931 |
926 |
0.34 |
|
|
|
| 34 |
A |
902 |
896 |
0.52 |
|
|
|
| 35 |
A |
891 |
886 |
1.30 |
|
|
|
| 36 |
A |
780 |
776 |
0.17 |
|
|
|
| 37 |
A |
749 |
745 |
2.87 |
|
|
|
| 38 |
A |
445 |
442 |
0.26 |
|
|
|
| 39 |
A |
403 |
400 |
0.24 |
|
|
|
| 40 |
A |
354 |
352 |
0.00 |
|
|
|
| 41 |
A |
261 |
259 |
0.01 |
|
|
|
| 42 |
A |
246 |
245 |
0.00 |
|
|
|
| 43 |
A |
222 |
221 |
0.01 |
|
|
|
| 44 |
A |
213 |
211 |
0.07 |
|
|
|
| 45 |
A |
95 |
94 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34004.5 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 33807.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.027 |
|
|
|
| 2 |
H |
0.029 |
|
|
|
| 3 |
H |
0.034 |
|
|
|
| 4 |
C |
-0.049 |
|
|
|
| 5 |
H |
0.012 |
|
|
|
| 6 |
H |
0.013 |
|
|
|
| 7 |
C |
-0.032 |
|
|
|
| 8 |
H |
0.003 |
|
|
|
| 9 |
C |
-0.158 |
|
|
|
| 10 |
H |
0.026 |
|
|
|
| 11 |
H |
0.024 |
|
|
|
| 12 |
H |
0.025 |
|
|
|
| 13 |
C |
-0.012 |
|
|
|
| 14 |
H |
0.022 |
|
|
|
| 15 |
H |
0.022 |
|
|
|
| 16 |
H |
0.025 |
|
|
|
| 17 |
C |
-0.009 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.036 |
-0.023 |
-0.064 |
0.077 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.546 |
0.228 |
-0.193 |
| y |
0.228 |
-35.372 |
0.054 |
| z |
-0.193 |
0.054 |
-34.978 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.371 |
0.228 |
-0.193 |
| y |
0.228 |
-0.110 |
0.054 |
| z |
-0.193 |
0.054 |
0.481 |
|
| Polar |
|---|
| 3z2-r2 | 0.962 |
|---|
| x2-y2 | -0.174 |
|---|
| xy | 0.228 |
|---|
| xz | -0.193 |
|---|
| yz | 0.054 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.906 |
-0.020 |
-0.009 |
| y |
-0.020 |
8.911 |
-0.030 |
| z |
-0.009 |
-0.030 |
8.047 |
<r2> (average value of r
2) Å
2
| <r2> |
159.485 |
| (<r2>)1/2 |
12.629 |