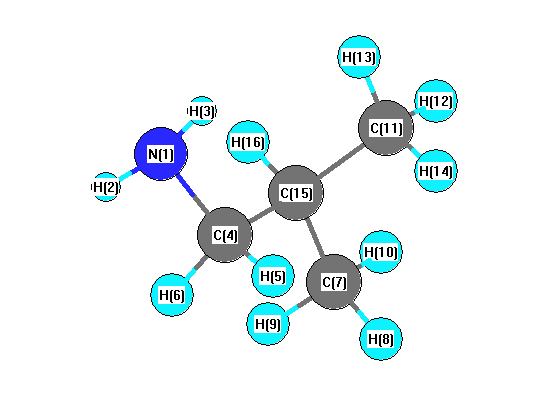
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3448 |
3428 |
0.34 |
|
|
|
| 2 |
A |
3372 |
3353 |
3.29 |
|
|
|
| 3 |
A |
3048 |
3030 |
33.35 |
|
|
|
| 4 |
A |
3047 |
3029 |
22.73 |
|
|
|
| 5 |
A |
3036 |
3019 |
54.89 |
|
|
|
| 6 |
A |
3032 |
3014 |
9.28 |
|
|
|
| 7 |
A |
2983 |
2966 |
36.56 |
|
|
|
| 8 |
A |
2960 |
2943 |
34.18 |
|
|
|
| 9 |
A |
2956 |
2939 |
16.93 |
|
|
|
| 10 |
A |
2953 |
2936 |
20.96 |
|
|
|
| 11 |
A |
2834 |
2818 |
83.85 |
|
|
|
| 12 |
A |
1591 |
1581 |
19.81 |
|
|
|
| 13 |
A |
1437 |
1429 |
12.27 |
|
|
|
| 14 |
A |
1427 |
1419 |
3.70 |
|
|
|
| 15 |
A |
1422 |
1414 |
2.34 |
|
|
|
| 16 |
A |
1412 |
1403 |
1.81 |
|
|
|
| 17 |
A |
1407 |
1399 |
1.13 |
|
|
|
| 18 |
A |
1370 |
1362 |
7.89 |
|
|
|
| 19 |
A |
1344 |
1336 |
1.42 |
|
|
|
| 20 |
A |
1334 |
1327 |
8.91 |
|
|
|
| 21 |
A |
1313 |
1305 |
0.82 |
|
|
|
| 22 |
A |
1292 |
1285 |
0.66 |
|
|
|
| 23 |
A |
1251 |
1243 |
7.34 |
|
|
|
| 24 |
A |
1188 |
1181 |
0.50 |
|
|
|
| 25 |
A |
1156 |
1149 |
1.61 |
|
|
|
| 26 |
A |
1130 |
1123 |
3.36 |
|
|
|
| 27 |
A |
1067 |
1060 |
8.89 |
|
|
|
| 28 |
A |
1061 |
1055 |
4.25 |
|
|
|
| 29 |
A |
934 |
929 |
0.34 |
|
|
|
| 30 |
A |
918 |
913 |
8.52 |
|
|
|
| 31 |
A |
901 |
896 |
42.10 |
|
|
|
| 32 |
A |
889 |
884 |
4.16 |
|
|
|
| 33 |
A |
833 |
828 |
64.76 |
|
|
|
| 34 |
A |
788 |
783 |
2.55 |
|
|
|
| 35 |
A |
472 |
470 |
6.26 |
|
|
|
| 36 |
A |
403 |
400 |
0.64 |
|
|
|
| 37 |
A |
349 |
347 |
0.24 |
|
|
|
| 38 |
A |
271 |
270 |
11.93 |
|
|
|
| 39 |
A |
254 |
253 |
0.65 |
|
|
|
| 40 |
A |
228 |
227 |
7.36 |
|
|
|
| 41 |
A |
209 |
208 |
27.48 |
|
|
|
| 42 |
A |
106 |
105 |
4.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31711.6 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 31527.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.165 |
|
|
|
| 2 |
H |
0.075 |
|
|
|
| 3 |
H |
0.074 |
|
|
|
| 4 |
C |
0.023 |
|
|
|
| 5 |
H |
-0.001 |
|
|
|
| 6 |
H |
0.019 |
|
|
|
| 7 |
C |
-0.015 |
|
|
|
| 8 |
H |
0.024 |
|
|
|
| 9 |
H |
0.023 |
|
|
|
| 10 |
H |
0.024 |
|
|
|
| 11 |
C |
-0.026 |
|
|
|
| 12 |
H |
0.027 |
|
|
|
| 13 |
H |
0.028 |
|
|
|
| 14 |
H |
0.021 |
|
|
|
| 15 |
C |
-0.140 |
|
|
|
| 16 |
H |
0.008 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.358 |
-0.145 |
1.024 |
1.094 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.173 |
0.617 |
-2.961 |
| y |
0.617 |
-33.386 |
0.509 |
| z |
-2.961 |
0.509 |
-35.433 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.236 |
0.617 |
-2.961 |
| y |
0.617 |
0.417 |
0.509 |
| z |
-2.961 |
0.509 |
-2.653 |
|
| Polar |
|---|
| 3z2-r2 | -5.306 |
|---|
| x2-y2 | 1.212 |
|---|
| xy | 0.617 |
|---|
| xz | -2.961 |
|---|
| yz | 0.509 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.236 |
-0.019 |
-0.069 |
| y |
-0.019 |
8.485 |
-0.003 |
| z |
-0.069 |
-0.003 |
7.286 |
<r2> (average value of r
2) Å
2
| <r2> |
151.541 |
| (<r2>)1/2 |
12.310 |