return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH(CH3)CN (Propanenitrile, 2-methyl-)

using model chemistry: QCISD(T)=FULL/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD(T)=FULL/cc-pVDZ
 hartrees
Energy at 0K-210.810838
Energy at 298.15K-210.818066
HF Energy-210.009072
Nuclear repulsion energy157.573122
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3153 3023        
2 A' 3147 3018        
3 A' 3065 2940        
4 A' 3056 2930        
5 A' 2269 2176        
6 A' 1501 1439        
7 A' 1491 1430        
8 A' 1417 1359        
9 A' 1350 1294        
10 A' 1194 1145        
11 A' 1122 1076        
12 A' 943 904        
13 A' 773 741        
14 A' 533 511        
15 A' 350 336        
16 A' 290 278        
17 A' 217 208        
18 A" 3151 3022        
19 A" 3143 3014        
20 A" 3054 2929        
21 A" 1481 1420        
22 A" 1479 1418        
23 A" 1398 1340        
24 A" 1330 1276        
25 A" 1148 1101        
26 A" 978 938        
27 A" 931 893        
28 A" 549 527        
29 A" 233 223        
30 A" 183 175        

Unscaled Zero Point Vibrational Energy (zpe) 22463.2 cm-1
Scaled (by 0.959) Zero Point Vibrational Energy (zpe) 21542.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)=FULL/cc-pVDZ
ABC
0.26249 0.12918 0.09495

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)=FULL/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.418 -2.203 0.000
C2 0.025 -1.093 0.000
C3 -0.448 0.318 0.000
C4 0.025 1.038 1.278
C5 0.025 1.038 -1.278
H6 -1.554 0.277 0.000
H7 -0.366 2.071 1.287
H8 -0.331 0.518 2.185
H9 1.129 1.080 1.312
H10 -0.366 2.071 -1.287
H11 -0.331 0.518 -2.185
H12 1.129 1.080 -1.312

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12
N11.17782.66523.50593.50593.16854.53223.56913.60624.53223.56913.6062
C21.17781.48742.48442.48442.09063.43802.73752.76743.43802.73752.7674
C32.66521.48741.54131.54131.10742.17692.19682.18792.17692.19682.1879
C43.50592.48441.54132.55642.16951.10491.10391.10512.79343.51972.8158
C53.50592.48441.54132.55642.16952.79343.51972.81581.10491.10391.1051
H63.16852.09061.10742.16952.16952.50772.51513.09262.50772.51513.0926
H74.53223.43802.17691.10492.79342.50771.79391.79372.57483.80373.1580
H83.56912.73752.19681.10393.51972.51511.79391.79163.80374.36913.8305
H93.60622.76742.18791.10512.81583.09261.79371.79163.15803.83052.6236
H104.53223.43802.17692.79341.10492.50772.57483.80373.15801.79391.7937
H113.56912.73752.19683.51971.10392.51513.80374.36913.83051.79391.7916
H123.60622.76742.18792.81581.10513.09263.15803.83052.62361.79371.7916

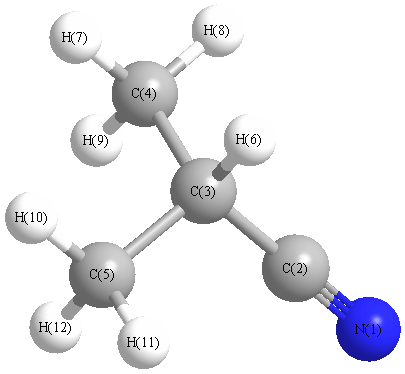
picture of Propanenitrile, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 179.069 C2 C3 C4 110.211
C2 C3 C5 110.211 C2 C3 H6 106.430
C3 C4 H7 109.593 C3 C4 H8 111.217
C3 C4 H9 110.444 C3 C5 H10 109.593
C3 C5 H11 111.217 C3 C5 H12 110.444
C4 C3 C5 112.052 C4 C3 H6 108.880
C5 C3 H6 108.880 H7 C4 H8 108.615
H7 C4 H9 108.512 H8 C4 H9 108.394
H10 C5 H11 108.615 H10 C5 H12 108.512
H11 C5 H12 108.394
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability