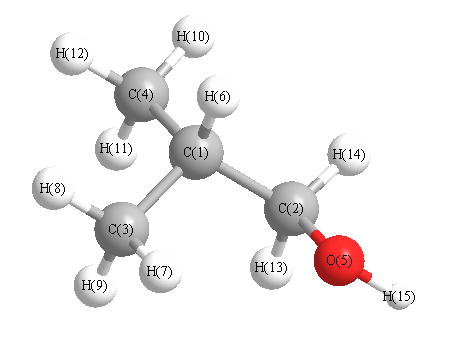
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3665 |
3644 |
11.52 |
|
|
|
| 2 |
A |
3049 |
3031 |
30.75 |
|
|
|
| 3 |
A |
3045 |
3027 |
21.56 |
|
|
|
| 4 |
A |
3037 |
3019 |
35.51 |
|
|
|
| 5 |
A |
3030 |
3012 |
28.63 |
|
|
|
| 6 |
A |
3003 |
2985 |
33.61 |
|
|
|
| 7 |
A |
2961 |
2944 |
28.42 |
|
|
|
| 8 |
A |
2955 |
2938 |
21.50 |
|
|
|
| 9 |
A |
2953 |
2935 |
21.91 |
|
|
|
| 10 |
A |
2868 |
2851 |
63.43 |
|
|
|
| 11 |
A |
1436 |
1428 |
11.13 |
|
|
|
| 12 |
A |
1427 |
1419 |
5.74 |
|
|
|
| 13 |
A |
1420 |
1412 |
0.65 |
|
|
|
| 14 |
A |
1414 |
1406 |
5.83 |
|
|
|
| 15 |
A |
1407 |
1399 |
0.36 |
|
|
|
| 16 |
A |
1368 |
1360 |
36.81 |
|
|
|
| 17 |
A |
1349 |
1341 |
3.36 |
|
|
|
| 18 |
A |
1339 |
1331 |
4.94 |
|
|
|
| 19 |
A |
1329 |
1321 |
4.96 |
|
|
|
| 20 |
A |
1305 |
1297 |
1.50 |
|
|
|
| 21 |
A |
1261 |
1254 |
1.07 |
|
|
|
| 22 |
A |
1198 |
1191 |
11.06 |
|
|
|
| 23 |
A |
1154 |
1147 |
0.55 |
|
|
|
| 24 |
A |
1120 |
1114 |
1.00 |
|
|
|
| 25 |
A |
1088 |
1081 |
0.28 |
|
|
|
| 26 |
A |
1051 |
1044 |
118.38 |
|
|
|
| 27 |
A |
938 |
932 |
0.33 |
|
|
|
| 28 |
A |
919 |
913 |
4.33 |
|
|
|
| 29 |
A |
891 |
886 |
0.40 |
|
|
|
| 30 |
A |
870 |
864 |
6.55 |
|
|
|
| 31 |
A |
803 |
798 |
6.58 |
|
|
|
| 32 |
A |
477 |
475 |
6.52 |
|
|
|
| 33 |
A |
408 |
406 |
2.82 |
|
|
|
| 34 |
A |
343 |
341 |
3.36 |
|
|
|
| 35 |
A |
292 |
290 |
84.81 |
|
|
|
| 36 |
A |
252 |
251 |
1.33 |
|
|
|
| 37 |
A |
244 |
243 |
27.31 |
|
|
|
| 38 |
A |
218 |
217 |
0.16 |
|
|
|
| 39 |
A |
115 |
115 |
3.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28999.1 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 28830.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.167 |
|
|
|
| 2 |
C |
0.136 |
|
|
|
| 3 |
C |
-0.023 |
|
|
|
| 4 |
C |
-0.034 |
|
|
|
| 5 |
O |
-0.243 |
|
|
|
| 6 |
H |
0.011 |
|
|
|
| 7 |
H |
0.028 |
|
|
|
| 8 |
H |
0.028 |
|
|
|
| 9 |
H |
0.027 |
|
|
|
| 10 |
H |
0.016 |
|
|
|
| 11 |
H |
0.027 |
|
|
|
| 12 |
H |
0.033 |
|
|
|
| 13 |
H |
0.025 |
|
|
|
| 14 |
H |
0.003 |
|
|
|
| 15 |
H |
0.134 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.008 |
0.765 |
0.644 |
1.420 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.310 |
-2.275 |
-1.800 |
| y |
-2.275 |
-31.930 |
0.405 |
| z |
-1.800 |
0.405 |
-33.387 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.652 |
-2.275 |
-1.800 |
| y |
-2.275 |
2.919 |
0.405 |
| z |
-1.800 |
0.405 |
0.733 |
|
| Polar |
|---|
| 3z2-r2 | 1.466 |
|---|
| x2-y2 | -4.380 |
|---|
| xy | -2.275 |
|---|
| xz | -1.800 |
|---|
| yz | 0.405 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.134 |
-0.192 |
-0.018 |
| y |
-0.192 |
8.092 |
0.019 |
| z |
-0.018 |
0.019 |
6.917 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |