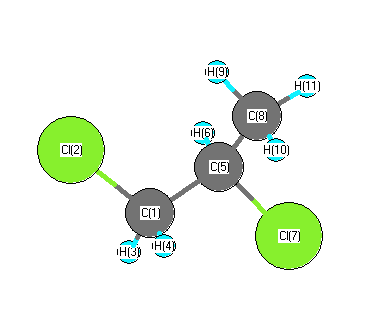
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3097 |
3079 |
3.46 |
|
|
|
| 2 |
A |
3086 |
3068 |
7.60 |
|
|
|
| 3 |
A |
3070 |
3053 |
8.18 |
|
|
|
| 4 |
A |
3030 |
3013 |
5.98 |
|
|
|
| 5 |
A |
3020 |
3003 |
4.17 |
|
|
|
| 6 |
A |
2984 |
2967 |
8.24 |
|
|
|
| 7 |
A |
1419 |
1411 |
3.95 |
|
|
|
| 8 |
A |
1409 |
1401 |
8.45 |
|
|
|
| 9 |
A |
1398 |
1390 |
3.58 |
|
|
|
| 10 |
A |
1342 |
1334 |
14.17 |
|
|
|
| 11 |
A |
1316 |
1308 |
1.21 |
|
|
|
| 12 |
A |
1233 |
1225 |
1.51 |
|
|
|
| 13 |
A |
1197 |
1190 |
6.66 |
|
|
|
| 14 |
A |
1148 |
1141 |
22.77 |
|
|
|
| 15 |
A |
1122 |
1115 |
2.96 |
|
|
|
| 16 |
A |
1033 |
1027 |
5.39 |
|
|
|
| 17 |
A |
986 |
980 |
20.67 |
|
|
|
| 18 |
A |
887 |
882 |
4.30 |
|
|
|
| 19 |
A |
845 |
840 |
1.37 |
|
|
|
| 20 |
A |
717 |
713 |
32.76 |
|
|
|
| 21 |
A |
649 |
646 |
59.80 |
|
|
|
| 22 |
A |
399 |
396 |
1.03 |
|
|
|
| 23 |
A |
338 |
336 |
2.44 |
|
|
|
| 24 |
A |
272 |
271 |
0.37 |
|
|
|
| 25 |
A |
249 |
248 |
0.36 |
|
|
|
| 26 |
A |
199 |
198 |
7.46 |
|
|
|
| 27 |
A |
112 |
111 |
4.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18277.6 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 18171.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.022 |
|
|
|
| 2 |
Cl |
-0.117 |
|
|
|
| 3 |
H |
0.090 |
|
|
|
| 4 |
H |
0.087 |
|
|
|
| 5 |
C |
-0.179 |
|
|
|
| 6 |
H |
0.078 |
|
|
|
| 7 |
Cl |
-0.107 |
|
|
|
| 8 |
C |
0.015 |
|
|
|
| 9 |
H |
0.056 |
|
|
|
| 10 |
H |
0.047 |
|
|
|
| 11 |
H |
0.052 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.029 |
0.537 |
0.073 |
0.542 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.296 |
0.810 |
-0.200 |
| y |
0.810 |
-42.932 |
-0.459 |
| z |
-0.200 |
-0.459 |
-43.696 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.981 |
0.810 |
-0.200 |
| y |
0.810 |
4.564 |
-0.459 |
| z |
-0.200 |
-0.459 |
3.417 |
|
| Polar |
|---|
| 3z2-r2 | 6.835 |
|---|
| x2-y2 | -8.364 |
|---|
| xy | 0.810 |
|---|
| xz | -0.200 |
|---|
| yz | -0.459 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.983 |
-0.717 |
0.315 |
| y |
-0.717 |
7.519 |
-0.168 |
| z |
0.315 |
-0.168 |
6.144 |
<r2> (average value of r
2) Å
2
| <r2> |
245.661 |
| (<r2>)1/2 |
15.674 |