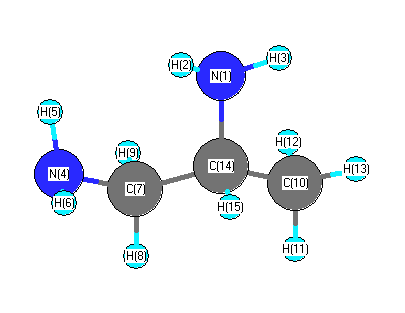
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3567 |
3442 |
0.03 |
|
|
|
| 2 |
A |
3556 |
3431 |
0.43 |
|
|
|
| 3 |
A |
3480 |
3358 |
2.61 |
|
|
|
| 4 |
A |
3477 |
3355 |
1.37 |
|
|
|
| 5 |
A |
3132 |
3022 |
24.21 |
|
|
|
| 6 |
A |
3117 |
3008 |
29.11 |
|
|
|
| 7 |
A |
3086 |
2978 |
29.24 |
|
|
|
| 8 |
A |
3038 |
2932 |
37.10 |
|
|
|
| 9 |
A |
3031 |
2925 |
28.26 |
|
|
|
| 10 |
A |
2900 |
2798 |
91.68 |
|
|
|
| 11 |
A |
1642 |
1585 |
1.21 |
|
|
|
| 12 |
A |
1632 |
1575 |
47.69 |
|
|
|
| 13 |
A |
1472 |
1420 |
1.92 |
|
|
|
| 14 |
A |
1460 |
1409 |
6.37 |
|
|
|
| 15 |
A |
1446 |
1396 |
0.39 |
|
|
|
| 16 |
A |
1410 |
1361 |
0.69 |
|
|
|
| 17 |
A |
1390 |
1341 |
14.84 |
|
|
|
| 18 |
A |
1379 |
1331 |
4.27 |
|
|
|
| 19 |
A |
1364 |
1317 |
10.53 |
|
|
|
| 20 |
A |
1327 |
1280 |
10.71 |
|
|
|
| 21 |
A |
1262 |
1218 |
1.26 |
|
|
|
| 22 |
A |
1198 |
1156 |
4.37 |
|
|
|
| 23 |
A |
1174 |
1133 |
6.54 |
|
|
|
| 24 |
A |
1129 |
1089 |
12.40 |
|
|
|
| 25 |
A |
1049 |
1013 |
13.41 |
|
|
|
| 26 |
A |
1028 |
992 |
1.13 |
|
|
|
| 27 |
A |
958 |
925 |
10.28 |
|
|
|
| 28 |
A |
916 |
884 |
98.48 |
|
|
|
| 29 |
A |
887 |
856 |
71.62 |
|
|
|
| 30 |
A |
849 |
819 |
70.11 |
|
|
|
| 31 |
A |
803 |
775 |
57.24 |
|
|
|
| 32 |
A |
491 |
474 |
0.50 |
|
|
|
| 33 |
A |
469 |
452 |
20.98 |
|
|
|
| 34 |
A |
366 |
353 |
12.27 |
|
|
|
| 35 |
A |
355 |
343 |
32.65 |
|
|
|
| 36 |
A |
257 |
248 |
8.52 |
|
|
|
| 37 |
A |
241 |
233 |
26.79 |
|
|
|
| 38 |
A |
224 |
216 |
22.17 |
|
|
|
| 39 |
A |
116 |
112 |
5.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30337.4 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 29275.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.181 |
|
|
|
| 2 |
H |
0.066 |
|
|
|
| 3 |
H |
0.085 |
|
|
|
| 4 |
N |
-0.218 |
|
|
|
| 5 |
H |
0.089 |
|
|
|
| 6 |
H |
0.083 |
|
|
|
| 7 |
C |
0.059 |
|
|
|
| 8 |
H |
0.032 |
|
|
|
| 9 |
H |
0.040 |
|
|
|
| 10 |
C |
-0.035 |
|
|
|
| 11 |
H |
0.038 |
|
|
|
| 12 |
H |
0.047 |
|
|
|
| 13 |
H |
0.031 |
|
|
|
| 14 |
C |
-0.134 |
|
|
|
| 15 |
H |
-0.002 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.775 |
1.132 |
1.581 |
2.093 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
143.522 |
| (<r2>)1/2 |
11.980 |