return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4O2 (Methyl glyoxal)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-266.454824
Energy at 298.15K-266.458967
HF Energy-265.660023
Nuclear repulsion energy158.943374
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3197 3067 5.37      
2 A' 3077 2952 1.46      
3 A' 2990 2868 87.97      
4 A' 1832 1758 8.27      
5 A' 1806 1732 168.91      
6 A' 1459 1400 12.34      
7 A' 1404 1347 26.52      
8 A' 1365 1310 2.85      
9 A' 1276 1224 18.62      
10 A' 1018 976 3.06      
11 A' 803 771 11.14      
12 A' 578 555 10.82      
13 A' 482 463 26.19      
14 A' 249 239 15.31      
15 A" 3158 3030 4.14      
16 A" 1469 1409 8.55      
17 A" 1081 1037 1.85      
18 A" 916 879 0.44      
19 A" 475 456 0.57      
20 A" 147 141 12.15      
21 A" 101 97 6.46      

Unscaled Zero Point Vibrational Energy (zpe) 14440.1 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 13853.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.30052 0.14647 0.10033

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.852 -0.724 0.000
C2 0.000 0.559 0.000
C3 1.502 0.372 0.000
O4 -0.370 -1.837 0.000
O5 -0.574 1.632 0.000
H6 -1.953 -0.524 0.000
H7 1.999 1.353 0.000
H8 1.804 -0.213 0.887
H9 1.804 -0.213 -0.887

Atom - Atom Distances (Å)
  C1 C2 C3 O4 O5 H6 H7 H8 H9
C11.53992.59651.21322.37271.11863.52792.84692.8469
C21.53991.51352.42431.21772.23282.15152.15352.1535
C32.59651.51352.89542.42913.56891.10051.10431.1043
O41.21322.42432.89543.47562.05663.97412.85532.8553
O52.37271.21772.42913.47562.55922.58893.13853.1385
H61.11862.23283.56892.05662.55924.37543.87293.8729
H73.52792.15151.10053.97412.58894.37541.81031.8103
H82.84692.15351.10432.85533.13853.87291.81031.7734
H92.84692.15351.10432.85533.13853.87291.81031.7734

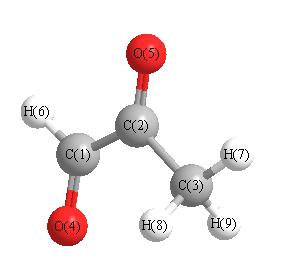
picture of Methyl glyoxal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 116.500 C1 C2 O5 118.256
C2 C1 O4 122.985 C2 C1 H6 113.298
C2 C3 H7 109.781 C2 C3 H8 109.711
C2 C3 H9 109.711 C3 C2 O5 125.244
O4 C1 H6 123.717 H7 C3 H8 110.383
H7 C3 H9 110.383 H8 C3 H9 106.826
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability