Jump to
S1C2
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -1071.506671 |
| Energy at 298.15K | -1071.509239 |
| HF Energy | -1070.756261 |
| Nuclear repulsion energy | 262.286902 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3194 |
3056 |
1.28 |
|
|
|
| 2 |
A' |
3026 |
2895 |
43.71 |
|
|
|
| 3 |
A' |
1875 |
1794 |
120.45 |
|
|
|
| 4 |
A' |
1417 |
1355 |
4.93 |
|
|
|
| 5 |
A' |
1234 |
1181 |
10.33 |
|
|
|
| 6 |
A' |
1101 |
1053 |
20.20 |
|
|
|
| 7 |
A' |
810 |
775 |
23.45 |
|
|
|
| 8 |
A' |
446 |
427 |
4.51 |
|
|
|
| 9 |
A' |
330 |
315 |
22.69 |
|
|
|
| 10 |
A' |
261 |
250 |
1.81 |
|
|
|
| 11 |
A" |
1261 |
1206 |
26.02 |
|
|
|
| 12 |
A" |
1029 |
985 |
19.30 |
|
|
|
| 13 |
A" |
780 |
746 |
92.75 |
|
|
|
| 14 |
A" |
290 |
278 |
2.32 |
|
|
|
| 15 |
A" |
84 |
80 |
7.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8568.3 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 8197.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.388 |
-0.017 |
0.000 |
| C2 |
-0.172 |
1.407 |
0.000 |
| H3 |
1.486 |
0.004 |
0.000 |
| Cl4 |
-0.172 |
-0.847 |
1.479 |
| Cl5 |
-0.172 |
-0.847 |
-1.479 |
| O6 |
0.542 |
2.376 |
0.000 |
| H7 |
-1.286 |
1.459 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5307 | 1.0979 | 1.7859 | 1.7859 | 2.3984 | 2.2328 |
C2 | 1.5307 | | 2.1721 | 2.6962 | 2.6962 | 1.2036 | 1.1159 | H3 | 1.0979 | 2.1721 | | 2.3791 | 2.3791 | 2.5531 | 3.1314 | Cl4 | 1.7859 | 2.6962 | 2.3791 | | 2.9575 | 3.6176 | 2.9580 | Cl5 | 1.7859 | 2.6962 | 2.3791 | 2.9575 | | 3.6176 | 2.9580 | O6 | 2.3984 | 1.2036 | 2.5531 | 3.6176 | 3.6176 | | 2.0456 | H7 | 2.2328 | 1.1159 | 3.1314 | 2.9580 | 2.9580 | 2.0456 | |
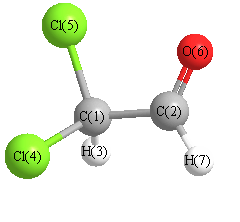 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
122.147 |
|
C1 |
C2 |
H7 |
114.142 |
| C2 |
C1 |
H3 |
110.361 |
|
C2 |
C1 |
Cl4 |
108.525 |
| C2 |
C1 |
Cl5 |
108.525 |
|
H3 |
C1 |
Cl4 |
108.820 |
| H3 |
C1 |
Cl5 |
108.820 |
|
Cl4 |
C1 |
Cl5 |
111.791 |
| O6 |
C2 |
H7 |
123.711 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -1071.504360 |
| Energy at 298.15K | |
| HF Energy | -1070.753341 |
| Nuclear repulsion energy | 265.018646 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3147 |
3011 |
5.18 |
|
|
|
| 2 |
A |
3011 |
2881 |
55.76 |
|
|
|
| 3 |
A |
1885 |
1803 |
106.59 |
|
|
|
| 4 |
A |
1416 |
1354 |
15.91 |
|
|
|
| 5 |
A |
1290 |
1234 |
10.74 |
|
|
|
| 6 |
A |
1245 |
1191 |
14.15 |
|
|
|
| 7 |
A |
1057 |
1011 |
11.10 |
|
|
|
| 8 |
A |
958 |
917 |
5.19 |
|
|
|
| 9 |
A |
857 |
820 |
67.82 |
|
|
|
| 10 |
A |
662 |
633 |
30.57 |
|
|
|
| 11 |
A |
630 |
603 |
29.20 |
|
|
|
| 12 |
A |
353 |
337 |
0.91 |
|
|
|
| 13 |
A |
280 |
268 |
3.05 |
|
|
|
| 14 |
A |
226 |
216 |
2.69 |
|
|
|
| 15 |
A |
92 |
88 |
9.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8553.8 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 8183.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.102 |
-0.030 |
0.520 |
| C2 |
-0.727 |
-1.284 |
0.189 |
| H3 |
0.227 |
0.048 |
1.611 |
| Cl4 |
1.741 |
-0.268 |
-0.168 |
| Cl5 |
-0.667 |
1.456 |
-0.057 |
| O6 |
-1.816 |
-1.266 |
-0.315 |
| H7 |
-0.204 |
-2.227 |
0.480 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5386 | 1.1015 | 1.7941 | 1.7700 | 2.4298 | 2.2179 |
C2 | 1.5386 | | 2.1696 | 2.6931 | 2.7514 | 1.2006 | 1.1168 | H3 | 1.1015 | 2.1696 | | 2.3582 | 2.3589 | 3.1007 | 2.5770 | Cl4 | 1.7941 | 2.6931 | 2.3582 | | 2.9641 | 3.6982 | 2.8358 | Cl5 | 1.7700 | 2.7514 | 2.3589 | 2.9641 | | 2.9660 | 3.7503 | O6 | 2.4298 | 1.2006 | 3.1007 | 3.6982 | 2.9660 | | 2.0383 | H7 | 2.2179 | 1.1168 | 2.5770 | 2.8358 | 3.7503 | 2.0383 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
124.547 |
|
C1 |
C2 |
H7 |
112.311 |
| C2 |
C1 |
H3 |
109.412 |
|
C2 |
C1 |
Cl4 |
107.570 |
| C2 |
C1 |
Cl5 |
112.334 |
|
H3 |
C1 |
Cl4 |
106.601 |
| H3 |
C1 |
Cl5 |
108.207 |
|
Cl4 |
C1 |
Cl5 |
112.536 |
| O6 |
C2 |
H7 |
123.140 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
