Jump to
S1C2
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -1072.483210 |
| Energy at 298.15K | |
| HF Energy | -1072.483210 |
| Nuclear repulsion energy | 263.535004 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3023 |
3006 |
3.60 |
99.20 |
0.18 |
0.31 |
| 2 |
A |
2832 |
2816 |
70.58 |
170.22 |
0.32 |
0.48 |
| 3 |
A |
1792 |
1782 |
135.72 |
17.19 |
0.46 |
0.63 |
| 4 |
A |
1328 |
1320 |
10.73 |
5.52 |
0.63 |
0.77 |
| 5 |
A |
1194 |
1187 |
13.18 |
4.56 |
0.74 |
0.85 |
| 6 |
A |
1139 |
1133 |
9.07 |
9.48 |
0.73 |
0.85 |
| 7 |
A |
984 |
978 |
12.18 |
1.95 |
0.64 |
0.78 |
| 8 |
A |
892 |
886 |
9.16 |
6.09 |
0.60 |
0.75 |
| 9 |
A |
763 |
759 |
81.27 |
4.41 |
0.67 |
0.80 |
| 10 |
A |
622 |
618 |
38.83 |
9.55 |
0.09 |
0.16 |
| 11 |
A |
590 |
587 |
43.74 |
7.74 |
0.44 |
0.61 |
| 12 |
A |
326 |
324 |
2.30 |
3.54 |
0.21 |
0.35 |
| 13 |
A |
264 |
262 |
2.95 |
7.17 |
0.64 |
0.78 |
| 14 |
A |
209 |
208 |
1.80 |
1.89 |
0.70 |
0.82 |
| 15 |
A |
89 |
88 |
6.91 |
2.25 |
0.70 |
0.82 |
Unscaled Zero Point Vibrational Energy (zpe) 8022.6 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 7976.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.096 |
-0.012 |
0.530 |
| C2 |
-0.678 |
-1.306 |
0.215 |
| H3 |
0.242 |
0.091 |
1.624 |
| Cl4 |
1.755 |
-0.219 |
-0.174 |
| Cl5 |
-0.732 |
1.444 |
-0.059 |
| O6 |
-1.753 |
-1.349 |
-0.336 |
| H7 |
-0.116 |
-2.221 |
0.562 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5401 | 1.1086 | 1.8134 | 1.7761 | 2.4404 | 2.2186 |
C2 | 1.5401 | | 2.1871 | 2.6927 | 2.7641 | 1.2085 | 1.1284 | H3 | 1.1086 | 2.1871 | | 2.3707 | 2.3686 | 3.1452 | 2.5693 | Cl4 | 1.8134 | 2.6927 | 2.3707 | | 2.9937 | 3.6887 | 2.8369 | Cl5 | 1.7761 | 2.7641 | 2.3686 | 2.9937 | | 2.9866 | 3.7677 | O6 | 2.4404 | 1.2085 | 3.1452 | 3.6887 | 2.9866 | | 2.0604 | H7 | 2.2186 | 1.1284 | 2.5693 | 2.8369 | 3.7677 | 2.0604 | |
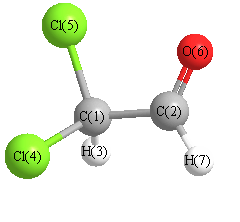 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
124.776 |
|
C1 |
C2 |
H7 |
111.563 |
| C2 |
C1 |
H3 |
110.260 |
|
C2 |
C1 |
Cl4 |
106.546 |
| C2 |
C1 |
Cl5 |
112.731 |
|
H3 |
C1 |
Cl4 |
105.916 |
| H3 |
C1 |
Cl5 |
108.151 |
|
Cl4 |
C1 |
Cl5 |
113.024 |
| O6 |
C2 |
H7 |
123.661 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.211 |
|
|
|
| 2 |
C |
0.206 |
|
|
|
| 3 |
H |
0.133 |
|
|
|
| 4 |
Cl |
-0.035 |
|
|
|
| 5 |
Cl |
-0.002 |
|
|
|
| 6 |
O |
-0.116 |
|
|
|
| 7 |
H |
0.025 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.994 |
-0.629 |
1.815 |
2.163 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.117 |
-1.958 |
-1.056 |
| y |
-1.958 |
-41.572 |
-1.423 |
| z |
-1.056 |
-1.423 |
-39.936 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.363 |
-1.958 |
-1.056 |
| y |
-1.958 |
1.455 |
-1.423 |
| z |
-1.056 |
-1.423 |
3.908 |
|
| Polar |
|---|
| 3z2-r2 | 7.817 |
|---|
| x2-y2 | -4.545 |
|---|
| xy | -1.958 |
|---|
| xz | -1.056 |
|---|
| yz | -1.423 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.081 |
-0.725 |
0.211 |
| y |
-0.725 |
7.086 |
-0.396 |
| z |
0.211 |
-0.396 |
4.475 |
<r2> (average value of r
2) Å
2
| <r2> |
187.876 |
| (<r2>)1/2 |
13.707 |
