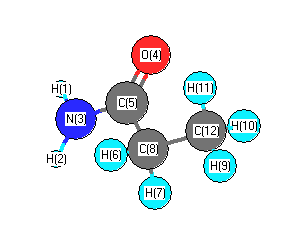
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3571 |
3576 |
15.59 |
|
|
|
| 2 |
A |
3435 |
3440 |
10.49 |
|
|
|
| 3 |
A |
3048 |
3053 |
20.29 |
|
|
|
| 4 |
A |
3034 |
3039 |
28.56 |
|
|
|
| 5 |
A |
2969 |
2974 |
23.70 |
|
|
|
| 6 |
A |
2965 |
2970 |
22.36 |
|
|
|
| 7 |
A |
2931 |
2935 |
24.86 |
|
|
|
| 8 |
A |
1715 |
1718 |
223.77 |
|
|
|
| 9 |
A |
1540 |
1542 |
79.18 |
|
|
|
| 10 |
A |
1435 |
1437 |
8.20 |
|
|
|
| 11 |
A |
1423 |
1425 |
5.14 |
|
|
|
| 12 |
A |
1401 |
1404 |
3.83 |
|
|
|
| 13 |
A |
1359 |
1361 |
4.02 |
|
|
|
| 14 |
A |
1337 |
1339 |
49.83 |
|
|
|
| 15 |
A |
1238 |
1240 |
15.24 |
|
|
|
| 16 |
A |
1234 |
1236 |
81.41 |
|
|
|
| 17 |
A |
1084 |
1086 |
2.37 |
|
|
|
| 18 |
A |
1053 |
1054 |
0.35 |
|
|
|
| 19 |
A |
1025 |
1027 |
3.22 |
|
|
|
| 20 |
A |
974 |
975 |
2.59 |
|
|
|
| 21 |
A |
793 |
794 |
6.81 |
|
|
|
| 22 |
A |
757 |
759 |
6.34 |
|
|
|
| 23 |
A |
614 |
615 |
5.14 |
|
|
|
| 24 |
A |
583 |
584 |
11.58 |
|
|
|
| 25 |
A |
482 |
483 |
8.14 |
|
|
|
| 26 |
A |
430 |
431 |
2.62 |
|
|
|
| 27 |
A |
259 |
259 |
163.38 |
|
|
|
| 28 |
A |
252 |
252 |
15.82 |
|
|
|
| 29 |
A |
201 |
201 |
0.77 |
|
|
|
| 30 |
A |
28 |
28 |
1.53 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21583.2 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 21617.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.082 |
|
|
|
| 2 |
H |
0.074 |
|
|
|
| 3 |
N |
-0.035 |
|
|
|
| 4 |
O |
-0.225 |
|
|
|
| 5 |
C |
0.039 |
|
|
|
| 6 |
H |
0.019 |
|
|
|
| 7 |
H |
0.007 |
|
|
|
| 8 |
C |
-0.029 |
|
|
|
| 9 |
H |
-0.008 |
|
|
|
| 10 |
H |
0.015 |
|
|
|
| 11 |
H |
0.022 |
|
|
|
| 12 |
C |
0.038 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.027 |
-3.204 |
0.389 |
3.387 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.736 |
2.492 |
-0.722 |
| y |
2.492 |
-32.721 |
-0.544 |
| z |
-0.722 |
-0.544 |
-31.256 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.252 |
2.492 |
-0.722 |
| y |
2.492 |
-4.225 |
-0.544 |
| z |
-0.722 |
-0.544 |
-2.027 |
|
| Polar |
|---|
| 3z2-r2 | -4.054 |
|---|
| x2-y2 | 6.984 |
|---|
| xy | 2.492 |
|---|
| xz | -0.722 |
|---|
| yz | -0.544 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.051 |
0.356 |
0.069 |
| y |
0.356 |
7.052 |
-0.017 |
| z |
0.069 |
-0.017 |
4.741 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |