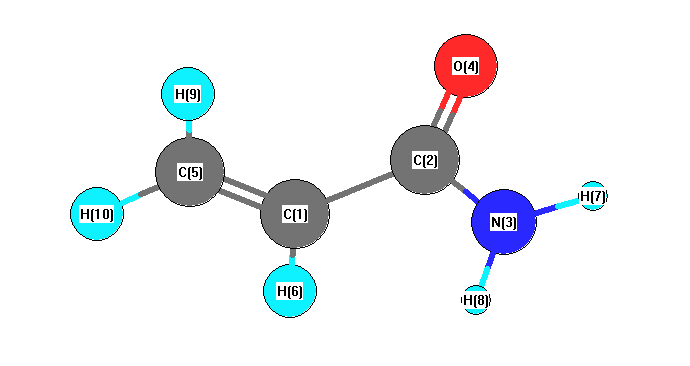
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3624 |
3603 |
28.94 |
|
|
|
| 2 |
A |
3480 |
3460 |
27.60 |
|
|
|
| 3 |
A |
3190 |
3171 |
1.76 |
|
|
|
| 4 |
A |
3087 |
3069 |
12.60 |
|
|
|
| 5 |
A |
3074 |
3056 |
9.66 |
|
|
|
| 6 |
A |
1734 |
1724 |
193.84 |
|
|
|
| 7 |
A |
1649 |
1640 |
24.22 |
|
|
|
| 8 |
A |
1533 |
1524 |
113.59 |
|
|
|
| 9 |
A |
1370 |
1362 |
93.17 |
|
|
|
| 10 |
A |
1308 |
1301 |
21.61 |
|
|
|
| 11 |
A |
1236 |
1229 |
47.02 |
|
|
|
| 12 |
A |
1070 |
1064 |
2.23 |
|
|
|
| 13 |
A |
991 |
985 |
1.83 |
|
|
|
| 14 |
A |
980 |
975 |
24.55 |
|
|
|
| 15 |
A |
944 |
939 |
8.78 |
|
|
|
| 16 |
A |
797 |
793 |
6.86 |
|
|
|
| 17 |
A |
790 |
785 |
13.15 |
|
|
|
| 18 |
A |
608 |
605 |
9.71 |
|
|
|
| 19 |
A |
589 |
586 |
4.78 |
|
|
|
| 20 |
A |
457 |
455 |
10.02 |
|
|
|
| 21 |
A |
448 |
445 |
5.64 |
|
|
|
| 22 |
A |
266 |
265 |
6.79 |
|
|
|
| 23 |
A |
177 |
176 |
142.59 |
|
|
|
| 24 |
A |
124 |
123 |
36.56 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16763.5 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 16666.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.099 |
|
|
|
| 2 |
C |
0.103 |
|
|
|
| 3 |
N |
-0.073 |
|
|
|
| 4 |
O |
-0.235 |
|
|
|
| 5 |
C |
0.055 |
|
|
|
| 6 |
H |
-0.014 |
|
|
|
| 7 |
H |
0.099 |
|
|
|
| 8 |
H |
0.092 |
|
|
|
| 9 |
H |
0.041 |
|
|
|
| 10 |
H |
0.031 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.679 |
-3.300 |
0.352 |
3.387 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.098 |
-2.836 |
0.736 |
| y |
-2.836 |
-29.816 |
-0.378 |
| z |
0.736 |
-0.378 |
-31.366 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
8.493 |
-2.836 |
0.736 |
| y |
-2.836 |
-3.084 |
-0.378 |
| z |
0.736 |
-0.378 |
-5.409 |
|
| Polar |
|---|
| 3z2-r2 | -10.818 |
|---|
| x2-y2 | 7.718 |
|---|
| xy | -2.836 |
|---|
| xz | 0.736 |
|---|
| yz | -0.378 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.465 |
-0.828 |
0.008 |
| y |
-0.828 |
6.857 |
0.011 |
| z |
0.008 |
0.011 |
2.854 |
<r2> (average value of r
2) Å
2
| <r2> |
116.425 |
| (<r2>)1/2 |
10.790 |