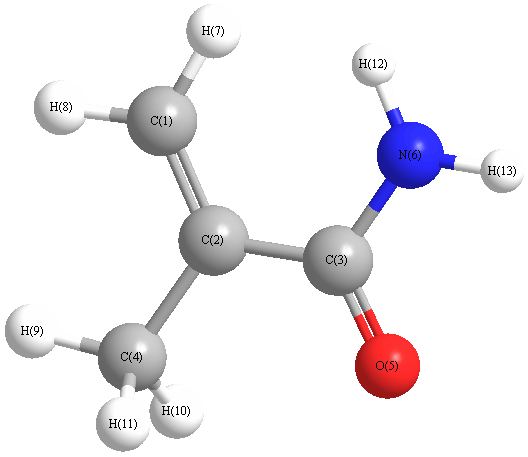
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3604 |
3583 |
24.03 |
|
|
|
| 2 |
A |
3468 |
3448 |
18.38 |
|
|
|
| 3 |
A |
3169 |
3151 |
10.39 |
|
|
|
| 4 |
A |
3084 |
3066 |
9.93 |
|
|
|
| 5 |
A |
3074 |
3056 |
5.40 |
|
|
|
| 6 |
A |
3045 |
3027 |
7.62 |
|
|
|
| 7 |
A |
2975 |
2958 |
14.57 |
|
|
|
| 8 |
A |
1721 |
1711 |
210.92 |
|
|
|
| 9 |
A |
1653 |
1643 |
34.03 |
|
|
|
| 10 |
A |
1527 |
1518 |
97.36 |
|
|
|
| 11 |
A |
1412 |
1404 |
23.04 |
|
|
|
| 12 |
A |
1385 |
1377 |
11.68 |
|
|
|
| 13 |
A |
1374 |
1366 |
14.28 |
|
|
|
| 14 |
A |
1353 |
1345 |
23.76 |
|
|
|
| 15 |
A |
1323 |
1315 |
34.99 |
|
|
|
| 16 |
A |
1197 |
1190 |
34.81 |
|
|
|
| 17 |
A |
1057 |
1051 |
3.74 |
|
|
|
| 18 |
A |
1016 |
1010 |
0.94 |
|
|
|
| 19 |
A |
972 |
966 |
4.84 |
|
|
|
| 20 |
A |
930 |
924 |
3.71 |
|
|
|
| 21 |
A |
897 |
892 |
27.87 |
|
|
|
| 22 |
A |
795 |
790 |
11.10 |
|
|
|
| 23 |
A |
733 |
729 |
6.45 |
|
|
|
| 24 |
A |
647 |
644 |
8.79 |
|
|
|
| 25 |
A |
560 |
557 |
21.49 |
|
|
|
| 26 |
A |
542 |
539 |
7.21 |
|
|
|
| 27 |
A |
494 |
491 |
4.26 |
|
|
|
| 28 |
A |
386 |
384 |
136.80 |
|
|
|
| 29 |
A |
361 |
359 |
20.07 |
|
|
|
| 30 |
A |
316 |
314 |
4.51 |
|
|
|
| 31 |
A |
260 |
258 |
10.60 |
|
|
|
| 32 |
A |
159 |
158 |
0.41 |
|
|
|
| 33 |
A |
73 |
72 |
4.40 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22780.3 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 22648.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.081 |
|
|
|
| 2 |
H |
0.099 |
|
|
|
| 3 |
H |
0.098 |
|
|
|
| 4 |
C |
0.019 |
|
|
|
| 5 |
H |
0.050 |
|
|
|
| 6 |
H |
0.064 |
|
|
|
| 7 |
H |
0.023 |
|
|
|
| 8 |
C |
0.033 |
|
|
|
| 9 |
H |
0.016 |
|
|
|
| 10 |
H |
0.022 |
|
|
|
| 11 |
C |
0.094 |
|
|
|
| 12 |
O |
-0.230 |
|
|
|
| 13 |
C |
-0.206 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.092 |
2.657 |
1.467 |
3.037 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.243 |
-4.137 |
-2.374 |
| y |
-4.137 |
-37.049 |
0.158 |
| z |
-2.374 |
0.158 |
-36.814 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.689 |
-4.137 |
-2.374 |
| y |
-4.137 |
-3.021 |
0.158 |
| z |
-2.374 |
0.158 |
-2.668 |
|
| Polar |
|---|
| 3z2-r2 | -5.336 |
|---|
| x2-y2 | 5.806 |
|---|
| xy | -4.137 |
|---|
| xz | -2.374 |
|---|
| yz | 0.158 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.295 |
0.267 |
-0.225 |
| y |
0.267 |
10.149 |
-0.197 |
| z |
-0.225 |
-0.197 |
4.800 |
<r2> (average value of r
2) Å
2
| <r2> |
165.317 |
| (<r2>)1/2 |
12.858 |