return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H4N2S2 (Ethanedithioamide)

using model chemistry: MP3=FULL/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1AG
Energy calculated at MP3=FULL/cc-pVDZ
 hartrees
Energy at 0K-982.957734
Energy at 298.15K-982.963781
HF Energy-982.046059
Nuclear repulsion energy336.716515
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3=FULL/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3768 3543 0.00      
2 Ag 3568 3355 0.00      
3 Ag 1671 1572 0.00      
4 Ag 1460 1373 0.00      
5 Ag 1376 1294 0.00      
6 Ag 990 931 0.00      
7 Ag 694 652 0.00      
8 Ag 419 394 0.00      
9 Ag 340 320 0.00      
10 Au 674 633 1.93      
11 Au 477 448 262.98      
12 Au 394 371 81.23      
13 Au 37 35 6.70      
14 Bg 720 677 0.00      
15 Bg 661 621 0.00      
16 Bg 448 421 0.00      
17 Bu 3770 3545 219.86      
18 Bu 3577 3364 272.45      
19 Bu 1625 1528 613.68      
20 Bu 1466 1379 319.42      
21 Bu 1259 1184 98.56      
22 Bu 908 854 60.97      
23 Bu 462 434 0.34      
24 Bu 285 268 27.41      

Unscaled Zero Point Vibrational Energy (zpe) 15523.9 cm-1
Scaled (by 0.9403) Zero Point Vibrational Energy (zpe) 14597.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3=FULL/cc-pVDZ
ABC
0.14972 0.05325 0.03928

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3=FULL/cc-pVDZ

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
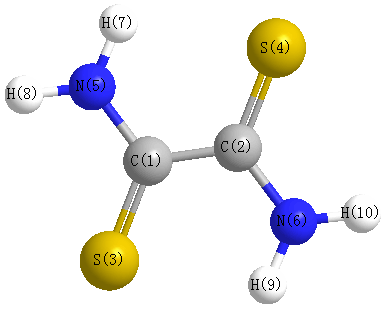
C1 -0.047 0.768 0.000
C2 0.047 -0.768 0.000
S3 1.301 1.744 0.000
S4 -1.301 -1.744 0.000
N5 -1.301 1.217 0.000
N6 1.301 -1.217 0.000
H7 -2.058 0.538 0.000
H8 -1.473 2.212 0.000
H9 2.058 -0.538 0.000
H10 1.473 -2.212 0.000

Atom - Atom Distances (Å)
  C1 C2 S3 S4 N5 N6 H7 H8 H9 H10
C11.53881.66432.80741.33232.39972.02412.02972.47683.3455
C21.53882.80741.66432.39971.33232.47683.34552.02412.0297
S31.66432.80744.35142.65522.96093.56912.81352.40343.9598
S42.80741.66434.35142.96092.65522.40343.95983.56912.8135
N51.33232.39972.65522.96093.56361.01711.00993.78984.4111
N62.39971.33232.96092.65523.56363.78984.41111.01711.0099
H72.02412.47683.56912.40341.01713.78981.77404.25384.4753
H82.02973.34552.81353.95981.00994.41111.77404.47535.3157
H92.47682.02412.40343.56913.78981.01714.25384.47531.7740
H103.34552.02973.95982.81354.41111.00994.47535.31571.7740

picture of Ethanedithioamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 S4 122.394 C1 C2 N6 113.206
C1 N5 H7 118.362 C1 N5 H8 119.494
C2 C1 S3 122.394 C2 C1 N5 113.206
C2 N6 H9 118.362 C2 N6 H10 119.494
S3 C1 N5 124.400 S4 C2 N6 124.400
H7 N5 H8 122.145 H9 N6 H10 122.145
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability