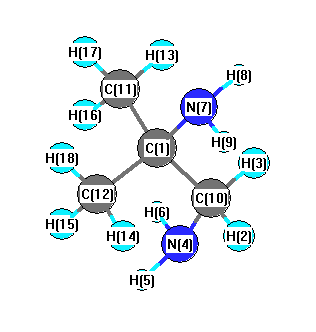
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3335 |
3340 |
3.00 |
|
|
|
| 2 |
A |
3300 |
3305 |
8.44 |
|
|
|
| 3 |
A |
3036 |
3041 |
45.51 |
|
|
|
| 4 |
A |
3010 |
3015 |
54.35 |
|
|
|
| 5 |
A |
2949 |
2954 |
1.65 |
|
|
|
| 6 |
A |
2915 |
2920 |
50.06 |
|
|
|
| 7 |
A |
1605 |
1607 |
9.31 |
|
|
|
| 8 |
A |
1596 |
1598 |
17.52 |
|
|
|
| 9 |
A |
1438 |
1441 |
5.84 |
|
|
|
| 10 |
A |
1431 |
1434 |
5.07 |
|
|
|
| 11 |
A |
1405 |
1407 |
0.67 |
|
|
|
| 12 |
A |
1357 |
1359 |
7.64 |
|
|
|
| 13 |
A |
1316 |
1318 |
9.17 |
|
|
|
| 14 |
A |
1201 |
1203 |
7.79 |
|
|
|
| 15 |
A |
1160 |
1162 |
22.90 |
|
|
|
| 16 |
A |
1061 |
1063 |
8.11 |
|
|
|
| 17 |
A |
985 |
987 |
9.10 |
|
|
|
| 18 |
A |
898 |
900 |
190.96 |
|
|
|
| 19 |
A |
862 |
863 |
8.64 |
|
|
|
| 20 |
A |
843 |
844 |
139.23 |
|
|
|
| 21 |
A |
704 |
705 |
6.31 |
|
|
|
| 22 |
A |
491 |
491 |
8.14 |
|
|
|
| 23 |
A |
410 |
411 |
7.20 |
|
|
|
| 24 |
A |
347 |
347 |
0.10 |
|
|
|
| 25 |
A |
255 |
256 |
2.59 |
|
|
|
| 26 |
A |
240 |
240 |
2.15 |
|
|
|
| 27 |
A |
3410 |
3416 |
1.25 |
|
|
|
| 28 |
A |
3373 |
3378 |
4.60 |
|
|
|
| 29 |
A |
3031 |
3036 |
0.62 |
|
|
|
| 30 |
A |
3008 |
3013 |
15.72 |
|
|
|
| 31 |
A |
2955 |
2959 |
51.75 |
|
|
|
| 32 |
A |
2943 |
2948 |
38.44 |
|
|
|
| 33 |
A |
1421 |
1423 |
0.08 |
|
|
|
| 34 |
A |
1413 |
1416 |
0.07 |
|
|
|
| 35 |
A |
1357 |
1359 |
0.23 |
|
|
|
| 36 |
A |
1337 |
1340 |
7.52 |
|
|
|
| 37 |
A |
1298 |
1300 |
0.21 |
|
|
|
| 38 |
A |
1151 |
1153 |
0.35 |
|
|
|
| 39 |
A |
1024 |
1026 |
1.60 |
|
|
|
| 40 |
A |
960 |
962 |
0.89 |
|
|
|
| 41 |
A |
906 |
907 |
0.13 |
|
|
|
| 42 |
A |
831 |
832 |
0.24 |
|
|
|
| 43 |
A |
442 |
442 |
0.98 |
|
|
|
| 44 |
A |
344 |
345 |
21.14 |
|
|
|
| 45 |
A |
311 |
312 |
0.69 |
|
|
|
| 46 |
A |
287 |
287 |
50.42 |
|
|
|
| 47 |
A |
202 |
203 |
0.01 |
|
|
|
| 48 |
A |
118 |
118 |
1.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35135.3 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 35191.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.209 |
|
|
|
| 2 |
H |
-0.019 |
|
|
|
| 3 |
H |
-0.019 |
|
|
|
| 4 |
N |
-0.132 |
|
|
|
| 5 |
H |
0.060 |
|
|
|
| 6 |
H |
0.060 |
|
|
|
| 7 |
N |
-0.102 |
|
|
|
| 8 |
H |
0.054 |
|
|
|
| 9 |
H |
0.054 |
|
|
|
| 10 |
C |
0.103 |
|
|
|
| 11 |
C |
0.097 |
|
|
|
| 12 |
C |
0.097 |
|
|
|
| 13 |
H |
-0.012 |
|
|
|
| 14 |
H |
-0.012 |
|
|
|
| 15 |
H |
-0.014 |
|
|
|
| 16 |
H |
-0.014 |
|
|
|
| 17 |
H |
0.005 |
|
|
|
| 18 |
H |
0.005 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.227 |
-0.078 |
0.000 |
0.240 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.026 |
4.006 |
0.000 |
| y |
4.006 |
-47.308 |
0.000 |
| z |
0.000 |
0.000 |
-38.188 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.722 |
4.006 |
0.000 |
| y |
4.006 |
-9.201 |
0.000 |
| z |
0.000 |
0.000 |
4.479 |
|
| Polar |
|---|
| 3z2-r2 | 8.959 |
|---|
| x2-y2 | 9.282 |
|---|
| xy | 4.006 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.397 |
0.283 |
0.000 |
| y |
0.283 |
9.469 |
0.000 |
| z |
0.000 |
0.000 |
9.526 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |