Jump to
S1C2
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -216.568524 |
| Energy at 298.15K | |
| HF Energy | -215.933203 |
| Nuclear repulsion energy | 115.790704 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3283 |
3150 |
5.73 |
|
|
|
| 2 |
A' |
3192 |
3063 |
6.66 |
|
|
|
| 3 |
A' |
3176 |
3047 |
8.59 |
|
|
|
| 4 |
A' |
3055 |
2931 |
43.60 |
|
|
|
| 5 |
A' |
1733 |
1662 |
2.07 |
|
|
|
| 6 |
A' |
1505 |
1444 |
1.31 |
|
|
|
| 7 |
A' |
1455 |
1396 |
13.05 |
|
|
|
| 8 |
A' |
1430 |
1372 |
12.33 |
|
|
|
| 9 |
A' |
1310 |
1257 |
0.35 |
|
|
|
| 10 |
A' |
1155 |
1108 |
52.06 |
|
|
|
| 11 |
A' |
1019 |
978 |
27.62 |
|
|
|
| 12 |
A' |
927 |
890 |
2.73 |
|
|
|
| 13 |
A' |
609 |
585 |
5.97 |
|
|
|
| 14 |
A' |
276 |
265 |
2.65 |
|
|
|
| 15 |
A" |
3100 |
2974 |
40.37 |
|
|
|
| 16 |
A" |
1273 |
1222 |
0.24 |
|
|
|
| 17 |
A" |
1050 |
1008 |
14.18 |
|
|
|
| 18 |
A" |
1013 |
972 |
11.65 |
|
|
|
| 19 |
A" |
943 |
905 |
29.59 |
|
|
|
| 20 |
A" |
558 |
535 |
7.70 |
|
|
|
| 21 |
A" |
175 |
167 |
2.79 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16118.3 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 15463.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.953 |
-0.205 |
0.000 |
| C2 |
0.000 |
0.961 |
0.000 |
| C3 |
1.339 |
0.835 |
0.000 |
| F4 |
-0.271 |
-1.413 |
0.000 |
| H5 |
1.991 |
1.716 |
0.000 |
| H6 |
1.811 |
-0.154 |
0.000 |
| H7 |
-0.472 |
1.955 |
0.000 |
| H8 |
-1.604 |
-0.174 |
0.896 |
| H9 |
-1.604 |
-0.174 |
-0.896 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5064 | 2.5171 | 1.3867 | 3.5149 | 2.7643 | 2.2127 | 1.1074 | 1.1074 |
C2 | 1.5064 | | 1.3453 | 2.3894 | 2.1287 | 2.1266 | 1.0995 | 2.1594 | 2.1594 | C3 | 2.5171 | 1.3453 | | 2.7645 | 1.0956 | 1.0951 | 2.1294 | 3.2373 | 3.2373 | F4 | 1.3867 | 2.3894 | 2.7645 | | 3.8600 | 2.4327 | 3.3730 | 2.0279 | 2.0279 | H5 | 3.5149 | 2.1287 | 1.0956 | 3.8600 | | 1.8780 | 2.4740 | 4.1583 | 4.1583 | H6 | 2.7643 | 2.1266 | 1.0951 | 2.4327 | 1.8780 | | 3.1073 | 3.5300 | 3.5300 | H7 | 2.2127 | 1.0995 | 2.1294 | 3.3730 | 2.4740 | 3.1073 | | 2.5716 | 2.5716 | H8 | 1.1074 | 2.1594 | 3.2373 | 2.0279 | 4.1583 | 3.5300 | 2.5716 | | 1.7910 | H9 | 1.1074 | 2.1594 | 3.2373 | 2.0279 | 4.1583 | 3.5300 | 2.5716 | 1.7910 | |
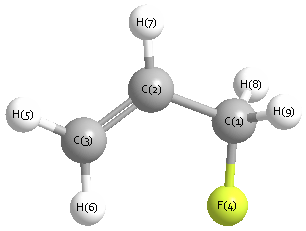 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.839 |
|
C1 |
C2 |
H7 |
115.339 |
| C2 |
C1 |
F4 |
111.289 |
|
C2 |
C1 |
H8 |
110.481 |
| C2 |
C1 |
H9 |
110.481 |
|
C2 |
C3 |
H5 |
121.068 |
| C2 |
C3 |
H6 |
120.908 |
|
C3 |
C2 |
H7 |
120.822 |
| F4 |
C1 |
H8 |
108.276 |
|
F4 |
C1 |
H9 |
108.276 |
| H5 |
C3 |
H6 |
118.024 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -216.566963 |
| Energy at 298.15K | |
| HF Energy | -215.932009 |
| Nuclear repulsion energy | 113.367559 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3264 |
3131 |
12.19 |
|
|
|
| 2 |
A |
3192 |
3063 |
6.02 |
|
|
|
| 3 |
A |
3166 |
3037 |
8.93 |
|
|
|
| 4 |
A |
3120 |
2993 |
33.50 |
|
|
|
| 5 |
A |
3064 |
2940 |
40.73 |
|
|
|
| 6 |
A |
1730 |
1660 |
0.37 |
|
|
|
| 7 |
A |
1508 |
1447 |
1.02 |
|
|
|
| 8 |
A |
1469 |
1410 |
22.70 |
|
|
|
| 9 |
A |
1404 |
1347 |
18.83 |
|
|
|
| 10 |
A |
1308 |
1255 |
0.12 |
|
|
|
| 11 |
A |
1276 |
1224 |
3.19 |
|
|
|
| 12 |
A |
1188 |
1139 |
3.54 |
|
|
|
| 13 |
A |
1090 |
1046 |
105.14 |
|
|
|
| 14 |
A |
1017 |
976 |
29.83 |
|
|
|
| 15 |
A |
991 |
951 |
4.42 |
|
|
|
| 16 |
A |
949 |
911 |
28.99 |
|
|
|
| 17 |
A |
931 |
893 |
1.93 |
|
|
|
| 18 |
A |
647 |
621 |
6.41 |
|
|
|
| 19 |
A |
434 |
416 |
2.07 |
|
|
|
| 20 |
A |
328 |
314 |
6.03 |
|
|
|
| 21 |
A |
111 |
107 |
1.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16092.0 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 15438.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.615 |
0.436 |
0.309 |
| C2 |
0.644 |
-0.386 |
0.239 |
| C3 |
1.820 |
0.093 |
-0.208 |
| F4 |
-1.656 |
-0.227 |
-0.337 |
| H5 |
2.726 |
-0.526 |
-0.214 |
| H6 |
1.915 |
1.122 |
-0.580 |
| H7 |
0.563 |
-1.420 |
0.600 |
| H8 |
-0.924 |
0.594 |
1.359 |
| H9 |
-0.471 |
1.421 |
-0.172 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5052 | 2.5135 | 1.3927 | 3.5155 | 2.7682 | 2.2179 | 1.1066 | 1.1054 |
C2 | 1.5052 | | 1.3464 | 2.3766 | 2.1349 | 2.1349 | 1.0989 | 2.1616 | 2.1622 | C3 | 2.5135 | 1.3464 | | 3.4935 | 1.0960 | 1.0980 | 2.1269 | 3.2001 | 2.6487 | F4 | 1.3927 | 2.3766 | 3.4935 | | 4.3934 | 3.8253 | 2.6880 | 2.0221 | 2.0367 | H5 | 3.5155 | 2.1349 | 1.0960 | 4.3934 | | 1.8715 | 2.4781 | 4.1289 | 3.7425 | H6 | 2.7682 | 2.1349 | 1.0980 | 3.8253 | 1.8715 | | 3.1115 | 3.4783 | 2.4391 | H7 | 2.2179 | 1.0989 | 2.1269 | 2.6880 | 2.4781 | 3.1115 | | 2.6164 | 3.1205 | H8 | 1.1066 | 2.1616 | 3.2001 | 2.0221 | 4.1289 | 3.4783 | 2.6164 | | 1.7985 | H9 | 1.1054 | 2.1622 | 2.6487 | 2.0367 | 3.7425 | 2.4391 | 3.1205 | 1.7985 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.539 |
|
C1 |
C2 |
H7 |
115.917 |
| C2 |
C1 |
F4 |
110.126 |
|
C2 |
C1 |
H8 |
110.787 |
| C2 |
C1 |
H9 |
110.910 |
|
C2 |
C3 |
H5 |
121.539 |
| C2 |
C3 |
H6 |
121.380 |
|
C3 |
C2 |
H7 |
120.533 |
| F4 |
C1 |
H8 |
107.452 |
|
F4 |
C1 |
H9 |
108.684 |
| H5 |
C3 |
H6 |
117.081 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
