Jump to
S1C2
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -216.562869 |
| Energy at 298.15K | |
| HF Energy | -215.933503 |
| Nuclear repulsion energy | 115.924512 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3292 |
3150 |
5.39 |
|
|
|
| 2 |
A' |
3201 |
3062 |
6.40 |
|
|
|
| 3 |
A' |
3184 |
3046 |
7.89 |
|
|
|
| 4 |
A' |
3065 |
2932 |
41.69 |
|
|
|
| 5 |
A' |
1747 |
1671 |
1.81 |
|
|
|
| 6 |
A' |
1509 |
1444 |
1.84 |
|
|
|
| 7 |
A' |
1461 |
1398 |
14.14 |
|
|
|
| 8 |
A' |
1436 |
1374 |
11.33 |
|
|
|
| 9 |
A' |
1316 |
1259 |
0.33 |
|
|
|
| 10 |
A' |
1170 |
1119 |
55.55 |
|
|
|
| 11 |
A' |
1027 |
983 |
24.67 |
|
|
|
| 12 |
A' |
932 |
892 |
2.44 |
|
|
|
| 13 |
A' |
613 |
587 |
6.09 |
|
|
|
| 14 |
A' |
278 |
265 |
2.72 |
|
|
|
| 15 |
A" |
3111 |
2976 |
38.97 |
|
|
|
| 16 |
A" |
1281 |
1225 |
0.33 |
|
|
|
| 17 |
A" |
1055 |
1009 |
14.57 |
|
|
|
| 18 |
A" |
1017 |
973 |
10.72 |
|
|
|
| 19 |
A" |
952 |
911 |
31.16 |
|
|
|
| 20 |
A" |
560 |
536 |
7.72 |
|
|
|
| 21 |
A" |
175 |
167 |
2.87 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16190.7 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 15489.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.951 |
-0.207 |
0.000 |
| C2 |
0.000 |
0.960 |
0.000 |
| C3 |
1.337 |
0.836 |
0.000 |
| F4 |
-0.270 |
-1.411 |
0.000 |
| H5 |
1.986 |
1.719 |
0.000 |
| H6 |
1.812 |
-0.150 |
0.000 |
| H7 |
-0.473 |
1.952 |
0.000 |
| H8 |
-1.602 |
-0.176 |
0.895 |
| H9 |
-1.602 |
-0.176 |
-0.895 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5060 | 2.5152 | 1.3830 | 3.5125 | 2.7635 | 2.2118 | 1.1069 | 1.1069 |
C2 | 1.5060 | | 1.3428 | 2.3866 | 2.1259 | 2.1247 | 1.0991 | 2.1584 | 2.1584 | C3 | 2.5152 | 1.3428 | | 2.7632 | 1.0952 | 1.0947 | 2.1265 | 3.2350 | 3.2350 | F4 | 1.3830 | 2.3866 | 2.7632 | | 3.8584 | 2.4341 | 3.3693 | 2.0246 | 2.0246 | H5 | 3.5125 | 2.1259 | 1.0952 | 3.8584 | | 1.8770 | 2.4703 | 4.1552 | 4.1552 | H6 | 2.7635 | 2.1247 | 1.0947 | 2.4341 | 1.8770 | | 3.1048 | 3.5291 | 3.5291 | H7 | 2.2118 | 1.0991 | 2.1265 | 3.3693 | 2.4703 | 3.1048 | | 2.5700 | 2.5700 | H8 | 1.1069 | 2.1584 | 3.2350 | 2.0246 | 4.1552 | 3.5291 | 2.5700 | | 1.7898 | H9 | 1.1069 | 2.1584 | 3.2350 | 2.0246 | 4.1552 | 3.5291 | 2.5700 | 1.7898 | |
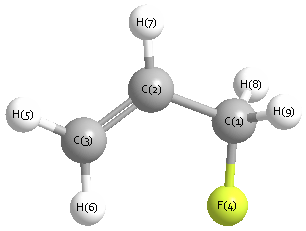 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.892 |
|
C1 |
C2 |
H7 |
115.319 |
| C2 |
C1 |
F4 |
111.328 |
|
C2 |
C1 |
H8 |
110.461 |
| C2 |
C1 |
H9 |
110.461 |
|
C2 |
C3 |
H5 |
121.047 |
| C2 |
C3 |
H6 |
120.969 |
|
C3 |
C2 |
H7 |
120.789 |
| F4 |
C1 |
H8 |
108.295 |
|
F4 |
C1 |
H9 |
108.295 |
| H5 |
C3 |
H6 |
117.983 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -216.561375 |
| Energy at 298.15K | |
| HF Energy | -215.932312 |
| Nuclear repulsion energy | 113.513779 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3273 |
3131 |
11.52 |
|
|
|
| 2 |
A |
3200 |
3062 |
5.74 |
|
|
|
| 3 |
A |
3175 |
3037 |
8.39 |
|
|
|
| 4 |
A |
3130 |
2994 |
32.77 |
|
|
|
| 5 |
A |
3073 |
2940 |
39.10 |
|
|
|
| 6 |
A |
1745 |
1669 |
0.37 |
|
|
|
| 7 |
A |
1512 |
1446 |
0.78 |
|
|
|
| 8 |
A |
1475 |
1411 |
24.19 |
|
|
|
| 9 |
A |
1411 |
1349 |
17.55 |
|
|
|
| 10 |
A |
1313 |
1256 |
0.13 |
|
|
|
| 11 |
A |
1283 |
1228 |
3.17 |
|
|
|
| 12 |
A |
1191 |
1140 |
3.79 |
|
|
|
| 13 |
A |
1113 |
1065 |
105.45 |
|
|
|
| 14 |
A |
1023 |
978 |
27.31 |
|
|
|
| 15 |
A |
995 |
952 |
3.74 |
|
|
|
| 16 |
A |
958 |
917 |
31.09 |
|
|
|
| 17 |
A |
934 |
893 |
1.81 |
|
|
|
| 18 |
A |
650 |
622 |
6.49 |
|
|
|
| 19 |
A |
436 |
418 |
2.02 |
|
|
|
| 20 |
A |
330 |
315 |
5.95 |
|
|
|
| 21 |
A |
111 |
107 |
1.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16164.7 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 15464.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.616 |
0.434 |
0.307 |
| C2 |
0.644 |
-0.386 |
0.237 |
| C3 |
1.819 |
0.094 |
-0.207 |
| F4 |
-1.655 |
-0.226 |
-0.335 |
| H5 |
2.724 |
-0.523 |
-0.213 |
| H6 |
1.914 |
1.123 |
-0.574 |
| H7 |
0.564 |
-1.422 |
0.594 |
| H8 |
-0.922 |
0.591 |
1.359 |
| H9 |
-0.472 |
1.419 |
-0.171 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5051 | 2.5114 | 1.3883 | 3.5132 | 2.7662 | 2.2174 | 1.1062 | 1.1050 |
C2 | 1.5051 | | 1.3439 | 2.3745 | 2.1324 | 2.1325 | 1.0986 | 2.1605 | 2.1612 | C3 | 2.5114 | 1.3439 | | 3.4903 | 1.0956 | 1.0976 | 2.1243 | 3.1956 | 2.6465 | F4 | 1.3883 | 2.3745 | 3.4903 | | 4.3904 | 3.8223 | 2.6860 | 2.0184 | 2.0331 | H5 | 3.5132 | 2.1324 | 1.0956 | 4.3904 | | 1.8705 | 2.4752 | 4.1244 | 3.7400 | H6 | 2.7662 | 2.1325 | 1.0976 | 3.8223 | 1.8705 | | 3.1090 | 3.4732 | 2.4372 | H7 | 2.2174 | 1.0986 | 2.1243 | 2.6860 | 2.4752 | 3.1090 | | 2.6164 | 3.1191 | H8 | 1.1062 | 2.1605 | 3.1956 | 2.0184 | 4.1244 | 3.4732 | 2.6164 | | 1.7969 | H9 | 1.1050 | 2.1612 | 2.6465 | 2.0331 | 3.7400 | 2.4372 | 3.1191 | 1.7969 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.559 |
|
C1 |
C2 |
H7 |
115.902 |
| C2 |
C1 |
F4 |
110.239 |
|
C2 |
C1 |
H8 |
110.731 |
| C2 |
C1 |
H9 |
110.864 |
|
C2 |
C3 |
H5 |
121.547 |
| C2 |
C3 |
H6 |
121.403 |
|
C3 |
C2 |
H7 |
120.529 |
| F4 |
C1 |
H8 |
107.482 |
|
F4 |
C1 |
H9 |
108.723 |
| H5 |
C3 |
H6 |
117.049 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
