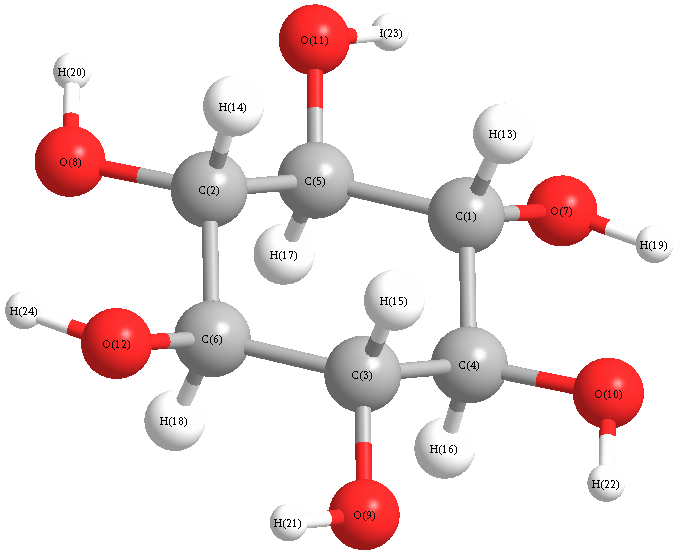
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
3642 |
3621 |
0.00 |
|
|
|
| 2 |
A1g |
2908 |
2891 |
0.00 |
|
|
|
| 3 |
A1g |
1389 |
1381 |
0.00 |
|
|
|
| 4 |
A1g |
1304 |
1296 |
0.00 |
|
|
|
| 5 |
A1g |
1156 |
1149 |
0.00 |
|
|
|
| 6 |
A1g |
977 |
971 |
0.00 |
|
|
|
| 7 |
A1g |
434 |
432 |
0.00 |
|
|
|
| 8 |
A1g |
120 |
119 |
0.00 |
|
|
|
| 9 |
A1u |
1264 |
1256 |
0.00 |
|
|
|
| 10 |
A1u |
987 |
981 |
0.00 |
|
|
|
| 11 |
A1u |
308 |
306 |
0.00 |
|
|
|
| 12 |
A1u |
122 |
121 |
0.00 |
|
|
|
| 13 |
A2g |
1117 |
1110 |
0.00 |
|
|
|
| 14 |
A2g |
623 |
620 |
0.00 |
|
|
|
| 15 |
A2g |
167 |
166 |
0.00 |
|
|
|
| 16 |
A2u |
3640 |
3619 |
56.81 |
|
|
|
| 17 |
A2u |
2937 |
2920 |
190.87 |
|
|
|
| 18 |
A2u |
1396 |
1388 |
28.97 |
|
|
|
| 19 |
A2u |
1244 |
1237 |
0.46 |
|
|
|
| 20 |
A2u |
1099 |
1092 |
22.29 |
|
|
|
| 21 |
A2u |
560 |
556 |
0.42 |
|
|
|
| 22 |
A2u |
226 |
225 |
17.36 |
|
|
|
| 23 |
Eg |
3640 |
3619 |
0.00 |
|
|
|
| 23 |
Eg |
3640 |
3619 |
0.00 |
|
|
|
| 24 |
Eg |
2913 |
2896 |
0.00 |
|
|
|
| 24 |
Eg |
2913 |
2896 |
0.00 |
|
|
|
| 25 |
Eg |
1380 |
1372 |
0.00 |
|
|
|
| 25 |
Eg |
1380 |
1372 |
0.00 |
|
|
|
| 26 |
Eg |
1319 |
1312 |
0.00 |
|
|
|
| 26 |
Eg |
1319 |
1312 |
0.00 |
|
|
|
| 27 |
Eg |
1172 |
1165 |
0.00 |
|
|
|
| 27 |
Eg |
1172 |
1165 |
0.00 |
|
|
|
| 28 |
Eg |
1106 |
1099 |
0.00 |
|
|
|
| 28 |
Eg |
1106 |
1099 |
0.00 |
|
|
|
| 29 |
Eg |
987 |
981 |
0.00 |
|
|
|
| 29 |
Eg |
987 |
981 |
0.00 |
|
|
|
| 30 |
Eg |
408 |
405 |
0.00 |
|
|
|
| 30 |
Eg |
408 |
405 |
0.00 |
|
|
|
| 31 |
Eg |
366 |
364 |
0.00 |
|
|
|
| 31 |
Eg |
366 |
364 |
0.00 |
|
|
|
| 32 |
Eg |
287 |
285 |
0.00 |
|
|
|
| 32 |
Eg |
287 |
285 |
0.00 |
|
|
|
| 33 |
Eg |
135 |
134 |
0.00 |
|
|
|
| 33 |
Eg |
135 |
134 |
0.00 |
|
|
|
| 34 |
Eu |
3641 |
3620 |
14.37 |
|
|
|
| 34 |
Eu |
3641 |
3620 |
14.37 |
|
|
|
| 35 |
Eu |
2901 |
2884 |
0.68 |
|
|
|
| 35 |
Eu |
2901 |
2884 |
0.68 |
|
|
|
| 36 |
Eu |
1374 |
1366 |
148.84 |
|
|
|
| 36 |
Eu |
1374 |
1366 |
148.85 |
|
|
|
| 37 |
Eu |
1315 |
1307 |
23.27 |
|
|
|
| 37 |
Eu |
1315 |
1307 |
23.26 |
|
|
|
| 38 |
Eu |
1154 |
1147 |
124.46 |
|
|
|
| 38 |
Eu |
1154 |
1147 |
124.46 |
|
|
|
| 39 |
Eu |
1115 |
1108 |
72.81 |
|
|
|
| 39 |
Eu |
1115 |
1108 |
72.81 |
|
|
|
| 40 |
Eu |
960 |
955 |
217.92 |
|
|
|
| 40 |
Eu |
960 |
955 |
217.92 |
|
|
|
| 41 |
Eu |
618 |
614 |
7.78 |
|
|
|
| 41 |
Eu |
618 |
614 |
7.78 |
|
|
|
| 42 |
Eu |
309 |
307 |
114.91 |
|
|
|
| 42 |
Eu |
309 |
307 |
114.91 |
|
|
|
| 43 |
Eu |
207 |
206 |
197.18 |
|
|
|
| 43 |
Eu |
207 |
206 |
197.18 |
|
|
|
| 44 |
Eu |
74 |
73 |
4.14 |
|
|
|
| 44 |
Eu |
74 |
73 |
4.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41186.4 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 40947.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.090 |
|
|
|
| 2 |
C |
0.090 |
|
|
|
| 3 |
C |
0.090 |
|
|
|
| 4 |
C |
0.090 |
|
|
|
| 5 |
C |
0.090 |
|
|
|
| 6 |
C |
0.090 |
|
|
|
| 7 |
O |
-0.216 |
|
|
|
| 8 |
O |
-0.216 |
|
|
|
| 9 |
O |
-0.216 |
|
|
|
| 10 |
O |
-0.216 |
|
|
|
| 11 |
O |
-0.216 |
|
|
|
| 12 |
O |
-0.216 |
|
|
|
| 13 |
H |
-0.011 |
|
|
|
| 14 |
H |
-0.011 |
|
|
|
| 15 |
H |
-0.011 |
|
|
|
| 16 |
H |
-0.011 |
|
|
|
| 17 |
H |
-0.011 |
|
|
|
| 18 |
H |
-0.011 |
|
|
|
| 19 |
H |
0.136 |
|
|
|
| 20 |
H |
0.136 |
|
|
|
| 21 |
H |
0.136 |
|
|
|
| 22 |
H |
0.136 |
|
|
|
| 23 |
H |
0.136 |
|
|
|
| 24 |
H |
0.136 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-81.505 |
0.000 |
0.000 |
| y |
0.000 |
-81.505 |
0.000 |
| z |
0.000 |
0.000 |
-57.253 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-12.126 |
0.000 |
0.000 |
| y |
0.000 |
-12.126 |
0.000 |
| z |
0.000 |
0.000 |
24.252 |
|
| Polar |
|---|
| 3z2-r2 | 48.504 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.313 |
0.000 |
0.000 |
| y |
0.000 |
13.313 |
0.000 |
| z |
0.000 |
0.000 |
12.310 |
<r2> (average value of r
2) Å
2
| <r2> |
596.831 |
| (<r2>)1/2 |
24.430 |