return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4N2 (2H-Imidazole)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-225.473732
Energy at 298.15K-225.479743
HF Energy-224.740693
Nuclear repulsion energy165.896376
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3273 3132 10.31      
2 A1 3117 2982 0.82      
3 A1 1761 1684 20.98      
4 A1 1452 1390 0.38      
5 A1 1412 1351 17.13      
6 A1 1285 1229 3.11      
7 A1 1050 1005 1.13      
8 A1 943 902 15.62      
9 A2 1164 1114 0.00      
10 A2 910 870 0.00      
11 A2 538 515 0.00      
12 B1 3172 3035 0.20      
13 B1 1026 982 22.29      
14 B1 828 793 5.77      
15 B1 346 331 32.77      
16 B2 3257 3116 6.05      
17 B2 1819 1740 0.04      
18 B2 1446 1383 41.55      
19 B2 1257 1203 2.08      
20 B2 1101 1053 26.67      
21 B2 945 904 64.23      

Unscaled Zero Point Vibrational Energy (zpe) 16051.1 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 15356.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.35961 0.30033 0.16905

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.211
N2 0.000 0.997 0.282
N3 0.000 -0.997 0.282
C4 0.000 0.733 -0.952
C5 0.000 -0.733 -0.952
H6 -0.903 0.000 1.842
H7 0.903 0.000 1.842
H8 0.000 1.491 -1.739
H9 0.000 -1.491 -1.739

Atom - Atom Distances (Å)
  C1 N2 N3 C4 C5 H6 H7 H8 H9
C11.36261.36262.28312.28311.10191.10193.30483.3048
N21.36261.99491.26192.12522.06012.06012.08053.2056
N31.36261.99492.12521.26192.06012.06013.20562.0805
C42.28311.26192.12521.46593.02633.02631.09272.3589
C52.28312.12521.26191.46593.02633.02632.35891.0927
H61.10192.06012.06013.02633.02631.80573.98283.9828
H71.10192.06012.06013.02633.02631.80573.98283.9828
H83.30482.08053.20561.09272.35893.98283.98282.9814
H93.30483.20562.08052.35891.09273.98283.98282.9814

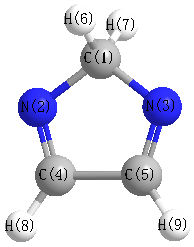
picture of 2H-Imidazole state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N2 C4 120.847 C1 N3 C5 120.847
N2 C1 N3 94.108 N2 C1 H6 112.991
N2 C1 H7 112.991 N2 C4 C5 102.098
N2 C4 H8 123.996 N3 C1 H6 112.991
N3 C1 H7 112.991 N3 C5 C4 102.098
N3 C5 H9 123.996 C4 C5 H9 133.906
C5 C4 H8 133.906 H6 C1 H7 110.040
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability