Jump to
S1C2
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Geometric Data calculated at QCISD(T)/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -115.617461 |
| Energy at 298.15K | -115.619138 |
| HF Energy | -115.200740 |
| Nuclear repulsion energy | 57.030244 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3303 |
3148 |
|
|
|
|
| 2 |
A' |
3085 |
2940 |
|
|
|
|
| 3 |
A' |
1720 |
1640 |
|
|
|
|
| 4 |
A' |
1221 |
1163 |
|
|
|
|
| 5 |
A' |
998 |
951 |
|
|
|
|
| 6 |
A' |
914 |
871 |
|
|
|
|
| 7 |
A' |
579 |
552 |
|
|
|
|
| 8 |
A" |
3269 |
3115 |
|
|
|
|
| 9 |
A" |
1157 |
1103 |
|
|
|
|
| 10 |
A" |
1070 |
1020 |
|
|
|
|
| 11 |
A" |
945 |
900 |
|
|
|
|
| 12 |
A" |
831 |
792 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9545.6 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 9097.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.039 |
0.904 |
0.000 |
| H2 |
0.759 |
1.670 |
0.000 |
| C3 |
-0.039 |
-0.425 |
0.669 |
| C4 |
-0.039 |
-0.425 |
-0.669 |
| H5 |
-0.032 |
-0.998 |
1.598 |
| H6 |
-0.032 |
-0.998 |
-1.598 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
| C1 | | 1.1058 | 1.4872 | 1.4872 | 2.4839 | 2.4839 |
H2 | 1.1058 | | 2.3389 | 2.3389 | 3.2089 | 3.2089 | C3 | 1.4872 | 2.3389 | | 1.3377 | 1.0918 | 2.3383 | C4 | 1.4872 | 2.3389 | 1.3377 | | 2.3383 | 1.0918 | H5 | 2.4839 | 3.2089 | 1.0918 | 2.3383 | | 3.1962 | H6 | 2.4839 | 3.2089 | 2.3383 | 1.0918 | 3.1962 | |
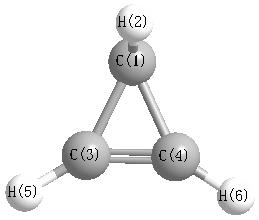 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C4 |
63.275 |
|
C1 |
C3 |
H5 |
148.389 |
| C1 |
C4 |
C3 |
63.275 |
|
C1 |
C4 |
H6 |
148.389 |
| H2 |
C1 |
C3 |
128.245 |
|
H2 |
C1 |
C4 |
128.245 |
| C3 |
C1 |
C4 |
53.451 |
|
C3 |
C4 |
H6 |
148.333 |
| C4 |
C3 |
H5 |
148.333 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
