Jump to
S1C2
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Geometric Data calculated at QCISD/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -115.601597 |
| Energy at 298.15K | -115.603149 |
| HF Energy | -115.201651 |
| Nuclear repulsion energy | 57.220094 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3319 |
3184 |
0.36 |
|
|
|
| 2 |
A' |
3106 |
2980 |
53.40 |
|
|
|
| 3 |
A' |
1656 |
1588 |
0.68 |
|
|
|
| 4 |
A' |
1242 |
1192 |
8.35 |
|
|
|
| 5 |
A' |
1009 |
969 |
16.17 |
|
|
|
| 6 |
A' |
923 |
886 |
4.13 |
|
|
|
| 7 |
A' |
601 |
577 |
62.11 |
|
|
|
| 8 |
A" |
3269 |
3136 |
9.83 |
|
|
|
| 9 |
A" |
1041 |
999 |
18.33 |
|
|
|
| 10 |
A" |
984 |
944 |
2.95 |
|
|
|
| 11 |
A" |
853 |
818 |
0.19 |
|
|
|
| 12 |
A" |
615 |
590 |
68.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9309.6 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 8931.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.038 |
0.901 |
0.000 |
| H2 |
0.757 |
1.667 |
0.000 |
| C3 |
-0.038 |
-0.424 |
0.666 |
| C4 |
-0.038 |
-0.424 |
-0.666 |
| H5 |
-0.038 |
-0.994 |
1.594 |
| H6 |
-0.038 |
-0.994 |
-1.594 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
| C1 | | 1.1039 | 1.4826 | 1.4826 | 2.4767 | 2.4767 |
H2 | 1.1039 | | 2.3335 | 2.3335 | 3.2023 | 3.2023 | C3 | 1.4826 | 2.3335 | | 1.3310 | 1.0899 | 2.3306 | C4 | 1.4826 | 2.3335 | 1.3310 | | 2.3306 | 1.0899 | H5 | 2.4767 | 3.2023 | 1.0899 | 2.3306 | | 3.1882 | H6 | 2.4767 | 3.2023 | 2.3306 | 1.0899 | 3.1882 | |
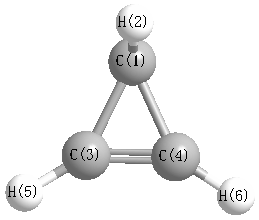 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C4 |
63.328 |
|
C1 |
C3 |
H5 |
148.246 |
| C1 |
C4 |
C3 |
63.328 |
|
C1 |
C4 |
H6 |
148.246 |
| H2 |
C1 |
C3 |
128.295 |
|
H2 |
C1 |
C4 |
128.295 |
| C3 |
C1 |
C4 |
53.345 |
|
C3 |
C4 |
H6 |
148.426 |
| C4 |
C3 |
H5 |
148.426 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
