Jump to
S1C2
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -115.803794 |
| Energy at 298.15K | -115.805305 |
| HF Energy | -115.803794 |
| Nuclear repulsion energy | 57.322216 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3216 |
3197 |
0.02 |
|
|
|
| 2 |
A' |
2977 |
2959 |
61.57 |
|
|
|
| 3 |
A' |
1631 |
1621 |
0.24 |
|
|
|
| 4 |
A' |
1223 |
1216 |
9.76 |
|
|
|
| 5 |
A' |
945 |
940 |
22.38 |
|
|
|
| 6 |
A' |
874 |
869 |
5.79 |
|
|
|
| 7 |
A' |
586 |
583 |
51.12 |
|
|
|
| 8 |
A" |
3168 |
3149 |
0.90 |
|
|
|
| 9 |
A" |
979 |
973 |
27.15 |
|
|
|
| 10 |
A" |
925 |
920 |
1.98 |
|
|
|
| 11 |
A" |
836 |
832 |
0.30 |
|
|
|
| 12 |
A" |
660 |
656 |
30.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9009.3 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 8957.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.038 |
0.896 |
0.000 |
| H2 |
0.759 |
1.670 |
0.000 |
| C3 |
-0.038 |
-0.421 |
0.665 |
| C4 |
-0.038 |
-0.421 |
-0.665 |
| H5 |
-0.038 |
-0.996 |
1.596 |
| H6 |
-0.038 |
-0.996 |
-1.596 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
| C1 | | 1.1106 | 1.4752 | 1.4752 | 2.4752 | 2.4752 |
H2 | 1.1106 | | 2.3341 | 2.3341 | 3.2077 | 3.2077 | C3 | 1.4752 | 2.3341 | | 1.3293 | 1.0947 | 2.3329 | C4 | 1.4752 | 2.3341 | 1.3293 | | 2.3329 | 1.0947 | H5 | 2.4752 | 3.2077 | 1.0947 | 2.3329 | | 3.1928 | H6 | 2.4752 | 3.2077 | 2.3329 | 1.0947 | 3.1928 | |
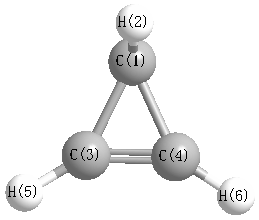 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C4 |
63.219 |
|
C1 |
C3 |
H5 |
148.447 |
| C1 |
C4 |
C3 |
63.219 |
|
C1 |
C4 |
H6 |
148.447 |
| H2 |
C1 |
C3 |
128.470 |
|
H2 |
C1 |
C4 |
128.470 |
| C3 |
C1 |
C4 |
53.561 |
|
C3 |
C4 |
H6 |
148.333 |
| C4 |
C3 |
H5 |
148.333 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.094 |
|
|
|
| 2 |
H |
0.035 |
|
|
|
| 3 |
C |
0.023 |
|
|
|
| 4 |
C |
0.023 |
|
|
|
| 5 |
H |
0.007 |
|
|
|
| 6 |
H |
0.007 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.614 |
-1.340 |
0.000 |
1.474 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.019 |
1.163 |
0.000 |
| y |
1.163 |
-18.288 |
0.000 |
| z |
0.000 |
0.000 |
-15.672 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.040 |
1.163 |
0.000 |
| y |
1.163 |
-0.942 |
0.000 |
| z |
0.000 |
0.000 |
2.982 |
|
| Polar |
|---|
| 3z2-r2 | 5.964 |
|---|
| x2-y2 | -0.732 |
|---|
| xy | 1.163 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.550 |
0.373 |
0.000 |
| y |
0.373 |
4.969 |
0.000 |
| z |
0.000 |
0.000 |
5.017 |
<r2> (average value of r
2) Å
2
| <r2> |
33.747 |
| (<r2>)1/2 |
5.809 |
