return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for HOCHCCHOH (allenediol)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-266.379675
Energy at 298.15K-266.383817
HF Energy-265.593454
Nuclear repulsion energy151.604231
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3861 3693 33.52      
2 A 3218 3079 9.32      
3 A 1561 1493 35.69      
4 A 1345 1287 0.20      
5 A 1240 1186 27.20      
6 A 969 927 83.76      
7 A 939 899 7.29      
8 A 530 507 75.20      
9 A 499 478 43.98      
10 A 341 326 18.37      
11 A 126 120 0.01      
12 B 3860 3693 42.76      
13 B 3219 3079 13.51      
14 B 2086 1996 295.23      
15 B 1429 1367 87.49      
16 B 1300 1244 5.24      
17 B 1154 1104 469.34      
18 B 871 833 10.47      
19 B 630 603 46.57      
20 B 479 459 107.69      
21 B 155 148 0.76      

Unscaled Zero Point Vibrational Energy (zpe) 14904.9 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 14259.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.78357 0.07409 0.07240

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.330
C2 0.000 1.325 0.357
C3 0.000 -1.325 0.357
O4 0.777 2.159 -0.401
O5 -0.777 -2.159 -0.401
H6 -0.657 1.899 1.022
H7 0.657 -1.899 1.022
H8 1.346 1.596 -0.943
H9 -1.346 -1.596 -0.943

Atom - Atom Distances (Å)
  C1 C2 C3 O4 O5 H6 H7 H8 H9
C11.32541.32542.40852.40852.12522.12522.44572.4457
C21.32542.65021.36943.64961.09673.35671.89123.4696
C31.32542.65023.64961.36943.35671.09673.46961.8912
O42.40851.36943.64964.58972.03734.30210.96654.3484
O52.40853.64961.36944.58974.30212.03734.34840.9665
H62.12521.09673.35672.03734.30214.01852.82244.0686
H72.12523.35671.09674.30212.03734.01854.06862.8224
H82.44571.89123.46960.96654.34842.82244.06864.1768
H92.44573.46961.89124.34840.96654.06862.82244.1768

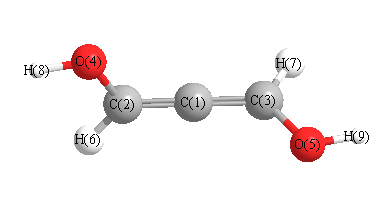
picture of allenediol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O4 126.696 C1 C2 H6 122.382
C1 C3 O5 126.696 C1 C3 H7 122.382
C2 C1 C3 177.636 C2 O4 H8 106.856
C3 O5 H9 106.856 O4 C2 H6 110.921
O5 C3 H7 110.921
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability