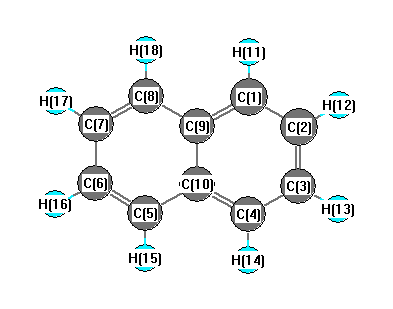
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3134 |
3116 |
0.00 |
|
|
|
| 2 |
Ag |
3107 |
3089 |
0.00 |
|
|
|
| 3 |
Ag |
1582 |
1573 |
0.00 |
|
|
|
| 4 |
Ag |
1441 |
1432 |
0.00 |
|
|
|
| 5 |
Ag |
1400 |
1392 |
0.00 |
|
|
|
| 6 |
Ag |
1134 |
1128 |
0.00 |
|
|
|
| 7 |
Ag |
1026 |
1020 |
0.00 |
|
|
|
| 8 |
Ag |
757 |
752 |
0.00 |
|
|
|
| 9 |
Ag |
506 |
503 |
0.00 |
|
|
|
| 10 |
Au |
962 |
957 |
0.00 |
|
|
|
| 11 |
Au |
826 |
822 |
0.00 |
|
|
|
| 12 |
Au |
619 |
615 |
0.00 |
|
|
|
| 13 |
Au |
181 |
180 |
0.00 |
|
|
|
| 14 |
B1g |
925 |
919 |
0.00 |
|
|
|
| 15 |
B1g |
707 |
703 |
0.00 |
|
|
|
| 16 |
B1g |
382 |
379 |
0.00 |
|
|
|
| 17 |
B1u |
3120 |
3102 |
50.22 |
|
|
|
| 18 |
B1u |
3101 |
3083 |
9.00 |
|
|
|
| 19 |
B1u |
1603 |
1594 |
3.11 |
|
|
|
| 20 |
B1u |
1362 |
1354 |
5.08 |
|
|
|
| 21 |
B1u |
1238 |
1231 |
7.29 |
|
|
|
| 22 |
B1u |
1109 |
1103 |
5.33 |
|
|
|
| 23 |
B1u |
783 |
778 |
0.06 |
|
|
|
| 24 |
B1u |
353 |
351 |
1.19 |
|
|
|
| 25 |
B2g |
968 |
962 |
0.00 |
|
|
|
| 26 |
B2g |
878 |
873 |
0.00 |
|
|
|
| 27 |
B2g |
772 |
768 |
0.00 |
|
|
|
| 28 |
B2g |
466 |
463 |
0.00 |
|
|
|
| 29 |
B2u |
3133 |
3115 |
38.29 |
|
|
|
| 30 |
B2u |
3104 |
3086 |
0.69 |
|
|
|
| 31 |
B2u |
1514 |
1505 |
6.69 |
|
|
|
| 32 |
B2u |
1398 |
1390 |
1.04 |
|
|
|
| 33 |
B2u |
1213 |
1206 |
1.79 |
|
|
|
| 34 |
B2u |
1134 |
1127 |
0.40 |
|
|
|
| 35 |
B2u |
1020 |
1014 |
7.72 |
|
|
|
| 36 |
B2u |
610 |
607 |
3.06 |
|
|
|
| 37 |
B3g |
3120 |
3102 |
0.00 |
|
|
|
| 38 |
B3g |
3100 |
3082 |
0.00 |
|
|
|
| 39 |
B3g |
1637 |
1627 |
0.00 |
|
|
|
| 40 |
B3g |
1445 |
1437 |
0.00 |
|
|
|
| 41 |
B3g |
1219 |
1212 |
0.00 |
|
|
|
| 42 |
B3g |
1129 |
1123 |
0.00 |
|
|
|
| 43 |
B3g |
914 |
909 |
0.00 |
|
|
|
| 44 |
B3g |
499 |
496 |
0.00 |
|
|
|
| 45 |
B3u |
942 |
936 |
2.27 |
|
|
|
| 46 |
B3u |
777 |
772 |
76.57 |
|
|
|
| 47 |
B3u |
472 |
469 |
14.97 |
|
|
|
| 48 |
B3u |
168 |
167 |
1.85 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31493.9 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 31311.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.007 |
|
|
|
| 2 |
C |
0.030 |
|
|
|
| 3 |
C |
0.030 |
|
|
|
| 4 |
C |
0.007 |
|
|
|
| 5 |
C |
0.007 |
|
|
|
| 6 |
C |
0.030 |
|
|
|
| 7 |
C |
0.030 |
|
|
|
| 8 |
C |
0.007 |
|
|
|
| 9 |
C |
0.038 |
|
|
|
| 10 |
C |
0.038 |
|
|
|
| 11 |
H |
-0.035 |
|
|
|
| 12 |
H |
-0.020 |
|
|
|
| 13 |
H |
-0.020 |
|
|
|
| 14 |
H |
-0.035 |
|
|
|
| 15 |
H |
-0.035 |
|
|
|
| 16 |
H |
-0.020 |
|
|
|
| 17 |
H |
-0.020 |
|
|
|
| 18 |
H |
-0.035 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-63.022 |
0.000 |
0.000 |
| y |
0.000 |
-51.292 |
0.000 |
| z |
0.000 |
0.000 |
-51.764 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-11.494 |
0.000 |
0.000 |
| y |
0.000 |
6.101 |
0.000 |
| z |
0.000 |
0.000 |
5.393 |
|
| Polar |
|---|
| 3z2-r2 | 10.786 |
|---|
| x2-y2 | -11.730 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.648 |
0.000 |
0.000 |
| y |
0.000 |
24.521 |
0.000 |
| z |
0.000 |
0.000 |
17.637 |
<r2> (average value of r
2) Å
2
| <r2> |
365.963 |
| (<r2>)1/2 |
19.130 |